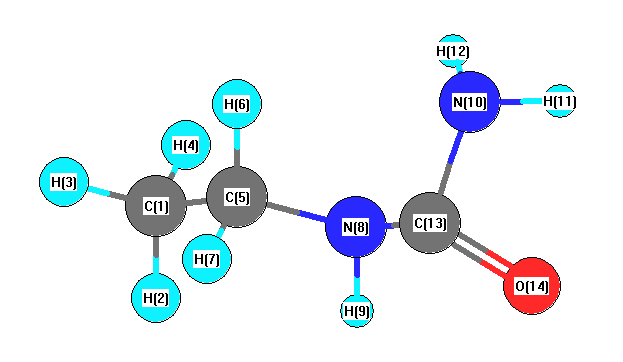
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3690 |
3561 |
29.38 |
|
|
|
| 2 |
A |
3586 |
3460 |
26.23 |
|
|
|
| 3 |
A |
3564 |
3439 |
23.75 |
|
|
|
| 4 |
A |
3132 |
3022 |
20.93 |
|
|
|
| 5 |
A |
3125 |
3015 |
28.78 |
|
|
|
| 6 |
A |
3059 |
2952 |
15.30 |
|
|
|
| 7 |
A |
3018 |
2912 |
40.92 |
|
|
|
| 8 |
A |
2994 |
2889 |
40.80 |
|
|
|
| 9 |
A |
1763 |
1701 |
356.03 |
|
|
|
| 10 |
A |
1670 |
1611 |
119.62 |
|
|
|
| 11 |
A |
1594 |
1538 |
8.28 |
|
|
|
| 12 |
A |
1572 |
1517 |
9.42 |
|
|
|
| 13 |
A |
1562 |
1507 |
7.58 |
|
|
|
| 14 |
A |
1478 |
1426 |
8.36 |
|
|
|
| 15 |
A |
1456 |
1405 |
1.34 |
|
|
|
| 16 |
A |
1411 |
1362 |
279.59 |
|
|
|
| 17 |
A |
1381 |
1333 |
92.84 |
|
|
|
| 18 |
A |
1331 |
1284 |
0.67 |
|
|
|
| 19 |
A |
1203 |
1161 |
16.84 |
|
|
|
| 20 |
A |
1196 |
1154 |
2.45 |
|
|
|
| 21 |
A |
1110 |
1071 |
22.46 |
|
|
|
| 22 |
A |
1058 |
1020 |
17.56 |
|
|
|
| 23 |
A |
981 |
946 |
0.89 |
|
|
|
| 24 |
A |
901 |
870 |
3.22 |
|
|
|
| 25 |
A |
864 |
834 |
0.75 |
|
|
|
| 26 |
A |
796 |
768 |
101.49 |
|
|
|
| 27 |
A |
565 |
545 |
10.28 |
|
|
|
| 28 |
A |
551 |
532 |
138.33 |
|
|
|
| 29 |
A |
538 |
519 |
14.85 |
|
|
|
| 30 |
A |
458 |
442 |
1.55 |
|
|
|
| 31 |
A |
366 |
353 |
1.05 |
|
|
|
| 32 |
A |
337 |
325 |
229.24 |
|
|
|
| 33 |
A |
269 |
259 |
1.89 |
|
|
|
| 34 |
A |
198 |
191 |
5.16 |
|
|
|
| 35 |
A |
72 |
69 |
1.18 |
|
|
|
| 36 |
A |
27 |
26 |
5.64 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26437.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 25509.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.576 |
|
|
-0.434 |
| 2 |
H |
0.203 |
|
|
0.130 |
| 3 |
H |
0.201 |
|
|
0.116 |
| 4 |
H |
0.204 |
|
|
0.131 |
| 5 |
C |
-0.195 |
|
|
0.239 |
| 6 |
H |
0.196 |
|
|
0.026 |
| 7 |
H |
0.192 |
|
|
0.024 |
| 8 |
N |
-0.709 |
|
|
-0.643 |
| 9 |
H |
0.319 |
|
|
0.340 |
| 10 |
N |
-0.780 |
|
|
-0.870 |
| 11 |
H |
0.318 |
|
|
0.386 |
| 12 |
H |
0.307 |
|
|
0.360 |
| 13 |
C |
0.862 |
|
|
0.763 |
| 14 |
O |
-0.541 |
|
|
-0.568 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
3.264 |
2.859 |
-0.115 |
4.341 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
3.277 |
2.848 |
-0.118 |
4.343 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.411 |
-7.178 |
0.097 |
| y |
-7.178 |
-33.312 |
-0.059 |
| z |
0.097 |
-0.059 |
-37.786 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.862 |
-7.178 |
0.097 |
| y |
-7.178 |
4.786 |
-0.059 |
| z |
0.097 |
-0.059 |
-1.925 |
|
| Polar |
|---|
| 3z2-r2 | -3.849 |
|---|
| x2-y2 | -5.098 |
|---|
| xy | -7.178 |
|---|
| xz | 0.097 |
|---|
| yz | -0.059 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.297 |
0.211 |
-0.098 |
| y |
0.211 |
6.822 |
-0.005 |
| z |
-0.098 |
-0.005 |
3.986 |
<r2> (average value of r
2) Å
2
| <r2> |
204.998 |
| (<r2>)1/2 |
14.318 |