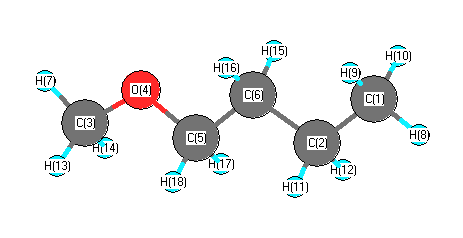
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3127 |
3017 |
29.55 |
|
|
|
| 2 |
A' |
3112 |
3003 |
36.11 |
|
|
|
| 3 |
A' |
3064 |
2956 |
29.59 |
|
|
|
| 4 |
A' |
3046 |
2939 |
21.40 |
|
|
|
| 5 |
A' |
3037 |
2930 |
19.08 |
|
|
|
| 6 |
A' |
2987 |
2883 |
66.53 |
|
|
|
| 7 |
A' |
2970 |
2866 |
27.67 |
|
|
|
| 8 |
A' |
1593 |
1537 |
0.68 |
|
|
|
| 9 |
A' |
1577 |
1521 |
6.93 |
|
|
|
| 10 |
A' |
1572 |
1517 |
13.54 |
|
|
|
| 11 |
A' |
1559 |
1504 |
1.56 |
|
|
|
| 12 |
A' |
1551 |
1497 |
0.30 |
|
|
|
| 13 |
A' |
1510 |
1457 |
1.72 |
|
|
|
| 14 |
A' |
1462 |
1410 |
2.96 |
|
|
|
| 15 |
A' |
1428 |
1378 |
33.19 |
|
|
|
| 16 |
A' |
1400 |
1351 |
0.77 |
|
|
|
| 17 |
A' |
1333 |
1286 |
0.11 |
|
|
|
| 18 |
A' |
1203 |
1161 |
16.73 |
|
|
|
| 19 |
A' |
1143 |
1103 |
2.68 |
|
|
|
| 20 |
A' |
1099 |
1060 |
126.77 |
|
|
|
| 21 |
A' |
1060 |
1023 |
1.64 |
|
|
|
| 22 |
A' |
1010 |
974 |
0.36 |
|
|
|
| 23 |
A' |
941 |
908 |
8.59 |
|
|
|
| 24 |
A' |
891 |
859 |
11.69 |
|
|
|
| 25 |
A' |
481 |
464 |
2.19 |
|
|
|
| 26 |
A' |
374 |
361 |
3.10 |
|
|
|
| 27 |
A' |
309 |
298 |
1.11 |
|
|
|
| 28 |
A' |
133 |
129 |
1.61 |
|
|
|
| 29 |
A" |
3117 |
3008 |
80.11 |
|
|
|
| 30 |
A" |
3103 |
2994 |
0.81 |
|
|
|
| 31 |
A" |
3063 |
2955 |
4.37 |
|
|
|
| 32 |
A" |
3021 |
2915 |
72.53 |
|
|
|
| 33 |
A" |
2997 |
2892 |
52.47 |
|
|
|
| 34 |
A" |
1568 |
1513 |
8.23 |
|
|
|
| 35 |
A" |
1550 |
1495 |
6.97 |
|
|
|
| 36 |
A" |
1363 |
1315 |
0.05 |
|
|
|
| 37 |
A" |
1350 |
1303 |
0.29 |
|
|
|
| 38 |
A" |
1274 |
1229 |
0.12 |
|
|
|
| 39 |
A" |
1206 |
1163 |
4.33 |
|
|
|
| 40 |
A" |
1158 |
1118 |
0.03 |
|
|
|
| 41 |
A" |
995 |
960 |
0.06 |
|
|
|
| 42 |
A" |
850 |
820 |
0.48 |
|
|
|
| 43 |
A" |
768 |
741 |
5.34 |
|
|
|
| 44 |
A" |
252 |
243 |
0.82 |
|
|
|
| 45 |
A" |
234 |
226 |
2.70 |
|
|
|
| 46 |
A" |
135 |
130 |
1.89 |
|
|
|
| 47 |
A" |
104 |
100 |
2.28 |
|
|
|
| 48 |
A" |
69 |
66 |
1.55 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36572.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35289.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.548 |
|
|
-0.315 |
| 2 |
C |
-0.389 |
|
|
0.093 |
| 3 |
C |
-0.309 |
|
|
-0.039 |
| 4 |
O |
-0.486 |
|
|
-0.360 |
| 5 |
C |
-0.099 |
|
|
0.078 |
| 6 |
C |
-0.384 |
|
|
-0.112 |
| 7 |
H |
0.203 |
|
|
0.091 |
| 8 |
H |
0.186 |
|
|
0.084 |
| 9 |
H |
0.188 |
|
|
0.082 |
| 10 |
H |
0.188 |
|
|
0.082 |
| 11 |
H |
0.186 |
|
|
0.000 |
| 12 |
H |
0.186 |
|
|
0.000 |
| 13 |
H |
0.171 |
|
|
0.058 |
| 14 |
H |
0.171 |
|
|
0.058 |
| 15 |
H |
0.200 |
|
|
0.059 |
| 16 |
H |
0.200 |
|
|
0.059 |
| 17 |
H |
0.168 |
|
|
0.042 |
| 18 |
H |
0.168 |
|
|
0.042 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.057 |
-0.809 |
0.000 |
1.331 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
1.034 |
-0.833 |
0.000 |
1.328 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.239 |
1.193 |
0.000 |
| y |
1.193 |
-35.317 |
0.000 |
| z |
0.000 |
0.000 |
-39.609 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.776 |
1.193 |
0.000 |
| y |
1.193 |
5.107 |
0.000 |
| z |
0.000 |
0.000 |
-1.331 |
|
| Polar |
|---|
| 3z2-r2 | -2.662 |
|---|
| x2-y2 | -5.922 |
|---|
| xy | 1.193 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.193 |
1.365 |
0.000 |
| y |
1.365 |
9.525 |
0.000 |
| z |
0.000 |
0.000 |
7.201 |
<r2> (average value of r
2) Å
2
| <r2> |
298.821 |
| (<r2>)1/2 |
17.286 |