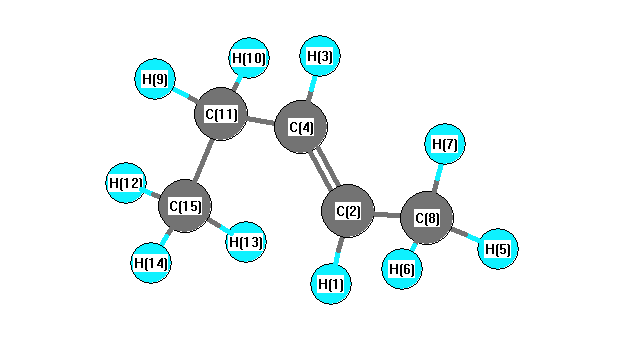
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3165 |
3054 |
26.05 |
|
|
|
| 2 |
A' |
3127 |
3017 |
20.56 |
|
|
|
| 3 |
A' |
3117 |
3008 |
31.57 |
|
|
|
| 4 |
A' |
3110 |
3001 |
12.67 |
|
|
|
| 5 |
A' |
3053 |
2946 |
17.12 |
|
|
|
| 6 |
A' |
3033 |
2926 |
30.81 |
|
|
|
| 7 |
A' |
3022 |
2916 |
25.88 |
|
|
|
| 8 |
A' |
1745 |
1684 |
0.18 |
|
|
|
| 9 |
A' |
1572 |
1517 |
6.47 |
|
|
|
| 10 |
A' |
1560 |
1505 |
10.71 |
|
|
|
| 11 |
A' |
1539 |
1485 |
2.61 |
|
|
|
| 12 |
A' |
1464 |
1412 |
2.66 |
|
|
|
| 13 |
A' |
1460 |
1409 |
7.87 |
|
|
|
| 14 |
A' |
1410 |
1361 |
6.61 |
|
|
|
| 15 |
A' |
1380 |
1332 |
0.55 |
|
|
|
| 16 |
A' |
1349 |
1301 |
1.69 |
|
|
|
| 17 |
A' |
1160 |
1120 |
3.01 |
|
|
|
| 18 |
A' |
1081 |
1043 |
4.18 |
|
|
|
| 19 |
A' |
1062 |
1025 |
1.06 |
|
|
|
| 20 |
A' |
961 |
927 |
8.81 |
|
|
|
| 21 |
A' |
853 |
823 |
0.01 |
|
|
|
| 22 |
A' |
590 |
569 |
1.11 |
|
|
|
| 23 |
A' |
374 |
361 |
0.24 |
|
|
|
| 24 |
A' |
211 |
203 |
0.34 |
|
|
|
| 25 |
A" |
3118 |
3009 |
32.16 |
|
|
|
| 26 |
A" |
3074 |
2966 |
24.08 |
|
|
|
| 27 |
A" |
3039 |
2933 |
16.82 |
|
|
|
| 28 |
A" |
1567 |
1512 |
7.12 |
|
|
|
| 29 |
A" |
1550 |
1496 |
8.01 |
|
|
|
| 30 |
A" |
1338 |
1291 |
0.00 |
|
|
|
| 31 |
A" |
1143 |
1102 |
1.05 |
|
|
|
| 32 |
A" |
1107 |
1068 |
0.61 |
|
|
|
| 33 |
A" |
1021 |
986 |
34.65 |
|
|
|
| 34 |
A" |
856 |
826 |
5.05 |
|
|
|
| 35 |
A" |
731 |
705 |
0.68 |
|
|
|
| 36 |
A" |
298 |
288 |
2.15 |
|
|
|
| 37 |
A" |
220 |
212 |
0.65 |
|
|
|
| 38 |
A" |
199 |
192 |
2.46 |
|
|
|
| 39 |
A" |
117 |
113 |
0.07 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30388.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29321.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.177 |
|
|
0.117 |
| 2 |
C |
-0.176 |
|
|
-0.039 |
| 3 |
H |
0.169 |
|
|
0.138 |
| 4 |
C |
-0.144 |
|
|
-0.258 |
| 5 |
H |
0.198 |
|
|
0.125 |
| 6 |
H |
0.198 |
|
|
0.125 |
| 7 |
H |
0.193 |
|
|
0.128 |
| 8 |
C |
-0.598 |
|
|
-0.411 |
| 9 |
H |
0.199 |
|
|
0.049 |
| 10 |
H |
0.199 |
|
|
0.049 |
| 11 |
C |
-0.428 |
|
|
0.015 |
| 12 |
H |
0.187 |
|
|
0.077 |
| 13 |
H |
0.192 |
|
|
0.077 |
| 14 |
H |
0.192 |
|
|
0.077 |
| 15 |
C |
-0.557 |
|
|
-0.269 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.071 |
-0.009 |
0.000 |
0.071 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.060 |
0.010 |
0.000 |
0.061 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.536 |
0.413 |
0.000 |
| y |
0.413 |
-32.090 |
0.000 |
| z |
0.000 |
0.000 |
-34.548 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.784 |
0.413 |
0.000 |
| y |
0.413 |
0.951 |
0.000 |
| z |
0.000 |
0.000 |
-2.735 |
|
| Polar |
|---|
| 3z2-r2 | -5.470 |
|---|
| x2-y2 | 0.555 |
|---|
| xy | 0.413 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.819 |
0.668 |
0.000 |
| y |
0.668 |
7.383 |
0.000 |
| z |
0.000 |
0.000 |
5.241 |
<r2> (average value of r
2) Å
2
| <r2> |
172.257 |
| (<r2>)1/2 |
13.125 |