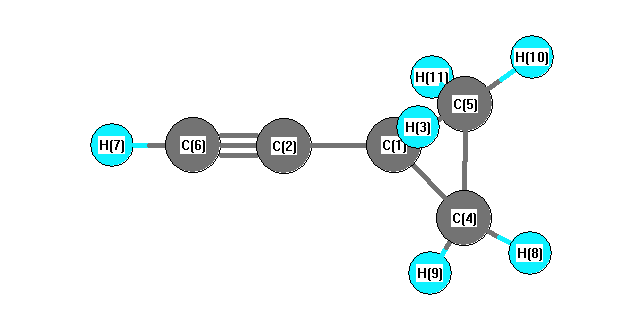
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3501 |
3378 |
65.85 |
|
|
|
| 2 |
A |
3272 |
3157 |
8.53 |
|
|
|
| 3 |
A |
3183 |
3072 |
4.35 |
|
|
|
| 4 |
A |
3173 |
3062 |
4.11 |
|
|
|
| 5 |
A |
2246 |
2167 |
11.98 |
|
|
|
| 6 |
A |
1528 |
1475 |
0.77 |
|
|
|
| 7 |
A |
1390 |
1341 |
2.95 |
|
|
|
| 8 |
A |
1177 |
1135 |
0.66 |
|
|
|
| 9 |
A |
1161 |
1120 |
2.17 |
|
|
|
| 10 |
A |
1084 |
1046 |
14.44 |
|
|
|
| 11 |
A |
938 |
905 |
16.79 |
|
|
|
| 12 |
A |
820 |
791 |
1.44 |
|
|
|
| 13 |
A |
761 |
734 |
5.21 |
|
|
|
| 14 |
A |
738 |
712 |
46.72 |
|
|
|
| 15 |
A |
491 |
474 |
8.51 |
|
|
|
| 16 |
A |
208 |
201 |
1.82 |
|
|
|
| 17 |
A |
3256 |
3141 |
0.04 |
|
|
|
| 18 |
A |
3170 |
3058 |
11.42 |
|
|
|
| 19 |
A |
1514 |
1461 |
5.14 |
|
|
|
| 20 |
A |
1218 |
1175 |
0.68 |
|
|
|
| 21 |
A |
1137 |
1097 |
0.03 |
|
|
|
| 22 |
A |
1108 |
1069 |
13.18 |
|
|
|
| 23 |
A |
900 |
869 |
0.64 |
|
|
|
| 24 |
A |
806 |
778 |
12.60 |
|
|
|
| 25 |
A |
701 |
677 |
55.14 |
|
|
|
| 26 |
A |
537 |
518 |
3.52 |
|
|
|
| 27 |
A |
219 |
212 |
2.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20117.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 19411.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.434 |
|
|
-0.127 |
| 2 |
C |
0.183 |
|
|
0.038 |
| 3 |
H |
0.228 |
|
|
0.175 |
| 4 |
C |
-0.356 |
|
|
-0.322 |
| 5 |
C |
-0.356 |
|
|
-0.322 |
| 6 |
C |
-0.361 |
|
|
-0.401 |
| 7 |
H |
0.239 |
|
|
0.273 |
| 8 |
H |
0.210 |
|
|
0.160 |
| 9 |
H |
0.219 |
|
|
0.182 |
| 10 |
H |
0.210 |
|
|
0.160 |
| 11 |
H |
0.219 |
|
|
0.182 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.812 |
0.337 |
0.000 |
0.879 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.810 |
0.356 |
0.000 |
0.885 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.797 |
2.696 |
0.000 |
| y |
2.696 |
-27.347 |
0.000 |
| z |
0.000 |
0.000 |
-31.327 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.540 |
2.696 |
0.000 |
| y |
2.696 |
2.715 |
0.000 |
| z |
0.000 |
0.000 |
-3.254 |
|
| Polar |
|---|
| 3z2-r2 | -6.509 |
|---|
| x2-y2 | -1.450 |
|---|
| xy | 2.696 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.305 |
2.330 |
0.000 |
| y |
2.330 |
6.055 |
0.000 |
| z |
0.000 |
0.000 |
4.676 |
<r2> (average value of r
2) Å
2
| <r2> |
120.795 |
| (<r2>)1/2 |
10.991 |