Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -747.291470 |
| Energy at 298.15K | -747.292662 |
| HF Energy | -747.291470 |
| Nuclear repulsion energy | 558.595494 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
1840 |
1775 |
0.00 |
|
|
|
| 2 |
Ag |
1449 |
1399 |
0.00 |
|
|
|
| 3 |
Ag |
1351 |
1304 |
0.00 |
|
|
|
| 4 |
Ag |
1213 |
1170 |
0.00 |
|
|
|
| 5 |
Ag |
707 |
682 |
0.00 |
|
|
|
| 6 |
Ag |
588 |
567 |
0.00 |
|
|
|
| 7 |
Ag |
371 |
358 |
0.00 |
|
|
|
| 8 |
Ag |
341 |
329 |
0.00 |
|
|
|
| 9 |
Ag |
217 |
210 |
0.00 |
|
|
|
| 10 |
Au |
605 |
584 |
3.34 |
|
|
|
| 11 |
Au |
356 |
344 |
4.87 |
|
|
|
| 12 |
Au |
119 |
115 |
0.03 |
|
|
|
| 13 |
Au |
17 |
16 |
0.03 |
|
|
|
| 14 |
Bg |
771 |
744 |
0.00 |
|
|
|
| 15 |
Bg |
489 |
472 |
0.00 |
|
|
|
| 16 |
Bg |
178 |
171 |
0.00 |
|
|
|
| 17 |
Bu |
1803 |
1740 |
247.37 |
|
|
|
| 18 |
Bu |
1365 |
1317 |
267.04 |
|
|
|
| 19 |
Bu |
1244 |
1201 |
166.62 |
|
|
|
| 20 |
Bu |
957 |
924 |
195.61 |
|
|
|
| 21 |
Bu |
607 |
586 |
2.48 |
|
|
|
| 22 |
Bu |
475 |
459 |
3.52 |
|
|
|
| 23 |
Bu |
279 |
269 |
5.76 |
|
|
|
| 24 |
Bu |
136 |
131 |
1.44 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8738.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 8431.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.164 |
1.830 |
0.000 |
| C2 |
0.477 |
0.536 |
0.000 |
| C3 |
-0.477 |
-0.536 |
0.000 |
| C4 |
-0.164 |
-1.830 |
0.000 |
| F5 |
1.091 |
2.809 |
0.000 |
| F6 |
-1.091 |
2.310 |
0.000 |
| F7 |
1.799 |
0.174 |
0.000 |
| F8 |
-1.799 |
-0.174 |
0.000 |
| F9 |
1.091 |
-2.310 |
0.000 |
| F10 |
-1.091 |
-2.809 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3311 | 2.4519 | 3.6749 | 1.3477 | 1.3435 | 2.3267 | 2.8054 | 4.2422 | 4.8057 |
C2 | 1.3311 | | 1.4360 | 2.4519 | 2.3538 | 2.3673 | 1.3700 | 2.3844 | 2.9114 | 3.6945 | C3 | 2.4519 | 1.4360 | | 1.3311 | 3.6945 | 2.9114 | 2.3844 | 1.3700 | 2.3673 | 2.3538 | C4 | 3.6749 | 2.4519 | 1.3311 | | 4.8057 | 4.2422 | 2.8054 | 2.3267 | 1.3435 | 1.3477 | F5 | 1.3477 | 2.3538 | 3.6945 | 4.8057 | | 2.2380 | 2.7280 | 4.1530 | 5.1185 | 6.0263 | F6 | 1.3435 | 2.3673 | 2.9114 | 4.2422 | 2.2380 | | 3.5930 | 2.5828 | 5.1086 | 5.1185 | F7 | 2.3267 | 1.3700 | 2.3844 | 2.8054 | 2.7280 | 3.5930 | | 3.6142 | 2.5828 | 4.1530 | F8 | 2.8054 | 2.3844 | 1.3700 | 2.3267 | 4.1530 | 2.5828 | 3.6142 | | 3.5930 | 2.7280 | F9 | 4.2422 | 2.9114 | 2.3673 | 1.3435 | 5.1185 | 5.1086 | 2.5828 | 3.5930 | | 2.2380 | F10 | 4.8057 | 3.6945 | 2.3538 | 1.3477 | 6.0263 | 5.1185 | 4.1530 | 2.7280 | 2.2380 | |
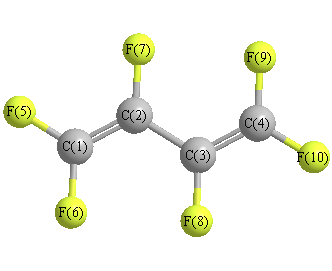 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.724 |
|
C1 |
C2 |
F7 |
118.931 |
| C2 |
C1 |
F5 |
122.961 |
|
C2 |
C1 |
F6 |
124.518 |
| C2 |
C3 |
C4 |
124.724 |
|
C2 |
C3 |
F8 |
116.345 |
| C3 |
C2 |
F7 |
116.345 |
|
C3 |
C4 |
F9 |
124.518 |
| C3 |
C4 |
F10 |
122.961 |
|
C4 |
C3 |
F8 |
118.931 |
| F5 |
C1 |
F6 |
112.521 |
|
F9 |
C4 |
F10 |
112.521 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.604 |
|
|
|
| 2 |
C |
0.143 |
|
|
|
| 3 |
C |
0.143 |
|
|
|
| 4 |
C |
0.604 |
|
|
|
| 5 |
F |
-0.248 |
|
|
|
| 6 |
F |
-0.243 |
|
|
|
| 7 |
F |
-0.255 |
|
|
|
| 8 |
F |
-0.255 |
|
|
|
| 9 |
F |
-0.243 |
|
|
|
| 10 |
F |
-0.248 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-55.122 |
-0.042 |
0.000 |
| y |
-0.042 |
-53.238 |
0.000 |
| z |
0.000 |
0.000 |
-48.647 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.180 |
-0.042 |
0.000 |
| y |
-0.042 |
-1.353 |
0.000 |
| z |
0.000 |
0.000 |
5.533 |
|
| Polar |
|---|
| 3z2-r2 | 11.066 |
|---|
| x2-y2 | -1.884 |
|---|
| xy | -0.042 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
5.513 |
-0.144 |
0.000 |
| y |
-0.144 |
11.147 |
0.000 |
| z |
0.000 |
0.000 |
1.988 |
<r2> (average value of r
2) Å
2
| <r2> |
419.037 |
| (<r2>)1/2 |
20.470 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -747.285407 |
| Energy at 298.15K | -747.286576 |
| HF Energy | -747.285407 |
| Nuclear repulsion energy | 564.832819 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
1821 |
1757 |
51.09 |
|
|
|
| 2 |
A |
1394 |
1346 |
21.34 |
|
|
|
| 3 |
A |
1372 |
1324 |
112.90 |
|
|
|
| 4 |
A |
1149 |
1109 |
170.34 |
|
|
|
| 5 |
A |
710 |
686 |
0.04 |
|
|
|
| 6 |
A |
690 |
666 |
1.37 |
|
|
|
| 7 |
A |
522 |
504 |
0.19 |
|
|
|
| 8 |
A |
462 |
446 |
0.14 |
|
|
|
| 9 |
A |
373 |
360 |
1.18 |
|
|
|
| 10 |
A |
254 |
245 |
0.37 |
|
|
|
| 11 |
A |
201 |
194 |
0.42 |
|
|
|
| 12 |
A |
120 |
116 |
0.27 |
|
|
|
| 13 |
A |
60 |
58 |
0.02 |
|
|
|
| 14 |
B |
1812 |
1748 |
170.47 |
|
|
|
| 15 |
B |
1351 |
1304 |
178.62 |
|
|
|
| 16 |
B |
1236 |
1193 |
47.80 |
|
|
|
| 17 |
B |
971 |
937 |
115.56 |
|
|
|
| 18 |
B |
608 |
586 |
3.61 |
|
|
|
| 19 |
B |
602 |
581 |
3.37 |
|
|
|
| 20 |
B |
556 |
536 |
2.23 |
|
|
|
| 21 |
B |
385 |
371 |
3.82 |
|
|
|
| 22 |
B |
300 |
289 |
2.80 |
|
|
|
| 23 |
B |
206 |
199 |
4.93 |
|
|
|
| 24 |
B |
114 |
110 |
0.58 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8635.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 8332.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.077 |
1.505 |
-0.392 |
| C2 |
0.224 |
0.685 |
0.644 |
| C3 |
-0.224 |
-0.685 |
0.644 |
| C4 |
-0.077 |
-1.505 |
-0.392 |
| F5 |
0.541 |
2.766 |
-0.431 |
| F6 |
-0.541 |
1.146 |
-1.532 |
| F7 |
0.809 |
1.154 |
1.795 |
| F8 |
-0.809 |
-1.154 |
1.795 |
| F9 |
0.541 |
-1.146 |
-1.532 |
| F10 |
-0.541 |
-2.766 |
-0.431 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
F5 |
F6 |
F7 |
F8 |
F9 |
F10 |
| C1 | | 1.3291 | 2.4415 | 3.0142 | 1.3442 | 1.3461 | 2.3325 | 3.5555 | 2.9230 | 4.3160 |
C2 | 1.3291 | | 1.4420 | 2.4415 | 2.3635 | 2.3525 | 1.3738 | 2.4037 | 2.8618 | 3.6950 | C3 | 2.4415 | 1.4420 | | 1.3291 | 3.6950 | 2.8618 | 2.4037 | 1.3738 | 2.3525 | 2.3635 | C4 | 3.0142 | 2.4415 | 1.3291 | | 4.3160 | 2.9230 | 3.5555 | 2.3325 | 1.3461 | 1.3442 | F5 | 1.3442 | 2.3635 | 3.6950 | 4.3160 | | 2.2380 | 2.7611 | 4.7063 | 4.0643 | 5.6373 | F6 | 1.3461 | 2.3525 | 2.8618 | 2.9230 | 2.2380 | | 3.5908 | 4.0540 | 2.5343 | 4.0643 | F7 | 2.3325 | 1.3738 | 2.4037 | 3.5555 | 2.7611 | 3.5908 | | 2.8199 | 4.0540 | 4.7063 | F8 | 3.5555 | 2.4037 | 1.3738 | 2.3325 | 4.7063 | 4.0540 | 2.8199 | | 3.5908 | 2.7611 | F9 | 2.9230 | 2.8618 | 2.3525 | 1.3461 | 4.0643 | 2.5343 | 4.0540 | 3.5908 | | 2.2380 | F10 | 4.3160 | 3.6950 | 2.3635 | 1.3442 | 5.6373 | 4.0643 | 4.7063 | 2.7611 | 2.2380 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.495 |
|
C1 |
C2 |
F7 |
119.291 |
| C2 |
C1 |
F5 |
124.281 |
|
C2 |
C1 |
F6 |
123.125 |
| C2 |
C3 |
C4 |
123.495 |
|
C2 |
C3 |
F8 |
117.203 |
| C3 |
C2 |
F7 |
117.203 |
|
C3 |
C4 |
F9 |
123.125 |
| C3 |
C4 |
F10 |
124.281 |
|
C4 |
C3 |
F8 |
119.291 |
| F5 |
C1 |
F6 |
112.586 |
|
F9 |
C4 |
F10 |
112.586 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.580 |
|
|
|
| 2 |
C |
0.168 |
|
|
|
| 3 |
C |
0.168 |
|
|
|
| 4 |
C |
0.580 |
|
|
|
| 5 |
F |
-0.246 |
|
|
|
| 6 |
F |
-0.249 |
|
|
|
| 7 |
F |
-0.254 |
|
|
|
| 8 |
F |
-0.254 |
|
|
|
| 9 |
F |
-0.249 |
|
|
|
| 10 |
F |
-0.246 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.660 |
0.660 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-50.016 |
-1.627 |
0.000 |
| y |
-1.627 |
-52.960 |
0.000 |
| z |
0.000 |
0.000 |
-54.544 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.736 |
-1.627 |
0.000 |
| y |
-1.627 |
-0.681 |
0.000 |
| z |
0.000 |
0.000 |
-3.056 |
|
| Polar |
|---|
| 3z2-r2 | -6.111 |
|---|
| x2-y2 | 2.944 |
|---|
| xy | -1.627 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
2.841 |
0.862 |
0.000 |
| y |
0.862 |
8.119 |
0.000 |
| z |
0.000 |
0.000 |
6.347 |
<r2> (average value of r
2) Å
2
| <r2> |
384.395 |
| (<r2>)1/2 |
19.606 |
