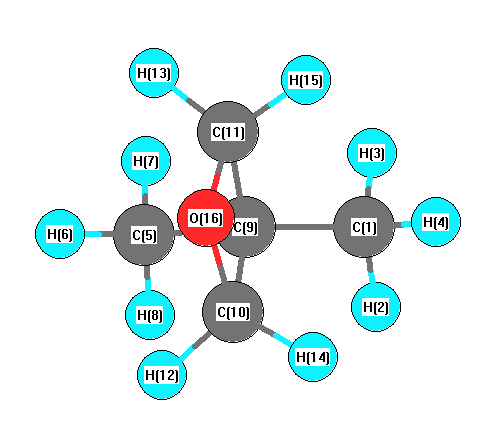
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3113 |
3004 |
61.51 |
|
|
|
| 2 |
A' |
3111 |
3002 |
42.93 |
|
|
|
| 3 |
A' |
3103 |
2994 |
24.25 |
|
|
|
| 4 |
A' |
3057 |
2950 |
5.60 |
|
|
|
| 5 |
A' |
3042 |
2935 |
22.73 |
|
|
|
| 6 |
A' |
3038 |
2932 |
17.32 |
|
|
|
| 7 |
A' |
1588 |
1532 |
0.95 |
|
|
|
| 8 |
A' |
1567 |
1512 |
5.21 |
|
|
|
| 9 |
A' |
1553 |
1499 |
4.61 |
|
|
|
| 10 |
A' |
1471 |
1420 |
3.73 |
|
|
|
| 11 |
A' |
1452 |
1401 |
12.44 |
|
|
|
| 12 |
A' |
1363 |
1315 |
2.32 |
|
|
|
| 13 |
A' |
1308 |
1262 |
1.64 |
|
|
|
| 14 |
A' |
1274 |
1229 |
5.76 |
|
|
|
| 15 |
A' |
1105 |
1066 |
0.07 |
|
|
|
| 16 |
A' |
1014 |
979 |
0.43 |
|
|
|
| 17 |
A' |
1004 |
969 |
1.63 |
|
|
|
| 18 |
A' |
944 |
911 |
0.36 |
|
|
|
| 19 |
A' |
916 |
884 |
2.22 |
|
|
|
| 20 |
A' |
821 |
792 |
11.25 |
|
|
|
| 21 |
A' |
624 |
602 |
0.55 |
|
|
|
| 22 |
A' |
406 |
392 |
0.03 |
|
|
|
| 23 |
A' |
344 |
332 |
0.32 |
|
|
|
| 24 |
A' |
20 |
20 |
5.52 |
|
|
|
| 25 |
A" |
3105 |
2996 |
42.44 |
|
|
|
| 26 |
A" |
3104 |
2995 |
4.51 |
|
|
|
| 27 |
A" |
3097 |
2989 |
0.84 |
|
|
|
| 28 |
A" |
3044 |
2937 |
94.62 |
|
|
|
| 29 |
A" |
1576 |
1521 |
12.27 |
|
|
|
| 30 |
A" |
1557 |
1502 |
0.37 |
|
|
|
| 31 |
A" |
1552 |
1498 |
0.00 |
|
|
|
| 32 |
A" |
1294 |
1248 |
10.91 |
|
|
|
| 33 |
A" |
1242 |
1199 |
2.32 |
|
|
|
| 34 |
A" |
1131 |
1091 |
0.00 |
|
|
|
| 35 |
A" |
1087 |
1049 |
0.04 |
|
|
|
| 36 |
A" |
966 |
932 |
5.66 |
|
|
|
| 37 |
A" |
958 |
925 |
44.57 |
|
|
|
| 38 |
A" |
877 |
846 |
8.45 |
|
|
|
| 39 |
A" |
390 |
376 |
0.45 |
|
|
|
| 40 |
A" |
321 |
309 |
0.09 |
|
|
|
| 41 |
A" |
274 |
264 |
0.02 |
|
|
|
| 42 |
A" |
239 |
231 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31524.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30418.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.523 |
|
|
-0.530 |
| 2 |
H |
0.189 |
|
|
0.142 |
| 3 |
H |
0.189 |
|
|
0.142 |
| 4 |
H |
0.197 |
|
|
0.147 |
| 5 |
C |
-0.534 |
|
|
-0.579 |
| 6 |
H |
0.192 |
|
|
0.155 |
| 7 |
H |
0.191 |
|
|
0.155 |
| 8 |
H |
0.191 |
|
|
0.155 |
| 9 |
C |
-0.226 |
|
|
0.209 |
| 10 |
C |
-0.071 |
|
|
0.187 |
| 11 |
C |
-0.071 |
|
|
0.187 |
| 12 |
H |
0.185 |
|
|
0.018 |
| 13 |
H |
0.185 |
|
|
0.018 |
| 14 |
H |
0.187 |
|
|
0.027 |
| 15 |
H |
0.187 |
|
|
0.027 |
| 16 |
O |
-0.465 |
|
|
-0.462 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.042 |
2.345 |
0.000 |
2.345 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.037 |
2.346 |
0.000 |
2.346 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.197 |
-0.015 |
0.002 |
| y |
-0.015 |
-37.488 |
-0.001 |
| z |
0.002 |
-0.001 |
-35.637 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-7.634 |
-0.015 |
0.002 |
| y |
-0.015 |
2.429 |
-0.001 |
| z |
0.002 |
-0.001 |
5.205 |
|
| Polar |
|---|
| 3z2-r2 | 10.410 |
|---|
| x2-y2 | -6.709 |
|---|
| xy | -0.015 |
|---|
| xz | 0.002 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.284 |
0.002 |
-0.001 |
| y |
0.002 |
7.340 |
-0.001 |
| z |
-0.001 |
-0.001 |
7.979 |
<r2> (average value of r
2) Å
2
| <r2> |
161.730 |
| (<r2>)1/2 |
12.717 |