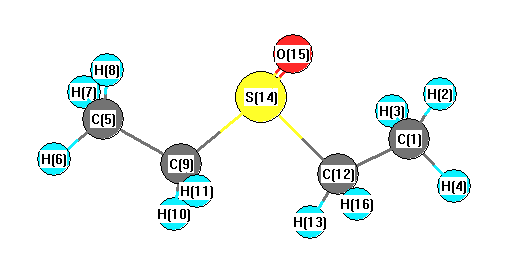
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3185 |
3073 |
9.19 |
|
|
|
| 2 |
A |
3173 |
3062 |
6.82 |
|
|
|
| 3 |
A |
3134 |
3024 |
15.33 |
|
|
|
| 4 |
A |
3122 |
3013 |
10.84 |
|
|
|
| 5 |
A |
3117 |
3008 |
17.31 |
|
|
|
| 6 |
A |
3115 |
3006 |
6.44 |
|
|
|
| 7 |
A |
3104 |
2995 |
2.86 |
|
|
|
| 8 |
A |
3096 |
2987 |
10.63 |
|
|
|
| 9 |
A |
3056 |
2949 |
15.26 |
|
|
|
| 10 |
A |
2950 |
2847 |
28.99 |
|
|
|
| 11 |
A |
1588 |
1532 |
13.44 |
|
|
|
| 12 |
A |
1567 |
1512 |
19.30 |
|
|
|
| 13 |
A |
1565 |
1510 |
6.25 |
|
|
|
| 14 |
A |
1563 |
1508 |
13.43 |
|
|
|
| 15 |
A |
1526 |
1472 |
4.60 |
|
|
|
| 16 |
A |
1515 |
1461 |
6.01 |
|
|
|
| 17 |
A |
1482 |
1430 |
36.74 |
|
|
|
| 18 |
A |
1468 |
1416 |
5.76 |
|
|
|
| 19 |
A |
1293 |
1247 |
4.32 |
|
|
|
| 20 |
A |
1287 |
1242 |
1.02 |
|
|
|
| 21 |
A |
1279 |
1234 |
4.83 |
|
|
|
| 22 |
A |
1257 |
1213 |
8.06 |
|
|
|
| 23 |
A |
1093 |
1054 |
13.28 |
|
|
|
| 24 |
A |
1076 |
1038 |
7.66 |
|
|
|
| 25 |
A |
1059 |
1022 |
6.10 |
|
|
|
| 26 |
A |
1028 |
992 |
0.80 |
|
|
|
| 27 |
A |
1003 |
968 |
3.07 |
|
|
|
| 28 |
A |
985 |
950 |
4.22 |
|
|
|
| 29 |
A |
862 |
832 |
37.32 |
|
|
|
| 30 |
A |
822 |
793 |
4.82 |
|
|
|
| 31 |
A |
788 |
761 |
5.36 |
|
|
|
| 32 |
A |
580 |
560 |
10.39 |
|
|
|
| 33 |
A |
494 |
476 |
0.95 |
|
|
|
| 34 |
A |
464 |
448 |
3.51 |
|
|
|
| 35 |
A |
327 |
316 |
1.05 |
|
|
|
| 36 |
A |
290 |
280 |
10.94 |
|
|
|
| 37 |
A |
265 |
256 |
1.15 |
|
|
|
| 38 |
A |
251 |
242 |
2.87 |
|
|
|
| 39 |
A |
193 |
186 |
1.02 |
|
|
|
| 40 |
A |
166 |
160 |
3.21 |
|
|
|
| 41 |
A |
111 |
107 |
2.42 |
|
|
|
| 42 |
A |
85 |
82 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30190.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29130.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.598 |
|
|
-0.196 |
| 2 |
H |
0.203 |
|
|
0.063 |
| 3 |
H |
0.189 |
|
|
0.067 |
| 4 |
H |
0.246 |
|
|
0.094 |
| 5 |
C |
-0.583 |
|
|
-0.299 |
| 6 |
H |
0.213 |
|
|
0.093 |
| 7 |
H |
0.217 |
|
|
0.099 |
| 8 |
H |
0.201 |
|
|
0.114 |
| 9 |
C |
-0.653 |
|
|
-0.101 |
| 10 |
H |
0.247 |
|
|
0.127 |
| 11 |
H |
0.241 |
|
|
0.105 |
| 12 |
C |
-0.632 |
|
|
-0.103 |
| 13 |
H |
0.224 |
|
|
0.062 |
| 14 |
S |
0.826 |
|
|
0.163 |
| 15 |
O |
-0.575 |
|
|
-0.401 |
| 16 |
H |
0.234 |
|
|
0.114 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.401 |
-3.186 |
-0.515 |
4.022 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
2.417 |
-3.236 |
-0.535 |
4.075 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.273 |
3.861 |
2.157 |
| y |
3.861 |
-50.219 |
-0.422 |
| z |
2.157 |
-0.422 |
-45.161 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.417 |
3.861 |
2.157 |
| y |
3.861 |
-5.502 |
-0.422 |
| z |
2.157 |
-0.422 |
2.085 |
|
| Polar |
|---|
| 3z2-r2 | 4.170 |
|---|
| x2-y2 | 5.946 |
|---|
| xy | 3.861 |
|---|
| xz | 2.157 |
|---|
| yz | -0.422 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.262 |
0.110 |
0.408 |
| y |
0.110 |
9.301 |
0.287 |
| z |
0.408 |
0.287 |
7.588 |
<r2> (average value of r
2) Å
2
| <r2> |
238.969 |
| (<r2>)1/2 |
15.459 |