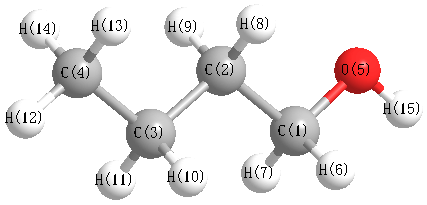
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3468 |
3346 |
1.78 |
|
|
|
| 2 |
A' |
3112 |
3003 |
35.99 |
|
|
|
| 3 |
A' |
3063 |
2956 |
30.45 |
|
|
|
| 4 |
A' |
3046 |
2939 |
21.25 |
|
|
|
| 5 |
A' |
3036 |
2929 |
20.63 |
|
|
|
| 6 |
A' |
2974 |
2870 |
48.21 |
|
|
|
| 7 |
A' |
1592 |
1536 |
2.78 |
|
|
|
| 8 |
A' |
1573 |
1518 |
8.06 |
|
|
|
| 9 |
A' |
1559 |
1505 |
1.85 |
|
|
|
| 10 |
A' |
1552 |
1497 |
0.14 |
|
|
|
| 11 |
A' |
1474 |
1422 |
3.33 |
|
|
|
| 12 |
A' |
1462 |
1411 |
4.86 |
|
|
|
| 13 |
A' |
1404 |
1355 |
3.23 |
|
|
|
| 14 |
A' |
1353 |
1305 |
13.10 |
|
|
|
| 15 |
A' |
1265 |
1221 |
47.45 |
|
|
|
| 16 |
A' |
1133 |
1093 |
0.53 |
|
|
|
| 17 |
A' |
1050 |
1013 |
0.01 |
|
|
|
| 18 |
A' |
1017 |
982 |
59.56 |
|
|
|
| 19 |
A' |
985 |
950 |
0.90 |
|
|
|
| 20 |
A' |
909 |
877 |
22.54 |
|
|
|
| 21 |
A' |
426 |
411 |
14.49 |
|
|
|
| 22 |
A' |
400 |
386 |
0.31 |
|
|
|
| 23 |
A' |
183 |
177 |
3.56 |
|
|
|
| 24 |
A" |
3116 |
3007 |
83.22 |
|
|
|
| 25 |
A" |
3103 |
2994 |
1.54 |
|
|
|
| 26 |
A" |
3061 |
2953 |
4.70 |
|
|
|
| 27 |
A" |
2997 |
2892 |
63.64 |
|
|
|
| 28 |
A" |
1567 |
1512 |
8.20 |
|
|
|
| 29 |
A" |
1366 |
1318 |
0.02 |
|
|
|
| 30 |
A" |
1357 |
1310 |
1.80 |
|
|
|
| 31 |
A" |
1273 |
1228 |
0.28 |
|
|
|
| 32 |
A" |
1191 |
1149 |
1.32 |
|
|
|
| 33 |
A" |
996 |
961 |
0.00 |
|
|
|
| 34 |
A" |
851 |
821 |
1.47 |
|
|
|
| 35 |
A" |
769 |
742 |
4.02 |
|
|
|
| 36 |
A" |
300 |
289 |
134.34 |
|
|
|
| 37 |
A" |
248 |
240 |
0.72 |
|
|
|
| 38 |
A" |
119 |
115 |
6.78 |
|
|
|
| 39 |
A" |
112 |
108 |
0.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30231.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29170.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.130 |
|
|
|
| 2 |
C |
-0.372 |
|
|
|
| 3 |
C |
-0.390 |
|
|
|
| 4 |
C |
-0.547 |
|
|
|
| 5 |
O |
-0.552 |
|
|
|
| 6 |
H |
0.165 |
|
|
|
| 7 |
H |
0.165 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.185 |
|
|
|
| 11 |
H |
0.185 |
|
|
|
| 12 |
H |
0.185 |
|
|
|
| 13 |
H |
0.188 |
|
|
|
| 14 |
H |
0.188 |
|
|
|
| 15 |
H |
0.332 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.375 |
-1.582 |
0.000 |
1.626 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-28.521 |
-3.553 |
0.000 |
| y |
-3.553 |
-34.788 |
0.000 |
| z |
0.000 |
0.000 |
-32.982 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.364 |
-3.553 |
0.000 |
| y |
-3.553 |
-4.036 |
0.000 |
| z |
0.000 |
0.000 |
-1.328 |
|
| Polar |
|---|
| 3z2-r2 | -2.655 |
|---|
| x2-y2 | 6.267 |
|---|
| xy | -3.553 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
8.045 |
0.127 |
0.000 |
| y |
0.127 |
6.162 |
0.000 |
| z |
0.000 |
0.000 |
5.766 |
<r2> (average value of r
2) Å
2
| <r2> |
188.630 |
| (<r2>)1/2 |
13.734 |