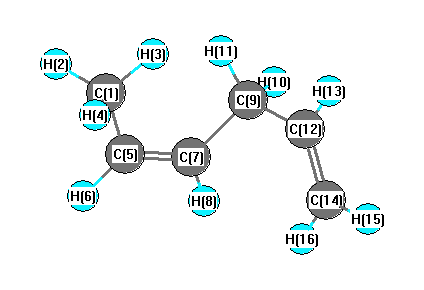
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3239 |
3125 |
16.87 |
|
|
|
| 2 |
A |
3162 |
3051 |
24.49 |
|
|
|
| 3 |
A |
3161 |
3050 |
21.45 |
|
|
|
| 4 |
A |
3145 |
3034 |
24.31 |
|
|
|
| 5 |
A |
3141 |
3031 |
12.74 |
|
|
|
| 6 |
A |
3128 |
3018 |
1.26 |
|
|
|
| 7 |
A |
3077 |
2969 |
19.65 |
|
|
|
| 8 |
A |
3067 |
2959 |
14.27 |
|
|
|
| 9 |
A |
3038 |
2932 |
19.79 |
|
|
|
| 10 |
A |
3022 |
2916 |
22.37 |
|
|
|
| 11 |
A |
1733 |
1672 |
0.64 |
|
|
|
| 12 |
A |
1718 |
1657 |
9.98 |
|
|
|
| 13 |
A |
1560 |
1505 |
10.22 |
|
|
|
| 14 |
A |
1555 |
1501 |
7.64 |
|
|
|
| 15 |
A |
1533 |
1479 |
8.41 |
|
|
|
| 16 |
A |
1496 |
1444 |
1.86 |
|
|
|
| 17 |
A |
1468 |
1416 |
7.54 |
|
|
|
| 18 |
A |
1463 |
1411 |
4.20 |
|
|
|
| 19 |
A |
1374 |
1326 |
2.59 |
|
|
|
| 20 |
A |
1353 |
1305 |
1.20 |
|
|
|
| 21 |
A |
1333 |
1287 |
0.75 |
|
|
|
| 22 |
A |
1279 |
1234 |
2.58 |
|
|
|
| 23 |
A |
1150 |
1110 |
1.07 |
|
|
|
| 24 |
A |
1127 |
1088 |
9.62 |
|
|
|
| 25 |
A |
1104 |
1065 |
0.06 |
|
|
|
| 26 |
A |
1050 |
1013 |
12.82 |
|
|
|
| 27 |
A |
1034 |
997 |
0.23 |
|
|
|
| 28 |
A |
1016 |
980 |
2.18 |
|
|
|
| 29 |
A |
980 |
945 |
40.24 |
|
|
|
| 30 |
A |
957 |
924 |
8.82 |
|
|
|
| 31 |
A |
913 |
881 |
5.85 |
|
|
|
| 32 |
A |
878 |
847 |
4.89 |
|
|
|
| 33 |
A |
738 |
712 |
33.09 |
|
|
|
| 34 |
A |
621 |
599 |
9.95 |
|
|
|
| 35 |
A |
577 |
557 |
3.40 |
|
|
|
| 36 |
A |
536 |
517 |
15.09 |
|
|
|
| 37 |
A |
401 |
387 |
0.24 |
|
|
|
| 38 |
A |
299 |
289 |
0.05 |
|
|
|
| 39 |
A |
213 |
205 |
0.07 |
|
|
|
| 40 |
A |
140 |
135 |
0.45 |
|
|
|
| 41 |
A |
124 |
120 |
0.04 |
|
|
|
| 42 |
A |
40 |
39 |
0.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31469.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30364.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.610 |
|
|
-0.217 |
| 2 |
H |
0.201 |
|
|
0.074 |
| 3 |
H |
0.201 |
|
|
0.094 |
| 4 |
H |
0.204 |
|
|
0.084 |
| 5 |
C |
-0.165 |
|
|
-0.060 |
| 6 |
H |
0.179 |
|
|
0.101 |
| 7 |
C |
-0.148 |
|
|
-0.349 |
| 8 |
H |
0.184 |
|
|
0.164 |
| 9 |
C |
-0.490 |
|
|
0.120 |
| 10 |
H |
0.212 |
|
|
0.040 |
| 11 |
H |
0.209 |
|
|
0.039 |
| 12 |
C |
-0.148 |
|
|
-0.139 |
| 13 |
H |
0.180 |
|
|
0.111 |
| 14 |
C |
-0.378 |
|
|
-0.401 |
| 15 |
H |
0.179 |
|
|
0.168 |
| 16 |
H |
0.190 |
|
|
0.170 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.341 |
0.225 |
-0.181 |
0.446 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.334 |
0.239 |
-0.197 |
0.456 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.801 |
1.349 |
-0.163 |
| y |
1.349 |
-37.592 |
-1.172 |
| z |
-0.163 |
-1.172 |
-38.665 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.328 |
1.349 |
-0.163 |
| y |
1.349 |
0.141 |
-1.172 |
| z |
-0.163 |
-1.172 |
-1.469 |
|
| Polar |
|---|
| 3z2-r2 | -2.937 |
|---|
| x2-y2 | 0.791 |
|---|
| xy | 1.349 |
|---|
| xz | -0.163 |
|---|
| yz | -1.172 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.906 |
0.535 |
0.185 |
| y |
0.535 |
8.448 |
-0.537 |
| z |
0.185 |
-0.537 |
6.495 |
<r2> (average value of r
2) Å
2
| <r2> |
211.051 |
| (<r2>)1/2 |
14.528 |