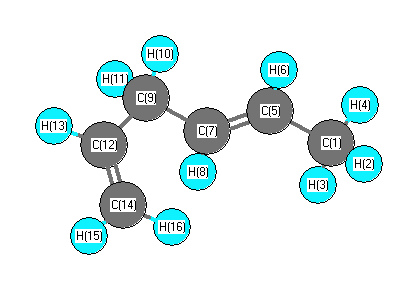
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3238 |
3125 |
17.61 |
|
|
|
| 2 |
A |
3162 |
3051 |
8.22 |
|
|
|
| 3 |
A |
3150 |
3039 |
36.46 |
|
|
|
| 4 |
A |
3144 |
3034 |
26.79 |
|
|
|
| 5 |
A |
3137 |
3027 |
2.41 |
|
|
|
| 6 |
A |
3111 |
3002 |
8.08 |
|
|
|
| 7 |
A |
3076 |
2968 |
21.14 |
|
|
|
| 8 |
A |
3044 |
2938 |
13.78 |
|
|
|
| 9 |
A |
3034 |
2928 |
26.89 |
|
|
|
| 10 |
A |
3016 |
2910 |
24.95 |
|
|
|
| 11 |
A |
1742 |
1681 |
0.68 |
|
|
|
| 12 |
A |
1719 |
1658 |
9.37 |
|
|
|
| 13 |
A |
1559 |
1505 |
9.90 |
|
|
|
| 14 |
A |
1550 |
1496 |
7.84 |
|
|
|
| 15 |
A |
1530 |
1477 |
12.72 |
|
|
|
| 16 |
A |
1497 |
1444 |
1.75 |
|
|
|
| 17 |
A |
1464 |
1412 |
4.50 |
|
|
|
| 18 |
A |
1392 |
1343 |
1.29 |
|
|
|
| 19 |
A |
1375 |
1326 |
1.14 |
|
|
|
| 20 |
A |
1367 |
1319 |
2.83 |
|
|
|
| 21 |
A |
1351 |
1303 |
0.30 |
|
|
|
| 22 |
A |
1270 |
1225 |
1.01 |
|
|
|
| 23 |
A |
1156 |
1116 |
5.40 |
|
|
|
| 24 |
A |
1131 |
1091 |
3.93 |
|
|
|
| 25 |
A |
1107 |
1068 |
0.36 |
|
|
|
| 26 |
A |
1051 |
1014 |
10.24 |
|
|
|
| 27 |
A |
1044 |
1008 |
2.51 |
|
|
|
| 28 |
A |
1020 |
985 |
36.70 |
|
|
|
| 29 |
A |
995 |
961 |
23.30 |
|
|
|
| 30 |
A |
971 |
937 |
32.20 |
|
|
|
| 31 |
A |
908 |
876 |
1.30 |
|
|
|
| 32 |
A |
905 |
873 |
2.85 |
|
|
|
| 33 |
A |
794 |
766 |
0.50 |
|
|
|
| 34 |
A |
619 |
597 |
5.83 |
|
|
|
| 35 |
A |
554 |
535 |
8.99 |
|
|
|
| 36 |
A |
472 |
455 |
0.26 |
|
|
|
| 37 |
A |
324 |
313 |
0.59 |
|
|
|
| 38 |
A |
286 |
276 |
1.82 |
|
|
|
| 39 |
A |
207 |
200 |
1.56 |
|
|
|
| 40 |
A |
157 |
152 |
0.56 |
|
|
|
| 41 |
A |
127 |
123 |
0.03 |
|
|
|
| 42 |
A |
75 |
73 |
0.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 31415.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 30312.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.600 |
|
|
-0.371 |
| 2 |
H |
0.201 |
|
|
0.121 |
| 3 |
H |
0.195 |
|
|
0.115 |
| 4 |
H |
0.201 |
|
|
0.116 |
| 5 |
C |
-0.163 |
|
|
-0.053 |
| 6 |
H |
0.176 |
|
|
0.129 |
| 7 |
C |
-0.147 |
|
|
-0.264 |
| 8 |
H |
0.179 |
|
|
0.178 |
| 9 |
C |
-0.477 |
|
|
-0.126 |
| 10 |
H |
0.208 |
|
|
0.087 |
| 11 |
H |
0.207 |
|
|
0.089 |
| 12 |
C |
-0.149 |
|
|
-0.014 |
| 13 |
H |
0.180 |
|
|
0.095 |
| 14 |
C |
-0.378 |
|
|
-0.480 |
| 15 |
H |
0.179 |
|
|
0.180 |
| 16 |
H |
0.189 |
|
|
0.197 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.317 |
-0.226 |
-0.037 |
0.391 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.326 |
-0.218 |
-0.046 |
0.395 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-36.203 |
1.429 |
-0.420 |
| y |
1.429 |
-37.714 |
0.730 |
| z |
-0.420 |
0.730 |
-38.957 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.132 |
1.429 |
-0.420 |
| y |
1.429 |
-0.134 |
0.730 |
| z |
-0.420 |
0.730 |
-1.998 |
|
| Polar |
|---|
| 3z2-r2 | -3.997 |
|---|
| x2-y2 | 1.511 |
|---|
| xy | 1.429 |
|---|
| xz | -0.420 |
|---|
| yz | 0.730 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.253 |
0.467 |
-0.878 |
| y |
0.467 |
8.612 |
0.234 |
| z |
-0.878 |
0.234 |
6.238 |
<r2> (average value of r
2) Å
2
| <r2> |
232.948 |
| (<r2>)1/2 |
15.263 |