Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -134.429921 |
| Energy at 298.15K | -134.438047 |
| Nuclear repulsion energy | 81.968580 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3384 |
3265 |
3.99 |
|
|
|
| 2 |
A' |
3104 |
2995 |
44.73 |
|
|
|
| 3 |
A' |
3055 |
2948 |
29.15 |
|
|
|
| 4 |
A' |
3037 |
2931 |
20.66 |
|
|
|
| 5 |
A' |
1724 |
1663 |
16.34 |
|
|
|
| 6 |
A' |
1566 |
1511 |
3.08 |
|
|
|
| 7 |
A' |
1545 |
1491 |
0.89 |
|
|
|
| 8 |
A' |
1454 |
1403 |
4.85 |
|
|
|
| 9 |
A' |
1402 |
1353 |
7.58 |
|
|
|
| 10 |
A' |
1157 |
1116 |
11.58 |
|
|
|
| 11 |
A' |
1027 |
991 |
5.60 |
|
|
|
| 12 |
A' |
885 |
854 |
25.16 |
|
|
|
| 13 |
A' |
720 |
695 |
238.49 |
|
|
|
| 14 |
A' |
393 |
379 |
7.89 |
|
|
|
| 15 |
A" |
3479 |
3357 |
2.41 |
|
|
|
| 16 |
A" |
3113 |
3004 |
59.65 |
|
|
|
| 17 |
A" |
3082 |
2974 |
0.67 |
|
|
|
| 18 |
A" |
1565 |
1510 |
8.33 |
|
|
|
| 19 |
A" |
1411 |
1362 |
0.16 |
|
|
|
| 20 |
A" |
1302 |
1257 |
0.00 |
|
|
|
| 21 |
A" |
1028 |
992 |
1.72 |
|
|
|
| 22 |
A" |
793 |
765 |
2.14 |
|
|
|
| 23 |
A" |
309 |
298 |
52.88 |
|
|
|
| 24 |
A" |
256 |
247 |
9.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20395.3 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 19679.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
-1.314 |
-0.101 |
0.000 |
| C2 |
0.000 |
0.585 |
0.000 |
| C3 |
1.231 |
-0.354 |
0.000 |
| H4 |
2.167 |
0.218 |
0.000 |
| H5 |
1.220 |
-0.997 |
0.889 |
| H6 |
1.220 |
-0.997 |
-0.889 |
| H7 |
0.035 |
1.235 |
-0.881 |
| H8 |
0.035 |
1.235 |
0.881 |
| H9 |
-1.431 |
-0.685 |
0.835 |
| H10 |
-1.431 |
-0.685 |
-0.835 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| N1 | | 1.4828 | 2.5582 | 3.4961 | 2.8307 | 2.8307 | 2.0936 | 2.0936 | 1.0253 | 1.0253 |
C2 | 1.4828 | | 1.5482 | 2.1979 | 2.1859 | 2.1859 | 1.0957 | 1.0957 | 2.0874 | 2.0874 | C3 | 2.5582 | 1.5482 | | 1.0966 | 1.0967 | 1.0967 | 2.1755 | 2.1755 | 2.8097 | 2.8097 | H4 | 3.4961 | 2.1979 | 1.0966 | | 1.7781 | 1.7781 | 2.5214 | 2.5214 | 3.8024 | 3.8024 | H5 | 2.8307 | 2.1859 | 1.0967 | 1.7781 | | 1.7774 | 3.0849 | 2.5266 | 2.6696 | 3.1773 | H6 | 2.8307 | 2.1859 | 1.0967 | 1.7781 | 1.7774 | | 2.5266 | 3.0849 | 3.1773 | 2.6696 | H7 | 2.0936 | 1.0957 | 2.1755 | 2.5214 | 3.0849 | 2.5266 | | 1.7625 | 2.9632 | 2.4160 | H8 | 2.0936 | 1.0957 | 2.1755 | 2.5214 | 2.5266 | 3.0849 | 1.7625 | | 2.4160 | 2.9632 | H9 | 1.0253 | 2.0874 | 2.8097 | 3.8024 | 2.6696 | 3.1773 | 2.9632 | 2.4160 | | 1.6702 | H10 | 1.0253 | 2.0874 | 2.8097 | 3.8024 | 3.1773 | 2.6696 | 2.4160 | 2.9632 | 1.6702 | |
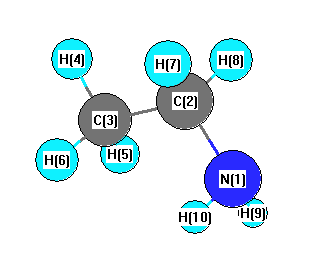 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C3 |
115.111 |
|
N1 |
C2 |
H7 |
107.628 |
| N1 |
C2 |
H8 |
107.628 |
|
C2 |
N1 |
H9 |
111.355 |
| C2 |
N1 |
H10 |
111.355 |
|
C2 |
C3 |
H4 |
111.258 |
| C2 |
C3 |
H5 |
110.298 |
|
C2 |
C3 |
H6 |
110.298 |
| C3 |
C2 |
H7 |
109.543 |
|
C3 |
C2 |
H8 |
109.543 |
| H4 |
C3 |
H5 |
108.322 |
|
H4 |
C3 |
H6 |
108.322 |
| H5 |
C3 |
H6 |
108.255 |
|
H7 |
C2 |
H8 |
107.073 |
| H9 |
N1 |
H10 |
109.066 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.646 |
|
|
-1.039 |
| 2 |
C |
-0.240 |
|
|
0.384 |
| 3 |
C |
-0.568 |
|
|
-0.391 |
| 4 |
H |
0.182 |
|
|
0.071 |
| 5 |
H |
0.180 |
|
|
0.099 |
| 6 |
H |
0.180 |
|
|
0.099 |
| 7 |
H |
0.199 |
|
|
0.004 |
| 8 |
H |
0.199 |
|
|
0.004 |
| 9 |
H |
0.257 |
|
|
0.385 |
| 10 |
H |
0.257 |
|
|
0.385 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.907 |
-1.082 |
0.000 |
1.412 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.855 |
-1.108 |
0.000 |
1.400 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.622 |
-0.082 |
-0.090 |
| y |
-0.082 |
4.052 |
-0.226 |
| z |
-0.090 |
-0.226 |
3.617 |
<r2> (average value of r
2) Å
2
| <r2> |
59.407 |
| (<r2>)1/2 |
7.708 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -134.428710 |
| Energy at 298.15K | |
| Nuclear repulsion energy | |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
