Jump to
S1C2
Energy calculated at BLYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -475.534106 |
| Energy at 298.15K | -475.540123 |
| HF Energy | -475.534106 |
| Nuclear repulsion energy | 103.926044 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3041 |
3024 |
26.78 |
|
|
|
| 2 |
A' |
3020 |
3003 |
8.00 |
|
|
|
| 3 |
A' |
2981 |
2964 |
16.33 |
|
|
|
| 4 |
A' |
2371 |
2358 |
47.02 |
|
|
|
| 5 |
A' |
1535 |
1526 |
4.08 |
|
|
|
| 6 |
A' |
1502 |
1494 |
3.25 |
|
|
|
| 7 |
A' |
1429 |
1421 |
6.76 |
|
|
|
| 8 |
A' |
1278 |
1271 |
48.45 |
|
|
|
| 9 |
A' |
1084 |
1078 |
1.82 |
|
|
|
| 10 |
A' |
958 |
953 |
8.74 |
|
|
|
| 11 |
A' |
802 |
798 |
3.62 |
|
|
|
| 12 |
A' |
581 |
577 |
4.45 |
|
|
|
| 13 |
A' |
292 |
291 |
4.55 |
|
|
|
| 14 |
A" |
3087 |
3070 |
25.48 |
|
|
|
| 15 |
A" |
3048 |
3032 |
5.60 |
|
|
|
| 16 |
A" |
1525 |
1516 |
8.14 |
|
|
|
| 17 |
A" |
1273 |
1266 |
0.96 |
|
|
|
| 18 |
A" |
1027 |
1021 |
0.05 |
|
|
|
| 19 |
A" |
787 |
783 |
6.55 |
|
|
|
| 20 |
A" |
242 |
241 |
0.39 |
|
|
|
| 21 |
A" |
117 |
116 |
24.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 15988.9 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 15901.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.537 |
0.758 |
0.000 |
| C2 |
0.000 |
0.892 |
0.000 |
| S3 |
-0.761 |
-0.901 |
0.000 |
| H4 |
1.996 |
1.762 |
0.000 |
| H5 |
1.881 |
0.217 |
0.895 |
| H6 |
1.881 |
0.217 |
-0.895 |
| H7 |
-0.361 |
1.404 |
0.902 |
| H8 |
-0.361 |
1.404 |
-0.902 |
| H9 |
-2.090 |
-0.494 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5432 | 2.8344 | 1.1037 | 1.1014 | 1.1014 | 2.1988 | 2.1988 | 3.8374 |
C2 | 1.5432 | | 1.9471 | 2.1779 | 2.1900 | 2.1900 | 1.0981 | 1.0981 | 2.5075 | S3 | 2.8344 | 1.9471 | | 3.8330 | 3.0050 | 3.0050 | 2.5065 | 2.5065 | 1.3898 | H4 | 1.1037 | 2.1779 | 3.8330 | | 1.7896 | 1.7896 | 2.5495 | 2.5495 | 4.6677 | H5 | 1.1014 | 2.1900 | 3.0050 | 1.7896 | | 1.7905 | 2.5372 | 3.1092 | 4.1323 | H6 | 1.1014 | 2.1900 | 3.0050 | 1.7896 | 1.7905 | | 3.1092 | 2.5372 | 4.1323 | H7 | 2.1988 | 1.0981 | 2.5065 | 2.5495 | 2.5372 | 3.1092 | | 1.8037 | 2.7208 | H8 | 2.1988 | 1.0981 | 2.5065 | 2.5495 | 3.1092 | 2.5372 | 1.8037 | | 2.7208 | H9 | 3.8374 | 2.5075 | 1.3898 | 4.6677 | 4.1323 | 4.1323 | 2.7208 | 2.7208 | |
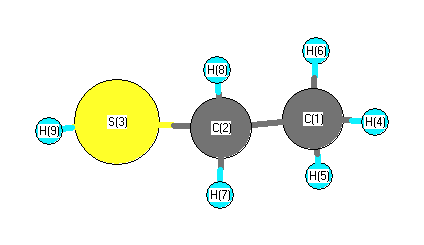 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.044 |
|
C1 |
C2 |
H7 |
111.590 |
| C1 |
C2 |
H8 |
111.590 |
|
C2 |
C1 |
H4 |
109.613 |
| C2 |
C1 |
H5 |
110.699 |
|
C2 |
C1 |
H6 |
110.699 |
| C2 |
S3 |
H9 |
95.993 |
|
S3 |
C2 |
H7 |
107.496 |
| S3 |
C2 |
H8 |
107.496 |
|
H4 |
C1 |
H5 |
108.508 |
| H4 |
C1 |
H6 |
108.508 |
|
H5 |
C1 |
H6 |
108.754 |
| H7 |
C2 |
H8 |
110.420 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.524 |
|
|
|
| 2 |
C |
-0.550 |
|
|
|
| 3 |
S |
-0.012 |
|
|
|
| 4 |
H |
0.190 |
|
|
|
| 5 |
H |
0.200 |
|
|
|
| 6 |
H |
0.200 |
|
|
|
| 7 |
H |
0.216 |
|
|
|
| 8 |
H |
0.216 |
|
|
|
| 9 |
H |
0.065 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.049 |
2.023 |
0.000 |
2.024 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.619 |
-0.138 |
0.000 |
| y |
-0.138 |
-28.829 |
0.000 |
| z |
0.000 |
0.000 |
-29.063 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.327 |
-0.138 |
0.000 |
| y |
-0.138 |
-1.988 |
0.000 |
| z |
0.000 |
0.000 |
-2.339 |
|
| Polar |
|---|
| 3z2-r2 | -4.679 |
|---|
| x2-y2 | 4.210 |
|---|
| xy | -0.138 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.351 |
0.971 |
0.000 |
| y |
0.971 |
5.934 |
0.000 |
| z |
0.000 |
0.000 |
3.786 |
<r2> (average value of r
2) Å
2
| <r2> |
88.125 |
| (<r2>)1/2 |
9.387 |
Jump to
S1C1
Energy calculated at BLYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -475.535280 |
| Energy at 298.15K | -475.541462 |
| HF Energy | -475.535280 |
| Nuclear repulsion energy | 103.813823 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at BLYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3086 |
3069 |
19.39 |
|
|
|
| 2 |
A |
3052 |
3036 |
8.60 |
|
|
|
| 3 |
A |
3032 |
3016 |
31.14 |
|
|
|
| 4 |
A |
3019 |
3003 |
3.05 |
|
|
|
| 5 |
A |
2969 |
2953 |
22.22 |
|
|
|
| 6 |
A |
2367 |
2354 |
43.38 |
|
|
|
| 7 |
A |
1531 |
1523 |
3.35 |
|
|
|
| 8 |
A |
1523 |
1514 |
10.38 |
|
|
|
| 9 |
A |
1500 |
1492 |
3.02 |
|
|
|
| 10 |
A |
1429 |
1421 |
4.84 |
|
|
|
| 11 |
A |
1286 |
1279 |
11.58 |
|
|
|
| 12 |
A |
1273 |
1266 |
16.99 |
|
|
|
| 13 |
A |
1097 |
1091 |
10.80 |
|
|
|
| 14 |
A |
1047 |
1041 |
0.63 |
|
|
|
| 15 |
A |
945 |
940 |
10.82 |
|
|
|
| 16 |
A |
855 |
850 |
12.49 |
|
|
|
| 17 |
A |
724 |
720 |
2.29 |
|
|
|
| 18 |
A |
573 |
570 |
9.01 |
|
|
|
| 19 |
A |
312 |
311 |
1.48 |
|
|
|
| 20 |
A |
257 |
255 |
1.81 |
|
|
|
| 21 |
A |
199 |
198 |
25.34 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16037.0 cm
-1
Scaled (by 0.9945) Zero Point Vibrational Energy (zpe) 15948.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at BLYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.675 |
-0.378 |
-0.056 |
| C2 |
0.560 |
0.677 |
0.092 |
| S3 |
-1.211 |
-0.098 |
-0.082 |
| H4 |
2.665 |
0.111 |
-0.002 |
| H5 |
1.618 |
-1.127 |
0.751 |
| H6 |
1.588 |
-0.898 |
-1.022 |
| H7 |
0.601 |
1.199 |
1.058 |
| H8 |
0.588 |
1.417 |
-0.720 |
| H9 |
-1.099 |
-0.925 |
1.029 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5419 | 2.8997 | 1.1052 | 1.1025 | 1.1005 | 2.2097 | 2.2004 | 3.0286 |
C2 | 1.5419 | | 1.9414 | 2.1810 | 2.1925 | 2.1859 | 1.0985 | 1.0988 | 2.4894 | S3 | 2.8997 | 1.9414 | | 3.8820 | 3.1234 | 3.0589 | 2.5030 | 2.4371 | 1.3895 | H4 | 1.1052 | 2.1810 | 3.8820 | | 1.7874 | 1.7939 | 2.5626 | 2.5554 | 4.0375 | H5 | 1.1025 | 2.1925 | 3.1234 | 1.7874 | | 1.7880 | 2.5576 | 3.1134 | 2.7388 | H6 | 1.1005 | 2.1859 | 3.0589 | 1.7939 | 1.7880 | | 3.1143 | 2.5393 | 3.3806 | H7 | 2.2097 | 1.0985 | 2.5030 | 2.5626 | 2.5576 | 3.1143 | | 1.7907 | 2.7206 | H8 | 2.2004 | 1.0988 | 2.4371 | 2.5554 | 3.1134 | 2.5393 | 1.7907 | | 3.3747 | H9 | 3.0286 | 2.4894 | 1.3895 | 4.0375 | 2.7388 | 3.3806 | 2.7206 | 3.3747 | |
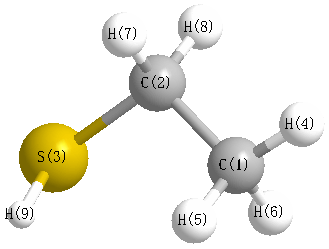 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
112.195 |
|
C1 |
C2 |
H7 |
112.542 |
| C1 |
C2 |
H8 |
111.778 |
|
C2 |
C1 |
H4 |
109.865 |
| C2 |
C1 |
H5 |
110.920 |
|
C2 |
C1 |
H6 |
110.518 |
| C2 |
S3 |
H9 |
95.290 |
|
S3 |
C2 |
H7 |
107.598 |
| S3 |
C2 |
H8 |
103.051 |
|
H4 |
C1 |
H5 |
108.125 |
| H4 |
C1 |
H6 |
108.842 |
|
H5 |
C1 |
H6 |
108.506 |
| H7 |
C2 |
H8 |
109.174 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at BLYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.516 |
|
|
|
| 2 |
C |
-0.559 |
|
|
|
| 3 |
S |
-0.007 |
|
|
|
| 4 |
H |
0.187 |
|
|
|
| 5 |
H |
0.184 |
|
|
|
| 6 |
H |
0.202 |
|
|
|
| 7 |
H |
0.217 |
|
|
|
| 8 |
H |
0.224 |
|
|
|
| 9 |
H |
0.067 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.881 |
0.034 |
0.939 |
2.102 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.512 |
1.307 |
-1.099 |
| y |
1.307 |
-27.255 |
-1.564 |
| z |
-1.099 |
-1.564 |
-27.158 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.305 |
1.307 |
-1.099 |
| y |
1.307 |
1.080 |
-1.564 |
| z |
-1.099 |
-1.564 |
1.225 |
|
| Polar |
|---|
| 3z2-r2 | 2.450 |
|---|
| x2-y2 | -2.257 |
|---|
| xy | 1.307 |
|---|
| xz | -1.099 |
|---|
| yz | -1.564 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.778 |
0.647 |
0.109 |
| y |
0.647 |
4.695 |
-0.604 |
| z |
0.109 |
-0.604 |
4.658 |
<r2> (average value of r
2) Å
2
| <r2> |
88.443 |
| (<r2>)1/2 |
9.404 |
