Jump to
S1C2
Energy calculated at wB97X-D/3-21G
| | hartrees |
|---|
| Energy at 0K | -475.575405 |
| Energy at 298.15K | |
| HF Energy | -475.575405 |
| Nuclear repulsion energy | 105.548446 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3169 |
3169 |
15.50 |
|
|
|
| 2 |
A' |
3139 |
3139 |
9.52 |
|
|
|
| 3 |
A' |
3093 |
3093 |
12.21 |
|
|
|
| 4 |
A' |
2503 |
2503 |
33.35 |
|
|
|
| 5 |
A' |
1573 |
1573 |
6.97 |
|
|
|
| 6 |
A' |
1545 |
1545 |
6.93 |
|
|
|
| 7 |
A' |
1480 |
1480 |
15.04 |
|
|
|
| 8 |
A' |
1339 |
1339 |
36.72 |
|
|
|
| 9 |
A' |
1134 |
1134 |
3.92 |
|
|
|
| 10 |
A' |
1013 |
1013 |
8.73 |
|
|
|
| 11 |
A' |
846 |
846 |
5.86 |
|
|
|
| 12 |
A' |
648 |
648 |
3.30 |
|
|
|
| 13 |
A' |
314 |
314 |
4.88 |
|
|
|
| 14 |
A" |
3209 |
3209 |
12.89 |
|
|
|
| 15 |
A" |
3168 |
3168 |
5.17 |
|
|
|
| 16 |
A" |
1564 |
1564 |
12.45 |
|
|
|
| 17 |
A" |
1318 |
1318 |
1.08 |
|
|
|
| 18 |
A" |
1077 |
1077 |
0.26 |
|
|
|
| 19 |
A" |
821 |
821 |
8.02 |
|
|
|
| 20 |
A" |
243 |
243 |
0.13 |
|
|
|
| 21 |
A" |
123i |
123i |
33.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16536.5 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16536.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.523 |
0.728 |
0.000 |
| C2 |
0.000 |
0.870 |
0.000 |
| S3 |
-0.755 |
-0.873 |
0.000 |
| H4 |
1.987 |
1.719 |
0.000 |
| H5 |
1.860 |
0.186 |
0.889 |
| H6 |
1.860 |
0.186 |
-0.889 |
| H7 |
-0.347 |
1.389 |
0.894 |
| H8 |
-0.347 |
1.389 |
-0.894 |
| H9 |
-2.067 |
-0.483 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5301 | 2.7850 | 1.0948 | 1.0937 | 1.0937 | 2.1761 | 2.1761 | 3.7894 |
C2 | 1.5301 | | 1.9001 | 2.1610 | 2.1717 | 2.1717 | 1.0905 | 1.0905 | 2.4708 | S3 | 2.7850 | 1.9001 | | 3.7741 | 2.9582 | 2.9582 | 2.4667 | 2.4667 | 1.3690 | H4 | 1.0948 | 2.1610 | 3.7741 | | 1.7768 | 1.7768 | 2.5213 | 2.5213 | 4.6141 | H5 | 1.0937 | 2.1717 | 2.9582 | 1.7768 | | 1.7774 | 2.5133 | 3.0815 | 4.0817 | H6 | 1.0937 | 2.1717 | 2.9582 | 1.7768 | 1.7774 | | 3.0815 | 2.5133 | 4.0817 | H7 | 2.1761 | 1.0905 | 2.4667 | 2.5213 | 2.5133 | 3.0815 | | 1.7883 | 2.6950 | H8 | 2.1761 | 1.0905 | 2.4667 | 2.5213 | 3.0815 | 2.5133 | 1.7883 | | 2.6950 | H9 | 3.7894 | 2.4708 | 1.3690 | 4.6141 | 4.0817 | 4.0817 | 2.6950 | 2.6950 | |
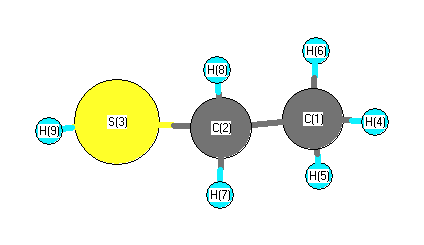 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
108.079 |
|
C1 |
C2 |
H7 |
111.171 |
| C1 |
C2 |
H8 |
111.171 |
|
C2 |
C1 |
H4 |
109.719 |
| C2 |
C1 |
H5 |
110.628 |
|
C2 |
C1 |
H6 |
110.628 |
| C2 |
S3 |
H9 |
96.848 |
|
S3 |
C2 |
H7 |
108.064 |
| S3 |
C2 |
H8 |
108.064 |
|
H4 |
C1 |
H5 |
108.558 |
| H4 |
C1 |
H6 |
108.558 |
|
H5 |
C1 |
H6 |
108.694 |
| H7 |
C2 |
H8 |
110.164 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.617 |
|
|
|
| 2 |
C |
-0.662 |
|
|
|
| 3 |
S |
0.004 |
|
|
|
| 4 |
H |
0.225 |
|
|
|
| 5 |
H |
0.233 |
|
|
|
| 6 |
H |
0.233 |
|
|
|
| 7 |
H |
0.252 |
|
|
|
| 8 |
H |
0.252 |
|
|
|
| 9 |
H |
0.081 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.139 |
2.123 |
0.000 |
2.128 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.321 |
-0.138 |
0.000 |
| y |
-0.138 |
-28.926 |
0.000 |
| z |
0.000 |
0.000 |
-28.917 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.601 |
-0.138 |
0.000 |
| y |
-0.138 |
-2.307 |
0.000 |
| z |
0.000 |
0.000 |
-2.293 |
|
| Polar |
|---|
| 3z2-r2 | -4.587 |
|---|
| x2-y2 | 4.605 |
|---|
| xy | -0.138 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.124 |
0.895 |
0.000 |
| y |
0.895 |
5.710 |
0.000 |
| z |
0.000 |
0.000 |
3.737 |
<r2> (average value of r
2) Å
2
| <r2> |
85.762 |
| (<r2>)1/2 |
9.261 |
Jump to
S1C1
Energy calculated at wB97X-D/3-21G
| | hartrees |
|---|
| Energy at 0K | -475.576546 |
| Energy at 298.15K | |
| HF Energy | -475.576546 |
| Nuclear repulsion energy | 105.432127 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at wB97X-D/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3201 |
3201 |
8.61 |
30.35 |
0.75 |
0.86 |
| 2 |
A |
3166 |
3166 |
7.18 |
73.50 |
0.74 |
0.85 |
| 3 |
A |
3148 |
3148 |
20.66 |
73.09 |
0.39 |
0.56 |
| 4 |
A |
3134 |
3134 |
3.55 |
99.46 |
0.32 |
0.48 |
| 5 |
A |
3078 |
3078 |
14.96 |
106.91 |
0.04 |
0.08 |
| 6 |
A |
2496 |
2496 |
31.06 |
117.96 |
0.37 |
0.54 |
| 7 |
A |
1568 |
1568 |
5.69 |
18.21 |
0.74 |
0.85 |
| 8 |
A |
1564 |
1564 |
15.58 |
21.78 |
0.75 |
0.86 |
| 9 |
A |
1541 |
1541 |
8.11 |
17.67 |
0.75 |
0.85 |
| 10 |
A |
1478 |
1478 |
12.60 |
5.45 |
0.67 |
0.80 |
| 11 |
A |
1343 |
1343 |
14.66 |
4.90 |
0.62 |
0.76 |
| 12 |
A |
1323 |
1323 |
4.73 |
10.37 |
0.75 |
0.86 |
| 13 |
A |
1144 |
1144 |
11.59 |
15.12 |
0.75 |
0.86 |
| 14 |
A |
1097 |
1097 |
1.89 |
7.09 |
0.58 |
0.74 |
| 15 |
A |
1004 |
1004 |
10.32 |
5.57 |
0.74 |
0.85 |
| 16 |
A |
896 |
896 |
17.43 |
10.42 |
0.75 |
0.86 |
| 17 |
A |
758 |
758 |
3.18 |
5.52 |
0.63 |
0.77 |
| 18 |
A |
640 |
640 |
7.88 |
22.70 |
0.31 |
0.47 |
| 19 |
A |
337 |
337 |
1.23 |
2.56 |
0.50 |
0.66 |
| 20 |
A |
265 |
265 |
7.25 |
3.33 |
0.74 |
0.85 |
| 21 |
A |
231 |
231 |
27.75 |
15.38 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16706.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 16706.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at wB97X-D/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.648 |
-0.372 |
-0.055 |
| C2 |
0.539 |
0.670 |
0.092 |
| S3 |
-1.187 |
-0.097 |
-0.080 |
| H4 |
2.628 |
0.114 |
0.002 |
| H5 |
1.589 |
-1.118 |
0.743 |
| H6 |
1.563 |
-0.885 |
-1.017 |
| H7 |
0.586 |
1.187 |
1.051 |
| H8 |
0.578 |
1.409 |
-0.710 |
| H9 |
-1.077 |
-0.936 |
0.997 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5284 | 2.8475 | 1.0958 | 1.0948 | 1.0930 | 2.1861 | 2.1775 | 2.9746 |
C2 | 1.5284 | | 1.8959 | 2.1641 | 2.1737 | 2.1672 | 1.0907 | 1.0911 | 2.4510 | S3 | 2.8475 | 1.8959 | | 3.8215 | 3.0699 | 3.0099 | 2.4640 | 2.4042 | 1.3698 | H4 | 1.0958 | 2.1641 | 3.8215 | | 1.7742 | 1.7806 | 2.5338 | 2.5269 | 3.9773 | H5 | 1.0948 | 2.1737 | 3.0699 | 1.7742 | | 1.7751 | 2.5326 | 3.0853 | 2.6840 | H6 | 1.0930 | 2.1672 | 3.0099 | 1.7806 | 1.7751 | | 3.0859 | 2.5153 | 3.3208 | H7 | 2.1861 | 1.0907 | 2.4640 | 2.5338 | 2.5326 | 3.0859 | | 1.7749 | 2.6971 | H8 | 2.1775 | 1.0911 | 2.4042 | 2.5269 | 3.0853 | 2.5153 | 1.7749 | | 3.3394 | H9 | 2.9746 | 2.4510 | 1.3698 | 3.9773 | 2.6840 | 3.3208 | 2.6971 | 3.3394 | |
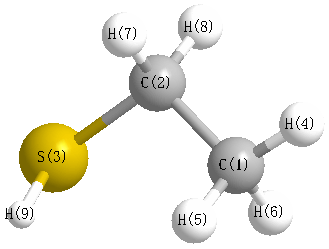 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
112.073 |
|
C1 |
C2 |
H7 |
112.082 |
| C1 |
C2 |
H8 |
111.363 |
|
C2 |
C1 |
H4 |
110.009 |
| C2 |
C1 |
H5 |
110.837 |
|
C2 |
C1 |
H6 |
110.426 |
| C2 |
S3 |
H9 |
95.935 |
|
S3 |
C2 |
H7 |
108.138 |
| S3 |
C2 |
H8 |
103.921 |
|
H4 |
C1 |
H5 |
108.167 |
| H4 |
C1 |
H6 |
108.879 |
|
H5 |
C1 |
H6 |
108.459 |
| H7 |
C2 |
H8 |
108.885 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at wB97X-D/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.608 |
|
|
|
| 2 |
C |
-0.671 |
|
|
|
| 3 |
S |
0.010 |
|
|
|
| 4 |
H |
0.222 |
|
|
|
| 5 |
H |
0.214 |
|
|
|
| 6 |
H |
0.235 |
|
|
|
| 7 |
H |
0.254 |
|
|
|
| 8 |
H |
0.263 |
|
|
|
| 9 |
H |
0.081 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.978 |
-0.007 |
1.023 |
2.227 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.652 |
1.417 |
-1.227 |
| y |
1.417 |
-27.007 |
-1.621 |
| z |
-1.227 |
-1.621 |
-27.076 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.610 |
1.417 |
-1.227 |
| y |
1.417 |
1.357 |
-1.621 |
| z |
-1.227 |
-1.621 |
1.254 |
|
| Polar |
|---|
| 3z2-r2 | 2.507 |
|---|
| x2-y2 | -2.645 |
|---|
| xy | 1.417 |
|---|
| xz | -1.227 |
|---|
| yz | -1.621 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.347 |
0.656 |
0.105 |
| y |
0.656 |
4.632 |
-0.605 |
| z |
0.105 |
-0.605 |
4.551 |
<r2> (average value of r
2) Å
2
| <r2> |
86.051 |
| (<r2>)1/2 |
9.276 |
