Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -475.620392 |
| Energy at 298.15K | -475.626516 |
| HF Energy | -475.620392 |
| Nuclear repulsion energy | 104.965785 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3123 |
3013 |
23.19 |
|
|
|
| 2 |
A' |
3102 |
2993 |
7.63 |
|
|
|
| 3 |
A' |
3059 |
2951 |
14.73 |
|
|
|
| 4 |
A' |
2459 |
2373 |
40.95 |
|
|
|
| 5 |
A' |
1570 |
1515 |
4.89 |
|
|
|
| 6 |
A' |
1539 |
1485 |
4.54 |
|
|
|
| 7 |
A' |
1469 |
1417 |
8.99 |
|
|
|
| 8 |
A' |
1322 |
1276 |
44.71 |
|
|
|
| 9 |
A' |
1119 |
1080 |
2.60 |
|
|
|
| 10 |
A' |
992 |
957 |
8.23 |
|
|
|
| 11 |
A' |
833 |
803 |
4.57 |
|
|
|
| 12 |
A' |
614 |
592 |
4.15 |
|
|
|
| 13 |
A' |
305 |
294 |
4.59 |
|
|
|
| 14 |
A" |
3169 |
3058 |
19.31 |
|
|
|
| 15 |
A" |
3129 |
3019 |
5.78 |
|
|
|
| 16 |
A" |
1561 |
1506 |
9.52 |
|
|
|
| 17 |
A" |
1310 |
1264 |
0.95 |
|
|
|
| 18 |
A" |
1062 |
1025 |
0.09 |
|
|
|
| 19 |
A" |
810 |
781 |
6.76 |
|
|
|
| 20 |
A" |
249 |
240 |
0.46 |
|
|
|
| 21 |
A" |
128 |
123 |
27.88 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16460.8 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 15883.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.527 |
0.742 |
0.000 |
| C2 |
0.000 |
0.880 |
0.000 |
| S3 |
-0.756 |
-0.885 |
0.000 |
| H4 |
1.988 |
1.736 |
0.000 |
| H5 |
1.866 |
0.201 |
0.889 |
| H6 |
1.866 |
0.201 |
-0.889 |
| H7 |
-0.354 |
1.394 |
0.895 |
| H8 |
-0.354 |
1.394 |
-0.895 |
| H9 |
-2.075 |
-0.491 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5328 | 2.8030 | 1.0960 | 1.0943 | 1.0943 | 2.1824 | 2.1824 | 3.8067 |
C2 | 1.5328 | | 1.9205 | 2.1644 | 2.1755 | 2.1755 | 1.0909 | 1.0909 | 2.4872 | S3 | 2.8030 | 1.9205 | | 3.7947 | 2.9740 | 2.9740 | 2.4813 | 2.4813 | 1.3765 | H4 | 1.0960 | 2.1644 | 3.7947 | | 1.7775 | 1.7775 | 2.5305 | 2.5305 | 4.6334 | H5 | 1.0943 | 2.1755 | 2.9740 | 1.7775 | | 1.7780 | 2.5200 | 3.0874 | 4.0987 | H6 | 1.0943 | 2.1755 | 2.9740 | 1.7775 | 1.7780 | | 3.0874 | 2.5200 | 4.0987 | H7 | 2.1824 | 1.0909 | 2.4813 | 2.5305 | 2.5200 | 3.0874 | | 1.7897 | 2.7049 | H8 | 2.1824 | 1.0909 | 2.4813 | 2.5305 | 3.0874 | 2.5200 | 1.7897 | | 2.7049 | H9 | 3.8067 | 2.4872 | 1.3765 | 4.6334 | 4.0987 | 4.0987 | 2.7049 | 2.7049 | |
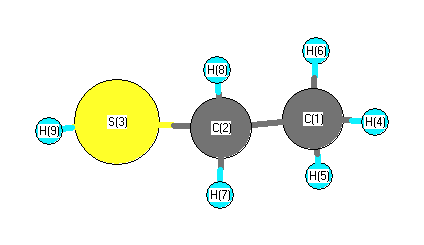 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
107.995 |
|
C1 |
C2 |
H7 |
111.454 |
| C1 |
C2 |
H8 |
111.454 |
|
C2 |
C1 |
H4 |
109.724 |
| C2 |
C1 |
H5 |
110.698 |
|
C2 |
C1 |
H6 |
110.698 |
| C2 |
S3 |
H9 |
96.551 |
|
S3 |
C2 |
H7 |
107.767 |
| S3 |
C2 |
H8 |
107.767 |
|
H4 |
C1 |
H5 |
108.499 |
| H4 |
C1 |
H6 |
108.499 |
|
H5 |
C1 |
H6 |
108.662 |
| H7 |
C2 |
H8 |
110.234 |
|
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.560 |
|
|
|
| 2 |
C |
-0.599 |
|
|
|
| 3 |
S |
0.001 |
|
|
|
| 4 |
H |
0.204 |
|
|
|
| 5 |
H |
0.212 |
|
|
|
| 6 |
H |
0.212 |
|
|
|
| 7 |
H |
0.231 |
|
|
|
| 8 |
H |
0.231 |
|
|
|
| 9 |
H |
0.069 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.088 |
2.054 |
0.000 |
2.056 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.066 |
2.059 |
0.000 |
2.060 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-24.572 |
-0.139 |
0.000 |
| y |
-0.139 |
-28.915 |
0.000 |
| z |
0.000 |
0.000 |
-29.027 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.399 |
-0.139 |
0.000 |
| y |
-0.139 |
-2.115 |
0.000 |
| z |
0.000 |
0.000 |
-2.284 |
|
| Polar |
|---|
| 3z2-r2 | -4.568 |
|---|
| x2-y2 | 4.343 |
|---|
| xy | -0.139 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.200 |
0.933 |
0.000 |
| y |
0.933 |
5.791 |
0.000 |
| z |
0.000 |
0.000 |
3.735 |
<r2> (average value of r
2) Å
2
| <r2> |
86.667 |
| (<r2>)1/2 |
9.310 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -475.621491 |
| Energy at 298.15K | -475.627777 |
| HF Energy | -475.621491 |
| Nuclear repulsion energy | 104.830748 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3168 |
3057 |
14.13 |
|
|
|
| 2 |
A |
3132 |
3022 |
8.45 |
|
|
|
| 3 |
A |
3114 |
3005 |
27.33 |
|
|
|
| 4 |
A |
3101 |
2992 |
2.92 |
|
|
|
| 5 |
A |
3049 |
2942 |
19.02 |
|
|
|
| 6 |
A |
2461 |
2375 |
37.36 |
|
|
|
| 7 |
A |
1566 |
1511 |
4.05 |
|
|
|
| 8 |
A |
1559 |
1505 |
12.03 |
|
|
|
| 9 |
A |
1536 |
1482 |
4.71 |
|
|
|
| 10 |
A |
1468 |
1417 |
6.85 |
|
|
|
| 11 |
A |
1327 |
1280 |
14.86 |
|
|
|
| 12 |
A |
1313 |
1267 |
10.78 |
|
|
|
| 13 |
A |
1132 |
1093 |
11.56 |
|
|
|
| 14 |
A |
1082 |
1044 |
0.96 |
|
|
|
| 15 |
A |
982 |
948 |
10.11 |
|
|
|
| 16 |
A |
882 |
851 |
14.35 |
|
|
|
| 17 |
A |
746 |
720 |
2.53 |
|
|
|
| 18 |
A |
607 |
585 |
8.77 |
|
|
|
| 19 |
A |
325 |
314 |
1.36 |
|
|
|
| 20 |
A |
263 |
254 |
2.41 |
|
|
|
| 21 |
A |
211 |
203 |
27.69 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16511.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 15931.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.659 |
-0.374 |
-0.055 |
| C2 |
0.550 |
0.671 |
0.092 |
| S3 |
-1.197 |
-0.098 |
-0.081 |
| H4 |
2.641 |
0.111 |
0.000 |
| H5 |
1.602 |
-1.120 |
0.745 |
| H6 |
1.574 |
-0.890 |
-1.016 |
| H7 |
0.592 |
1.190 |
1.051 |
| H8 |
0.583 |
1.408 |
-0.713 |
| H9 |
-1.094 |
-0.924 |
1.014 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
H4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5310 | 2.8689 | 1.0973 | 1.0953 | 1.0934 | 2.1922 | 2.1830 | 3.0044 |
C2 | 1.5310 | | 1.9158 | 2.1674 | 2.1773 | 2.1708 | 1.0911 | 1.0915 | 2.4696 | S3 | 2.8689 | 1.9158 | | 3.8446 | 3.0917 | 3.0293 | 2.4780 | 2.4151 | 1.3762 | H4 | 1.0973 | 2.1674 | 3.8446 | | 1.7750 | 1.7815 | 2.5426 | 2.5353 | 4.0071 | H5 | 1.0953 | 2.1773 | 3.0917 | 1.7750 | | 1.7756 | 2.5391 | 3.0908 | 2.7166 | H6 | 1.0934 | 2.1708 | 3.0293 | 1.7815 | 1.7756 | | 3.0917 | 2.5210 | 3.3527 | H7 | 2.1922 | 1.0911 | 2.4780 | 2.5426 | 2.5391 | 3.0917 | | 1.7767 | 2.7052 | H8 | 2.1830 | 1.0915 | 2.4151 | 2.5353 | 3.0908 | 2.5210 | 1.7767 | | 3.3520 | H9 | 3.0044 | 2.4696 | 1.3762 | 4.0071 | 2.7166 | 3.3527 | 2.7052 | 3.3520 | |
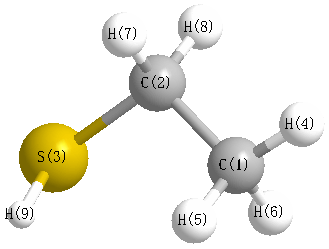 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S3 |
112.195 |
|
C1 |
C2 |
H7 |
112.369 |
| C1 |
C2 |
H8 |
111.598 |
|
C2 |
C1 |
H4 |
110.003 |
| C2 |
C1 |
H5 |
110.911 |
|
C2 |
C1 |
H6 |
110.507 |
| C2 |
S3 |
H9 |
95.822 |
|
S3 |
C2 |
H7 |
107.823 |
| S3 |
C2 |
H8 |
103.425 |
|
H4 |
C1 |
H5 |
108.097 |
| H4 |
C1 |
H6 |
108.817 |
|
H5 |
C1 |
H6 |
108.436 |
| H7 |
C2 |
H8 |
108.979 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.551 |
|
|
|
| 2 |
C |
-0.608 |
|
|
|
| 3 |
S |
0.006 |
|
|
|
| 4 |
H |
0.200 |
|
|
|
| 5 |
H |
0.195 |
|
|
|
| 6 |
H |
0.215 |
|
|
|
| 7 |
H |
0.232 |
|
|
|
| 8 |
H |
0.240 |
|
|
|
| 9 |
H |
0.071 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.909 |
0.023 |
0.974 |
2.143 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-29.613 |
1.349 |
-1.164 |
| y |
1.349 |
-27.205 |
-1.574 |
| z |
-1.164 |
-1.574 |
-27.160 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.431 |
1.349 |
-1.164 |
| y |
1.349 |
1.182 |
-1.574 |
| z |
-1.164 |
-1.574 |
1.249 |
|
| Polar |
|---|
| 3z2-r2 | 2.499 |
|---|
| x2-y2 | -2.409 |
|---|
| xy | 1.349 |
|---|
| xz | -1.164 |
|---|
| yz | -1.574 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.541 |
0.661 |
0.099 |
| y |
0.661 |
4.632 |
-0.601 |
| z |
0.099 |
-0.601 |
4.578 |
<r2> (average value of r
2) Å
2
| <r2> |
87.032 |
| (<r2>)1/2 |
9.329 |
