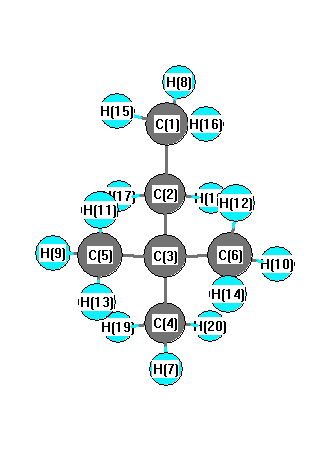
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3118 |
3009 |
52.78 |
|
|
|
| 2 |
A' |
3109 |
3000 |
74.14 |
|
|
|
| 3 |
A' |
3107 |
2998 |
37.13 |
|
|
|
| 4 |
A' |
3099 |
2990 |
0.32 |
|
|
|
| 5 |
A' |
3056 |
2949 |
15.37 |
|
|
|
| 6 |
A' |
3046 |
2939 |
10.46 |
|
|
|
| 7 |
A' |
3035 |
2929 |
33.06 |
|
|
|
| 8 |
A' |
3029 |
2922 |
18.18 |
|
|
|
| 9 |
A' |
1583 |
1528 |
14.52 |
|
|
|
| 10 |
A' |
1577 |
1522 |
8.63 |
|
|
|
| 11 |
A' |
1571 |
1516 |
6.84 |
|
|
|
| 12 |
A' |
1555 |
1500 |
0.28 |
|
|
|
| 13 |
A' |
1542 |
1488 |
0.05 |
|
|
|
| 14 |
A' |
1477 |
1425 |
2.99 |
|
|
|
| 15 |
A' |
1459 |
1408 |
14.54 |
|
|
|
| 16 |
A' |
1446 |
1395 |
7.01 |
|
|
|
| 17 |
A' |
1400 |
1351 |
0.26 |
|
|
|
| 18 |
A' |
1296 |
1250 |
6.46 |
|
|
|
| 19 |
A' |
1258 |
1214 |
8.90 |
|
|
|
| 20 |
A' |
1122 |
1083 |
4.77 |
|
|
|
| 21 |
A' |
1048 |
1011 |
4.89 |
|
|
|
| 22 |
A' |
1018 |
982 |
1.81 |
|
|
|
| 23 |
A' |
942 |
909 |
1.49 |
|
|
|
| 24 |
A' |
876 |
845 |
0.20 |
|
|
|
| 25 |
A' |
702 |
677 |
0.44 |
|
|
|
| 26 |
A' |
486 |
469 |
0.87 |
|
|
|
| 27 |
A' |
407 |
393 |
0.01 |
|
|
|
| 28 |
A' |
361 |
348 |
0.06 |
|
|
|
| 29 |
A' |
280 |
271 |
0.06 |
|
|
|
| 30 |
A' |
262 |
253 |
0.05 |
|
|
|
| 31 |
A" |
3127 |
3018 |
34.15 |
|
|
|
| 32 |
A" |
3111 |
3002 |
26.91 |
|
|
|
| 33 |
A" |
3102 |
2993 |
27.83 |
|
|
|
| 34 |
A" |
3098 |
2990 |
0.14 |
|
|
|
| 35 |
A" |
3062 |
2954 |
11.64 |
|
|
|
| 36 |
A" |
3039 |
2932 |
29.52 |
|
|
|
| 37 |
A" |
1580 |
1525 |
0.23 |
|
|
|
| 38 |
A" |
1569 |
1514 |
13.17 |
|
|
|
| 39 |
A" |
1553 |
1498 |
0.19 |
|
|
|
| 40 |
A" |
1547 |
1492 |
0.17 |
|
|
|
| 41 |
A" |
1447 |
1396 |
12.30 |
|
|
|
| 42 |
A" |
1374 |
1326 |
1.90 |
|
|
|
| 43 |
A" |
1270 |
1225 |
2.68 |
|
|
|
| 44 |
A" |
1139 |
1099 |
0.95 |
|
|
|
| 45 |
A" |
1039 |
1002 |
1.31 |
|
|
|
| 46 |
A" |
1000 |
965 |
0.01 |
|
|
|
| 47 |
A" |
946 |
913 |
0.09 |
|
|
|
| 48 |
A" |
811 |
782 |
4.05 |
|
|
|
| 49 |
A" |
418 |
404 |
0.00 |
|
|
|
| 50 |
A" |
343 |
331 |
0.02 |
|
|
|
| 51 |
A" |
293 |
283 |
0.00 |
|
|
|
| 52 |
A" |
261 |
252 |
0.00 |
|
|
|
| 53 |
A" |
222 |
214 |
0.03 |
|
|
|
| 54 |
A" |
99 |
95 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 41858.1 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 40388.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.565 |
|
|
-0.262 |
| 2 |
C |
-0.323 |
|
|
-0.055 |
| 3 |
C |
-0.150 |
|
|
0.522 |
| 4 |
C |
-0.498 |
|
|
-0.576 |
| 5 |
C |
-0.507 |
|
|
-0.494 |
| 6 |
C |
-0.507 |
|
|
-0.494 |
| 7 |
H |
0.179 |
|
|
0.149 |
| 8 |
H |
0.186 |
|
|
0.079 |
| 9 |
H |
0.179 |
|
|
0.118 |
| 10 |
H |
0.179 |
|
|
0.118 |
| 11 |
H |
0.185 |
|
|
0.119 |
| 12 |
H |
0.185 |
|
|
0.119 |
| 13 |
H |
0.181 |
|
|
0.115 |
| 14 |
H |
0.181 |
|
|
0.115 |
| 15 |
H |
0.188 |
|
|
0.065 |
| 16 |
H |
0.188 |
|
|
0.065 |
| 17 |
H |
0.180 |
|
|
0.022 |
| 18 |
H |
0.180 |
|
|
0.022 |
| 19 |
H |
0.180 |
|
|
0.127 |
| 20 |
H |
0.180 |
|
|
0.127 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.003 |
-0.031 |
0.000 |
0.031 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.014 |
-0.025 |
0.000 |
0.028 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.697 |
-0.210 |
0.000 |
| y |
-0.210 |
-41.724 |
0.000 |
| z |
0.000 |
0.000 |
-41.627 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.022 |
-0.210 |
0.000 |
| y |
-0.210 |
-0.062 |
0.000 |
| z |
0.000 |
0.000 |
0.083 |
|
| Polar |
|---|
| 3z2-r2 | 0.166 |
|---|
| x2-y2 | 0.027 |
|---|
| xy | -0.210 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
197.651 |
| (<r2>)1/2 |
14.059 |