Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -234.573654 |
| Energy at 298.15K | -234.585970 |
| HF Energy | -234.573654 |
| Nuclear repulsion energy | 229.329816 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3157 |
3046 |
56.26 |
|
|
|
| 2 |
A |
3136 |
3026 |
8.13 |
|
|
|
| 3 |
A |
3126 |
3016 |
19.80 |
|
|
|
| 4 |
A |
3113 |
3004 |
48.34 |
|
|
|
| 5 |
A |
3110 |
3001 |
36.13 |
|
|
|
| 6 |
A |
3095 |
2986 |
1.64 |
|
|
|
| 7 |
A |
3076 |
2968 |
7.08 |
|
|
|
| 8 |
A |
3074 |
2966 |
19.54 |
|
|
|
| 9 |
A |
3047 |
2940 |
26.00 |
|
|
|
| 10 |
A |
3044 |
2937 |
22.88 |
|
|
|
| 11 |
A |
3037 |
2931 |
23.17 |
|
|
|
| 12 |
A |
3028 |
2921 |
17.57 |
|
|
|
| 13 |
A |
1732 |
1671 |
0.60 |
|
|
|
| 14 |
A |
1572 |
1517 |
7.85 |
|
|
|
| 15 |
A |
1567 |
1512 |
8.49 |
|
|
|
| 16 |
A |
1562 |
1507 |
7.13 |
|
|
|
| 17 |
A |
1557 |
1502 |
4.97 |
|
|
|
| 18 |
A |
1555 |
1500 |
2.49 |
|
|
|
| 19 |
A |
1542 |
1488 |
0.85 |
|
|
|
| 20 |
A |
1470 |
1418 |
7.57 |
|
|
|
| 21 |
A |
1463 |
1411 |
4.06 |
|
|
|
| 22 |
A |
1460 |
1409 |
5.80 |
|
|
|
| 23 |
A |
1394 |
1345 |
1.12 |
|
|
|
| 24 |
A |
1364 |
1316 |
0.07 |
|
|
|
| 25 |
A |
1347 |
1299 |
0.75 |
|
|
|
| 26 |
A |
1332 |
1285 |
0.89 |
|
|
|
| 27 |
A |
1297 |
1252 |
0.39 |
|
|
|
| 28 |
A |
1197 |
1155 |
2.66 |
|
|
|
| 29 |
A |
1133 |
1093 |
2.91 |
|
|
|
| 30 |
A |
1103 |
1064 |
0.02 |
|
|
|
| 31 |
A |
1094 |
1056 |
1.08 |
|
|
|
| 32 |
A |
1036 |
1000 |
3.27 |
|
|
|
| 33 |
A |
1027 |
991 |
1.53 |
|
|
|
| 34 |
A |
994 |
959 |
7.29 |
|
|
|
| 35 |
A |
924 |
891 |
1.67 |
|
|
|
| 36 |
A |
904 |
872 |
10.44 |
|
|
|
| 37 |
A |
883 |
852 |
4.49 |
|
|
|
| 38 |
A |
769 |
742 |
2.85 |
|
|
|
| 39 |
A |
727 |
702 |
38.20 |
|
|
|
| 40 |
A |
587 |
566 |
8.84 |
|
|
|
| 41 |
A |
480 |
463 |
1.17 |
|
|
|
| 42 |
A |
370 |
357 |
0.04 |
|
|
|
| 43 |
A |
297 |
287 |
0.05 |
|
|
|
| 44 |
A |
240 |
232 |
0.00 |
|
|
|
| 45 |
A |
203 |
196 |
0.02 |
|
|
|
| 46 |
A |
133 |
129 |
0.35 |
|
|
|
| 47 |
A |
91 |
88 |
0.01 |
|
|
|
| 48 |
A |
47 |
45 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36745.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35455.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.472 |
0.958 |
-0.075 |
| C2 |
2.002 |
-0.476 |
-0.143 |
| C3 |
0.781 |
-0.942 |
0.135 |
| C4 |
-0.432 |
-0.149 |
0.565 |
| C5 |
-1.586 |
-0.250 |
-0.464 |
| C6 |
-2.847 |
0.509 |
-0.006 |
| H7 |
1.672 |
1.640 |
0.225 |
| H8 |
3.297 |
1.060 |
0.643 |
| H9 |
2.852 |
1.284 |
-1.053 |
| H10 |
2.767 |
-1.188 |
-0.453 |
| H11 |
0.603 |
-2.014 |
0.034 |
| H12 |
-0.174 |
0.906 |
0.715 |
| H13 |
-0.791 |
-0.533 |
1.532 |
| H14 |
-1.238 |
0.147 |
-1.426 |
| H15 |
-1.830 |
-1.309 |
-0.626 |
| H16 |
-3.650 |
0.424 |
-0.747 |
| H17 |
-3.219 |
0.108 |
0.945 |
| H18 |
-2.629 |
1.574 |
0.139 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
| C1 | | 1.5101 | 2.5523 | 3.1723 | 4.2513 | 5.3380 | 1.0935 | 1.0986 | 1.0984 | 2.1986 | 3.5119 | 2.7613 | 3.9308 | 4.0305 | 4.8940 | 6.1811 | 5.8434 | 5.1418 |
C2 | 1.5101 | | 1.3368 | 2.5558 | 3.6094 | 4.9499 | 2.1731 | 2.1566 | 2.1558 | 1.0899 | 2.0866 | 2.7166 | 3.2572 | 3.5404 | 3.9519 | 5.7548 | 5.3649 | 5.0720 | C3 | 2.5523 | 1.3368 | | 1.5114 | 2.5373 | 3.9096 | 2.7336 | 3.2552 | 3.2647 | 2.0860 | 1.0907 | 2.1596 | 2.1419 | 2.7749 | 2.7446 | 4.7195 | 4.2135 | 4.2373 | C4 | 3.1723 | 2.5558 | 1.5114 | | 1.5494 | 2.5672 | 2.7823 | 3.9201 | 3.9308 | 3.5138 | 2.1979 | 1.0963 | 1.1011 | 2.1682 | 2.1730 | 3.5221 | 2.8245 | 2.8240 | C5 | 4.2513 | 3.6094 | 2.5373 | 1.5494 | | 1.5416 | 3.8293 | 5.1750 | 4.7322 | 4.4524 | 2.8545 | 2.1730 | 2.1670 | 1.0975 | 1.0987 | 2.1897 | 2.1865 | 2.1862 | C6 | 5.3380 | 4.9499 | 3.9096 | 2.5672 | 1.5416 | | 4.6642 | 6.2023 | 5.8457 | 5.8815 | 4.2739 | 2.7971 | 2.7710 | 2.1762 | 2.1732 | 1.0962 | 1.0971 | 1.0970 | H7 | 1.0935 | 2.1731 | 2.7336 | 2.7823 | 3.8293 | 4.6642 | | 1.7751 | 1.7755 | 3.1077 | 3.8119 | 2.0457 | 3.5352 | 3.6643 | 4.6575 | 5.5448 | 5.1756 | 4.3020 | H8 | 1.0986 | 2.1566 | 3.2552 | 3.9201 | 5.1750 | 6.2023 | 1.7751 | | 1.7674 | 2.5561 | 4.1318 | 3.4745 | 4.4762 | 5.0674 | 5.7888 | 7.1125 | 6.5916 | 5.9689 | H9 | 1.0984 | 2.1558 | 3.2647 | 3.9308 | 4.7322 | 5.8457 | 1.7755 | 1.7674 | | 2.5452 | 4.1369 | 3.5243 | 4.8222 | 4.2615 | 5.3694 | 6.5650 | 6.4982 | 5.6159 | H10 | 2.1986 | 1.0899 | 2.0860 | 3.5138 | 4.4524 | 5.8815 | 3.1077 | 2.5561 | 2.5452 | | 2.3666 | 3.7942 | 4.1263 | 4.3322 | 4.6021 | 6.6223 | 6.2817 | 6.0900 | H11 | 3.5119 | 2.0866 | 1.0907 | 2.1979 | 2.8545 | 4.2739 | 3.8119 | 4.1318 | 4.1369 | 2.3666 | | 3.0972 | 2.5256 | 3.1922 | 2.6181 | 4.9638 | 4.4651 | 4.8297 | H12 | 2.7613 | 2.7166 | 2.1596 | 1.0963 | 2.1730 | 2.7971 | 2.0457 | 3.4745 | 3.5243 | 3.7942 | 3.0972 | | 1.7667 | 2.5087 | 3.0744 | 3.8017 | 3.1565 | 2.6085 | H13 | 3.9308 | 3.2572 | 2.1419 | 1.1011 | 2.1670 | 2.7710 | 3.5352 | 4.4762 | 4.8222 | 4.1263 | 2.5256 | 1.7667 | | 3.0681 | 2.5182 | 3.7797 | 2.5787 | 3.1239 | H14 | 4.0305 | 3.5404 | 2.7749 | 2.1682 | 1.0975 | 2.1762 | 3.6643 | 5.0674 | 4.2615 | 4.3322 | 3.1922 | 2.5087 | 3.0681 | | 1.7637 | 2.5203 | 3.0899 | 2.5336 | H15 | 4.8940 | 3.9519 | 2.7446 | 2.1730 | 1.0987 | 2.1732 | 4.6575 | 5.7888 | 5.3694 | 4.6021 | 2.6181 | 3.0744 | 2.5182 | 1.7637 | | 2.5157 | 2.5308 | 3.0879 | H16 | 6.1811 | 5.7548 | 4.7195 | 3.5221 | 2.1897 | 1.0962 | 5.5448 | 7.1125 | 6.5650 | 6.6223 | 4.9638 | 3.8017 | 3.7797 | 2.5203 | 2.5157 | | 1.7753 | 1.7748 | H17 | 5.8434 | 5.3649 | 4.2135 | 2.8245 | 2.1865 | 1.0971 | 5.1756 | 6.5916 | 6.4982 | 6.2817 | 4.4651 | 3.1565 | 2.5787 | 3.0899 | 2.5308 | 1.7753 | | 1.7743 | H18 | 5.1418 | 5.0720 | 4.2373 | 2.8240 | 2.1862 | 1.0970 | 4.3020 | 5.9689 | 5.6159 | 6.0900 | 4.8297 | 2.6085 | 3.1239 | 2.5336 | 3.0879 | 1.7748 | 1.7743 | |
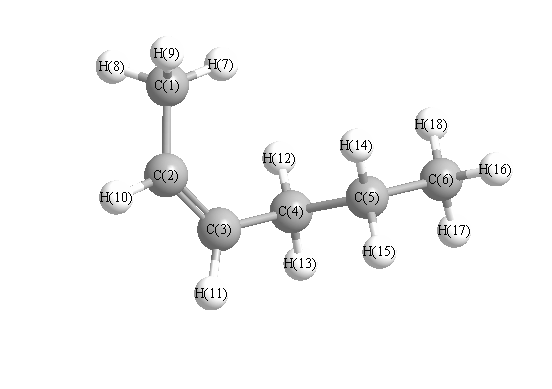 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
127.306 |
|
C1 |
C2 |
H10 |
114.513 |
| C2 |
C1 |
H7 |
112.179 |
|
C2 |
C1 |
H8 |
110.534 |
| C2 |
C1 |
H9 |
110.480 |
|
C2 |
C3 |
C4 |
127.507 |
| C2 |
C3 |
H11 |
118.179 |
|
C3 |
C2 |
H10 |
118.180 |
| C3 |
C4 |
C5 |
111.982 |
|
C3 |
C4 |
H12 |
110.813 |
| C3 |
C4 |
H13 |
109.132 |
|
C4 |
C3 |
H11 |
114.306 |
| C4 |
C5 |
C6 |
112.312 |
|
C4 |
C5 |
H14 |
108.808 |
| C4 |
C5 |
H15 |
109.108 |
|
C5 |
C4 |
H12 |
109.238 |
| C5 |
C4 |
H13 |
108.505 |
|
C5 |
C6 |
H16 |
111.099 |
| C5 |
C6 |
H17 |
110.785 |
|
C5 |
C6 |
H18 |
110.771 |
| C6 |
C5 |
H14 |
109.956 |
|
C6 |
C5 |
H15 |
109.650 |
| H7 |
C1 |
H8 |
108.152 |
|
H7 |
C1 |
H9 |
108.202 |
| H8 |
C1 |
H9 |
107.118 |
|
H12 |
C4 |
H13 |
107.020 |
| H14 |
C5 |
H16 |
94.206 |
|
H16 |
C6 |
H17 |
108.082 |
| H16 |
C6 |
H18 |
108.047 |
|
H17 |
C6 |
H18 |
107.934 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.611 |
|
|
|
| 2 |
C |
-0.168 |
|
|
|
| 3 |
C |
-0.146 |
|
|
|
| 4 |
C |
-0.430 |
|
|
|
| 5 |
C |
-0.367 |
|
|
|
| 6 |
C |
-0.553 |
|
|
|
| 7 |
H |
0.199 |
|
|
|
| 8 |
H |
0.200 |
|
|
|
| 9 |
H |
0.200 |
|
|
|
| 10 |
H |
0.177 |
|
|
|
| 11 |
H |
0.174 |
|
|
|
| 12 |
H |
0.193 |
|
|
|
| 13 |
H |
0.195 |
|
|
|
| 14 |
H |
0.194 |
|
|
|
| 15 |
H |
0.185 |
|
|
|
| 16 |
H |
0.187 |
|
|
|
| 17 |
H |
0.185 |
|
|
|
| 18 |
H |
0.185 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.045 |
0.191 |
-0.040 |
0.200 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.590 |
0.753 |
-0.721 |
| y |
0.753 |
-38.853 |
0.172 |
| z |
-0.721 |
0.172 |
-41.060 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.366 |
0.753 |
-0.721 |
| y |
0.753 |
0.972 |
0.172 |
| z |
-0.721 |
0.172 |
-2.338 |
|
| Polar |
|---|
| 3z2-r2 | -4.677 |
|---|
| x2-y2 | 0.263 |
|---|
| xy | 0.753 |
|---|
| xz | -0.721 |
|---|
| yz | 0.172 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.023 |
0.721 |
-0.711 |
| y |
0.721 |
8.439 |
0.024 |
| z |
-0.711 |
0.024 |
6.796 |
<r2> (average value of r
2) Å
2
| <r2> |
261.687 |
| (<r2>)1/2 |
16.177 |
