Jump to
S1C2
Vibrational Frequencies calculated at M06-2X/3-21G
Geometric Data calculated at M06-2X/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at M06-2X/3-21G
| | hartrees |
|---|
| Energy at 0K | -259.420603 |
| Energy at 298.15K | -259.425254 |
| HF Energy | -259.420603 |
| Nuclear repulsion energy | 123.762188 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at M06-2X/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3503 |
3359 |
77.64 |
|
|
|
| 2 |
A' |
1566 |
1501 |
72.97 |
|
|
|
| 3 |
A' |
1269 |
1217 |
247.61 |
|
|
|
| 4 |
A' |
952 |
913 |
5.98 |
|
|
|
| 5 |
A' |
712 |
683 |
25.47 |
|
|
|
| 6 |
A' |
643 |
616 |
19.82 |
|
|
|
| 7 |
A' |
321 |
307 |
413.76 |
|
|
|
| 8 |
A" |
3683 |
3531 |
102.84 |
|
|
|
| 9 |
A" |
1517 |
1454 |
131.81 |
|
|
|
| 10 |
A" |
1121 |
1075 |
53.62 |
|
|
|
| 11 |
A" |
536 |
514 |
6.10 |
|
|
|
| 12 |
A" |
454 |
435 |
7.81 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8138.4 cm
-1
Scaled (by 0.9587) Zero Point Vibrational Energy (zpe) 7802.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at M06-2X/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.036 |
-1.255 |
0.000 |
| N2 |
0.006 |
0.140 |
0.000 |
| O3 |
0.006 |
0.696 |
1.143 |
| O4 |
0.006 |
0.696 |
-1.143 |
| H5 |
-0.196 |
-1.662 |
-0.899 |
| H6 |
-0.196 |
-1.662 |
0.899 |
Atom - Atom Distances (Å)
| |
N1 |
N2 |
O3 |
O4 |
H5 |
H6 |
| N1 | | 1.3948 | 2.2611 | 2.2611 | 1.0141 | 1.0141 |
N2 | 1.3948 | | 1.2713 | 1.2713 | 2.0237 | 2.0237 | O3 | 2.2611 | 1.2713 | | 2.2865 | 3.1260 | 2.3788 | O4 | 2.2611 | 1.2713 | 2.2865 | | 2.3788 | 3.1260 | H5 | 1.0141 | 2.0237 | 3.1260 | 2.3788 | | 1.7988 | H6 | 1.0141 | 2.0237 | 2.3788 | 3.1260 | 1.7988 | |
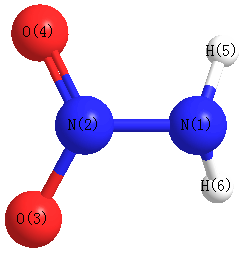 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
N2 |
O3 |
115.935 |
|
N1 |
N2 |
O4 |
115.935 |
| N2 |
N1 |
H5 |
113.354 |
|
N2 |
N1 |
H6 |
113.354 |
| O3 |
N2 |
O4 |
128.118 |
|
H5 |
N1 |
H6 |
124.969 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.481 |
|
|
|
| 2 |
N |
0.399 |
|
|
|
| 3 |
O |
-0.323 |
|
|
|
| 4 |
O |
-0.323 |
|
|
|
| 5 |
H |
0.364 |
|
|
|
| 6 |
H |
0.364 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.993 |
-4.086 |
0.000 |
4.205 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-22.296 |
1.694 |
0.000 |
| y |
1.694 |
-19.403 |
0.000 |
| z |
0.000 |
0.000 |
-23.937 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.627 |
1.694 |
0.000 |
| y |
1.694 |
3.714 |
0.000 |
| z |
0.000 |
0.000 |
-3.087 |
|
| Polar |
|---|
| 3z2-r2 | -6.175 |
|---|
| x2-y2 | -2.894 |
|---|
| xy | 1.694 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.791 |
0.064 |
0.000 |
| y |
0.064 |
3.547 |
0.000 |
| z |
0.000 |
0.000 |
4.097 |
<r2> (average value of r
2) Å
2
| <r2> |
60.709 |
| (<r2>)1/2 |
7.792 |
