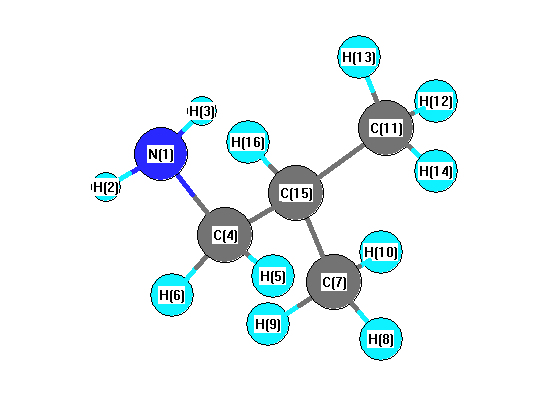
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3508 |
3385 |
0.67 |
|
|
|
| 2 |
A |
3416 |
3296 |
2.59 |
|
|
|
| 3 |
A |
3119 |
3009 |
27.25 |
|
|
|
| 4 |
A |
3109 |
3000 |
63.68 |
|
|
|
| 5 |
A |
3103 |
2994 |
40.44 |
|
|
|
| 6 |
A |
3095 |
2987 |
13.31 |
|
|
|
| 7 |
A |
3076 |
2968 |
20.43 |
|
|
|
| 8 |
A |
3063 |
2956 |
6.11 |
|
|
|
| 9 |
A |
3039 |
2933 |
20.07 |
|
|
|
| 10 |
A |
3035 |
2928 |
24.80 |
|
|
|
| 11 |
A |
2929 |
2826 |
74.29 |
|
|
|
| 12 |
A |
1720 |
1660 |
19.90 |
|
|
|
| 13 |
A |
1583 |
1527 |
15.83 |
|
|
|
| 14 |
A |
1570 |
1515 |
5.06 |
|
|
|
| 15 |
A |
1561 |
1506 |
2.62 |
|
|
|
| 16 |
A |
1559 |
1504 |
0.64 |
|
|
|
| 17 |
A |
1548 |
1494 |
1.15 |
|
|
|
| 18 |
A |
1465 |
1413 |
12.47 |
|
|
|
| 19 |
A |
1441 |
1390 |
8.84 |
|
|
|
| 20 |
A |
1426 |
1376 |
2.65 |
|
|
|
| 21 |
A |
1409 |
1360 |
5.69 |
|
|
|
| 22 |
A |
1373 |
1325 |
1.58 |
|
|
|
| 23 |
A |
1342 |
1294 |
6.77 |
|
|
|
| 24 |
A |
1261 |
1216 |
0.69 |
|
|
|
| 25 |
A |
1214 |
1172 |
4.59 |
|
|
|
| 26 |
A |
1176 |
1135 |
3.91 |
|
|
|
| 27 |
A |
1092 |
1054 |
2.57 |
|
|
|
| 28 |
A |
1070 |
1032 |
15.19 |
|
|
|
| 29 |
A |
991 |
956 |
0.51 |
|
|
|
| 30 |
A |
962 |
928 |
2.23 |
|
|
|
| 31 |
A |
928 |
896 |
0.56 |
|
|
|
| 32 |
A |
911 |
879 |
4.87 |
|
|
|
| 33 |
A |
799 |
771 |
9.27 |
|
|
|
| 34 |
A |
688 |
664 |
177.41 |
|
|
|
| 35 |
A |
485 |
468 |
10.37 |
|
|
|
| 36 |
A |
417 |
402 |
0.80 |
|
|
|
| 37 |
A |
363 |
350 |
0.43 |
|
|
|
| 38 |
A |
292 |
281 |
6.22 |
|
|
|
| 39 |
A |
265 |
256 |
2.01 |
|
|
|
| 40 |
A |
237 |
229 |
0.65 |
|
|
|
| 41 |
A |
186 |
179 |
50.01 |
|
|
|
| 42 |
A |
110 |
107 |
12.96 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32966.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 31809.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.663 |
|
|
-1.041 |
| 2 |
H |
0.262 |
|
|
0.390 |
| 3 |
H |
0.264 |
|
|
0.391 |
| 4 |
C |
-0.196 |
|
|
0.261 |
| 5 |
H |
0.153 |
|
|
-0.029 |
| 6 |
H |
0.190 |
|
|
0.023 |
| 7 |
C |
-0.536 |
|
|
-0.572 |
| 8 |
H |
0.179 |
|
|
0.132 |
| 9 |
H |
0.180 |
|
|
0.118 |
| 10 |
H |
0.185 |
|
|
0.156 |
| 11 |
C |
-0.539 |
|
|
-0.394 |
| 12 |
H |
0.184 |
|
|
0.102 |
| 13 |
H |
0.192 |
|
|
0.090 |
| 14 |
H |
0.172 |
|
|
0.101 |
| 15 |
C |
-0.227 |
|
|
0.257 |
| 16 |
H |
0.199 |
|
|
0.014 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.556 |
-0.411 |
1.128 |
1.323 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.571 |
-0.403 |
1.155 |
1.350 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.103 |
1.026 |
-2.785 |
| y |
1.026 |
-32.929 |
0.840 |
| z |
-2.785 |
0.840 |
-35.047 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.885 |
1.026 |
-2.785 |
| y |
1.026 |
0.146 |
0.840 |
| z |
-2.785 |
0.840 |
-3.031 |
|
| Polar |
|---|
| 3z2-r2 | -6.063 |
|---|
| x2-y2 | 1.826 |
|---|
| xy | 1.026 |
|---|
| xz | -2.785 |
|---|
| yz | 0.840 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.832 |
0.017 |
0.081 |
| y |
0.017 |
7.122 |
0.141 |
| z |
0.081 |
0.141 |
6.062 |
<r2> (average value of r
2) Å
2
| <r2> |
151.024 |
| (<r2>)1/2 |
12.289 |