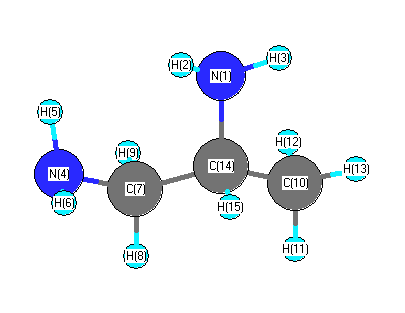
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3527 |
3404 |
0.25 |
|
|
|
| 2 |
A |
3480 |
3358 |
1.50 |
|
|
|
| 3 |
A |
3418 |
3298 |
1.65 |
|
|
|
| 4 |
A |
3387 |
3268 |
7.45 |
|
|
|
| 5 |
A |
3128 |
3019 |
31.69 |
|
|
|
| 6 |
A |
3107 |
2998 |
18.59 |
|
|
|
| 7 |
A |
3105 |
2996 |
29.69 |
|
|
|
| 8 |
A |
3050 |
2943 |
41.27 |
|
|
|
| 9 |
A |
3045 |
2938 |
18.65 |
|
|
|
| 10 |
A |
2907 |
2805 |
88.88 |
|
|
|
| 11 |
A |
1709 |
1649 |
2.01 |
|
|
|
| 12 |
A |
1703 |
1644 |
47.30 |
|
|
|
| 13 |
A |
1569 |
1514 |
4.80 |
|
|
|
| 14 |
A |
1554 |
1500 |
5.38 |
|
|
|
| 15 |
A |
1524 |
1471 |
1.72 |
|
|
|
| 16 |
A |
1452 |
1401 |
12.08 |
|
|
|
| 17 |
A |
1421 |
1371 |
6.03 |
|
|
|
| 18 |
A |
1419 |
1369 |
3.17 |
|
|
|
| 19 |
A |
1390 |
1341 |
8.36 |
|
|
|
| 20 |
A |
1369 |
1321 |
10.61 |
|
|
|
| 21 |
A |
1287 |
1242 |
1.33 |
|
|
|
| 22 |
A |
1220 |
1177 |
2.75 |
|
|
|
| 23 |
A |
1165 |
1124 |
0.23 |
|
|
|
| 24 |
A |
1086 |
1048 |
11.17 |
|
|
|
| 25 |
A |
1065 |
1027 |
5.30 |
|
|
|
| 26 |
A |
1024 |
988 |
1.01 |
|
|
|
| 27 |
A |
945 |
912 |
5.01 |
|
|
|
| 28 |
A |
892 |
861 |
3.50 |
|
|
|
| 29 |
A |
838 |
808 |
33.16 |
|
|
|
| 30 |
A |
729 |
703 |
138.52 |
|
|
|
| 31 |
A |
623 |
601 |
257.98 |
|
|
|
| 32 |
A |
491 |
474 |
5.62 |
|
|
|
| 33 |
A |
461 |
445 |
31.44 |
|
|
|
| 34 |
A |
387 |
373 |
75.47 |
|
|
|
| 35 |
A |
364 |
351 |
1.05 |
|
|
|
| 36 |
A |
258 |
249 |
6.82 |
|
|
|
| 37 |
A |
235 |
227 |
7.00 |
|
|
|
| 38 |
A |
210 |
203 |
92.61 |
|
|
|
| 39 |
A |
115 |
111 |
11.82 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30328.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 29263.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.651 |
|
|
-0.952 |
| 2 |
H |
0.244 |
|
|
0.336 |
| 3 |
H |
0.260 |
|
|
0.367 |
| 4 |
N |
-0.664 |
|
|
-0.977 |
| 5 |
H |
0.268 |
|
|
0.377 |
| 6 |
H |
0.258 |
|
|
0.364 |
| 7 |
C |
-0.181 |
|
|
0.196 |
| 8 |
H |
0.184 |
|
|
0.021 |
| 9 |
H |
0.206 |
|
|
0.051 |
| 10 |
C |
-0.535 |
|
|
-0.575 |
| 11 |
H |
0.183 |
|
|
0.119 |
| 12 |
H |
0.203 |
|
|
0.156 |
| 13 |
H |
0.178 |
|
|
0.160 |
| 14 |
C |
-0.102 |
|
|
0.381 |
| 15 |
H |
0.148 |
|
|
-0.024 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.732 |
1.286 |
1.677 |
2.236 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.711 |
1.292 |
1.658 |
2.219 |
Electric Quadrupole moment
Quadrupole components in D Å
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.139 |
-0.288 |
-0.139 |
| y |
-0.288 |
6.577 |
-0.204 |
| z |
-0.139 |
-0.204 |
5.630 |
<r2> (average value of r
2) Å
2
| <r2> |
144.572 |
| (<r2>)1/2 |
12.024 |