Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Electronic state
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -231.197031 |
| Energy at 298.15K | -231.205297 |
| Nuclear repulsion energy | 175.122052 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3160 |
3049 |
9.73 |
|
|
|
| 2 |
A |
3139 |
3029 |
21.20 |
|
|
|
| 3 |
A |
3135 |
3025 |
18.57 |
|
|
|
| 4 |
A |
3105 |
2996 |
8.82 |
|
|
|
| 5 |
A |
3070 |
2962 |
17.60 |
|
|
|
| 6 |
A |
3064 |
2957 |
5.50 |
|
|
|
| 7 |
A |
3053 |
2946 |
6.42 |
|
|
|
| 8 |
A |
3038 |
2931 |
11.08 |
|
|
|
| 9 |
A |
1748 |
1687 |
69.60 |
|
|
|
| 10 |
A |
1564 |
1509 |
8.85 |
|
|
|
| 11 |
A |
1563 |
1508 |
8.20 |
|
|
|
| 12 |
A |
1545 |
1491 |
14.11 |
|
|
|
| 13 |
A |
1523 |
1470 |
21.70 |
|
|
|
| 14 |
A |
1509 |
1456 |
9.39 |
|
|
|
| 15 |
A |
1470 |
1419 |
11.70 |
|
|
|
| 16 |
A |
1431 |
1380 |
35.18 |
|
|
|
| 17 |
A |
1377 |
1329 |
20.94 |
|
|
|
| 18 |
A |
1327 |
1280 |
0.00 |
|
|
|
| 19 |
A |
1192 |
1150 |
84.29 |
|
|
|
| 20 |
A |
1165 |
1124 |
0.34 |
|
|
|
| 21 |
A |
1124 |
1084 |
6.94 |
|
|
|
| 22 |
A |
1009 |
974 |
1.33 |
|
|
|
| 23 |
A |
1001 |
965 |
6.78 |
|
|
|
| 24 |
A |
946 |
913 |
15.08 |
|
|
|
| 25 |
A |
785 |
757 |
4.09 |
|
|
|
| 26 |
A |
751 |
725 |
2.58 |
|
|
|
| 27 |
A |
584 |
563 |
5.24 |
|
|
|
| 28 |
A |
493 |
475 |
0.44 |
|
|
|
| 29 |
A |
407 |
393 |
5.55 |
|
|
|
| 30 |
A |
258 |
249 |
5.82 |
|
|
|
| 31 |
A |
179 |
173 |
0.12 |
|
|
|
| 32 |
A |
112 |
108 |
0.08 |
|
|
|
| 33 |
A |
46 |
44 |
0.21 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 24934.9 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 24059.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
1.894 |
-0.496 |
0.000 |
| C2 |
0.524 |
0.173 |
-0.000 |
| C3 |
-0.668 |
-0.783 |
-0.000 |
| C4 |
-2.007 |
-0.029 |
0.000 |
| O5 |
0.379 |
1.398 |
-0.000 |
| H6 |
2.674 |
0.268 |
-0.000 |
| H7 |
2.003 |
-1.134 |
0.886 |
| H8 |
2.003 |
-1.136 |
-0.884 |
| H9 |
-2.848 |
-0.728 |
0.000 |
| H10 |
-2.079 |
0.613 |
0.883 |
| H11 |
-2.080 |
0.615 |
-0.881 |
| H12 |
-0.584 |
-1.436 |
-0.881 |
| H13 |
-0.584 |
-1.437 |
0.879 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
O5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.5247 | 2.5785 | 3.9290 | 2.4258 | 1.0916 | 1.0969 | 1.0969 | 4.7479 | 4.2189 | 4.2194 | 2.7935 | 2.7927 |
C2 | 1.5247 | | 1.5285 | 2.5393 | 1.2337 | 2.1517 | 2.1632 | 2.1633 | 3.4908 | 2.7843 | 2.7843 | 2.1436 | 2.1435 | C3 | 2.5785 | 1.5285 | | 1.5364 | 2.4196 | 3.5035 | 2.8359 | 2.8353 | 2.1804 | 2.1727 | 2.1727 | 1.0994 | 1.0994 | C4 | 3.9290 | 2.5393 | 1.5364 | | 2.7802 | 4.6902 | 4.2522 | 4.2525 | 1.0941 | 1.0937 | 1.0937 | 2.1865 | 2.1865 | O5 | 2.4258 | 1.2337 | 2.4196 | 2.7802 | | 2.5581 | 3.1359 | 3.1367 | 3.8649 | 2.7276 | 2.7270 | 3.1207 | 3.1210 | H6 | 1.0916 | 2.1517 | 3.5035 | 4.6902 | 2.5581 | | 1.7893 | 1.7895 | 5.6111 | 4.8469 | 4.8470 | 3.7810 | 3.7808 | H7 | 1.0969 | 2.1632 | 2.8359 | 4.2522 | 3.1359 | 1.7893 | | 1.7701 | 4.9474 | 4.4402 | 4.7797 | 3.1470 | 2.6041 | H8 | 1.0969 | 2.1633 | 2.8353 | 4.2525 | 3.1367 | 1.7895 | 1.7701 | | 4.9473 | 4.7795 | 4.4418 | 2.6044 | 3.1448 | H9 | 4.7479 | 3.4908 | 2.1804 | 1.0941 | 3.8649 | 5.6111 | 4.9474 | 4.9473 | | 1.7807 | 1.7806 | 2.5298 | 2.5299 | H10 | 4.2189 | 2.7843 | 2.1727 | 1.0937 | 2.7276 | 4.8469 | 4.4402 | 4.7795 | 1.7807 | | 1.7639 | 3.0896 | 2.5380 | H11 | 4.2194 | 2.7843 | 2.1727 | 1.0937 | 2.7270 | 4.8470 | 4.7797 | 4.4418 | 1.7806 | 1.7639 | | 2.5382 | 3.0896 | H12 | 2.7935 | 2.1436 | 1.0994 | 2.1865 | 3.1207 | 3.7810 | 3.1470 | 2.6044 | 2.5298 | 3.0896 | 2.5382 | | 1.7594 | H13 | 2.7927 | 2.1435 | 1.0994 | 2.1865 | 3.1210 | 3.7808 | 2.6041 | 3.1448 | 2.5299 | 2.5380 | 3.0896 | 1.7594 | |
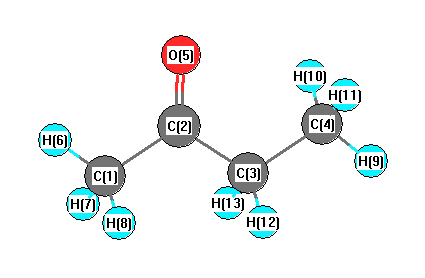 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
115.247 |
|
C1 |
C2 |
O5 |
122.795 |
| C2 |
C1 |
H6 |
109.546 |
|
C2 |
C1 |
H7 |
110.136 |
| C2 |
C1 |
H8 |
110.151 |
|
C2 |
C3 |
C4 |
111.889 |
| C2 |
C3 |
H12 |
108.205 |
|
C2 |
C3 |
H13 |
108.200 |
| C3 |
C2 |
O5 |
121.958 |
|
C3 |
C4 |
H9 |
110.844 |
| C3 |
C4 |
H10 |
110.262 |
|
C3 |
C4 |
H11 |
110.260 |
| C4 |
C3 |
H12 |
111.018 |
|
C4 |
C3 |
H13 |
111.013 |
| H6 |
C1 |
H7 |
109.694 |
|
H6 |
C1 |
H8 |
109.707 |
| H7 |
C1 |
H8 |
107.581 |
|
H9 |
C4 |
H10 |
108.958 |
| H9 |
C4 |
H11 |
108.950 |
|
H10 |
C4 |
H11 |
107.487 |
| H12 |
C3 |
H13 |
106.294 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.632 |
|
|
|
| 2 |
C |
0.428 |
|
|
|
| 3 |
C |
-0.481 |
|
|
|
| 4 |
C |
-0.540 |
|
|
|
| 5 |
O |
-0.437 |
|
|
|
| 6 |
H |
0.223 |
|
|
|
| 7 |
H |
0.212 |
|
|
|
| 8 |
H |
0.212 |
|
|
|
| 9 |
H |
0.182 |
|
|
|
| 10 |
H |
0.204 |
|
|
|
| 11 |
H |
0.204 |
|
|
|
| 12 |
H |
0.211 |
|
|
|
| 13 |
H |
0.211 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.313 |
-2.563 |
0.000 |
2.582 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.023 |
-1.043 |
0.001 |
| y |
-1.043 |
-34.731 |
0.001 |
| z |
0.001 |
0.001 |
-30.573 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.629 |
-1.043 |
0.001 |
| y |
-1.043 |
-4.433 |
0.001 |
| z |
0.001 |
0.001 |
1.804 |
|
| Polar |
|---|
| 3z2-r2 | 3.609 |
|---|
| x2-y2 | 4.708 |
|---|
| xy | -1.043 |
|---|
| xz | 0.001 |
|---|
| yz | 0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.826 |
-0.151 |
-0.000 |
| y |
-0.151 |
6.511 |
0.001 |
| z |
-0.000 |
0.001 |
4.917 |
<r2> (average value of r
2) Å
2
| <r2> |
137.346 |
| (<r2>)1/2 |
11.719 |
