Jump to
S1C2
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -1067.753905 |
| Energy at 298.15K | -1067.756235 |
| HF Energy | -1067.753905 |
| Nuclear repulsion energy | 255.575003 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3231 |
3118 |
0.68 |
|
|
|
| 2 |
A' |
3012 |
2906 |
40.54 |
|
|
|
| 3 |
A' |
1740 |
1678 |
87.91 |
|
|
|
| 4 |
A' |
1434 |
1384 |
2.96 |
|
|
|
| 5 |
A' |
1207 |
1165 |
8.59 |
|
|
|
| 6 |
A' |
1015 |
980 |
32.65 |
|
|
|
| 7 |
A' |
723 |
698 |
27.70 |
|
|
|
| 8 |
A' |
412 |
397 |
5.45 |
|
|
|
| 9 |
A' |
298 |
288 |
25.47 |
|
|
|
| 10 |
A' |
231 |
223 |
1.50 |
|
|
|
| 11 |
A" |
1201 |
1158 |
24.25 |
|
|
|
| 12 |
A" |
1040 |
1003 |
21.07 |
|
|
|
| 13 |
A" |
648 |
625 |
130.40 |
|
|
|
| 14 |
A" |
283 |
274 |
3.04 |
|
|
|
| 15 |
A" |
81 |
78 |
7.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 8278.4 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 7987.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.346 |
0.261 |
0.000 |
| C2 |
-0.941 |
1.069 |
0.000 |
| H3 |
1.246 |
0.866 |
0.000 |
| Cl4 |
0.346 |
-0.812 |
1.534 |
| Cl5 |
0.346 |
-0.812 |
-1.534 |
| O6 |
-0.952 |
2.293 |
0.000 |
| H7 |
-1.837 |
0.431 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5201 | 1.0838 | 1.8721 | 1.8721 | 2.4110 | 2.1898 |
C2 | 1.5201 | | 2.1962 | 2.7478 | 2.7478 | 1.2235 | 1.0998 | H3 | 1.0838 | 2.1962 | | 2.4450 | 2.4450 | 2.6200 | 3.1130 | Cl4 | 1.8721 | 2.7478 | 2.4450 | | 3.0675 | 3.6985 | 2.9436 | Cl5 | 1.8721 | 2.7478 | 2.4450 | 3.0675 | | 3.6985 | 2.9436 | O6 | 2.4110 | 1.2235 | 2.6200 | 3.6985 | 3.6985 | | 2.0611 | H7 | 2.1898 | 1.0998 | 3.1130 | 2.9436 | 2.9436 | 2.0611 | |
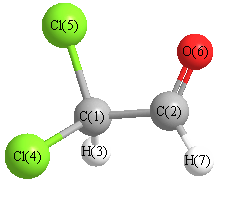 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
122.624 |
|
C1 |
C2 |
H7 |
112.417 |
| C2 |
C1 |
H3 |
113.951 |
|
C2 |
C1 |
Cl4 |
107.748 |
| C2 |
C1 |
Cl5 |
107.748 |
|
H3 |
C1 |
Cl4 |
108.669 |
| H3 |
C1 |
Cl5 |
108.669 |
|
Cl4 |
C1 |
Cl5 |
110.025 |
| O6 |
C2 |
H7 |
124.959 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.641 |
|
|
|
| 2 |
C |
0.322 |
|
|
|
| 3 |
H |
0.328 |
|
|
|
| 4 |
Cl |
0.074 |
|
|
|
| 5 |
Cl |
0.074 |
|
|
|
| 6 |
O |
-0.371 |
|
|
|
| 7 |
H |
0.214 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.070 |
0.030 |
0.000 |
0.076 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.941 |
2.814 |
0.000 |
| y |
2.814 |
-48.853 |
0.000 |
| z |
0.000 |
0.000 |
-44.645 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.808 |
2.814 |
0.000 |
| y |
2.814 |
-6.060 |
0.000 |
| z |
0.000 |
0.000 |
0.252 |
|
| Polar |
|---|
| 3z2-r2 | 0.504 |
|---|
| x2-y2 | 7.912 |
|---|
| xy | 2.814 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.936 |
-0.685 |
0.000 |
| y |
-0.685 |
6.436 |
0.000 |
| z |
0.000 |
0.000 |
7.145 |
<r2> (average value of r
2) Å
2
| <r2> |
202.952 |
| (<r2>)1/2 |
14.246 |
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -1067.752868 |
| Energy at 298.15K | |
| HF Energy | -1067.752868 |
| Nuclear repulsion energy | 258.913088 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3184 |
3072 |
1.85 |
82.41 |
0.23 |
0.37 |
| 2 |
A |
2981 |
2876 |
43.00 |
135.02 |
0.32 |
0.49 |
| 3 |
A |
1751 |
1690 |
74.47 |
14.26 |
0.44 |
0.61 |
| 4 |
A |
1424 |
1374 |
10.86 |
6.50 |
0.72 |
0.83 |
| 5 |
A |
1259 |
1215 |
11.95 |
9.57 |
0.65 |
0.79 |
| 6 |
A |
1183 |
1141 |
15.47 |
14.08 |
0.70 |
0.82 |
| 7 |
A |
1032 |
996 |
12.89 |
4.67 |
0.71 |
0.83 |
| 8 |
A |
917 |
885 |
10.25 |
5.92 |
0.54 |
0.70 |
| 9 |
A |
765 |
738 |
66.14 |
7.26 |
0.72 |
0.83 |
| 10 |
A |
594 |
574 |
47.49 |
13.39 |
0.17 |
0.28 |
| 11 |
A |
587 |
566 |
48.61 |
13.97 |
0.33 |
0.50 |
| 12 |
A |
330 |
318 |
3.10 |
4.52 |
0.26 |
0.41 |
| 13 |
A |
247 |
239 |
3.89 |
8.61 |
0.59 |
0.74 |
| 14 |
A |
219 |
212 |
1.53 |
2.64 |
0.74 |
0.85 |
| 15 |
A |
99 |
96 |
9.56 |
2.48 |
0.71 |
0.83 |
Unscaled Zero Point Vibrational Energy (zpe) 8286.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 7995.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.085 |
-0.036 |
0.576 |
| C2 |
-0.599 |
-1.351 |
0.222 |
| H3 |
0.237 |
0.095 |
1.644 |
| Cl4 |
1.813 |
-0.115 |
-0.181 |
| Cl5 |
-0.860 |
1.412 |
-0.062 |
| O6 |
-1.668 |
-1.451 |
-0.358 |
| H7 |
0.001 |
-2.211 |
0.561 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
Cl4 |
Cl5 |
O6 |
H7 |
| C1 | | 1.5240 | 1.0874 | 1.8877 | 1.8435 | 2.4381 | 2.1770 |
C2 | 1.5240 | | 2.1939 | 2.7397 | 2.7908 | 1.2201 | 1.1022 | H3 | 1.0874 | 2.1939 | | 2.4201 | 2.4192 | 3.1665 | 2.5590 | Cl4 | 1.8877 | 2.7397 | 2.4201 | | 3.0811 | 3.7319 | 2.8677 | Cl5 | 1.8435 | 2.7908 | 2.4192 | 3.0811 | | 2.9897 | 3.7767 | O6 | 2.4381 | 1.2201 | 3.1665 | 3.7319 | 2.9897 | | 2.0515 | H7 | 2.1770 | 1.1022 | 2.5590 | 2.8677 | 3.7767 | 2.0515 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
O6 |
125.007 |
|
C1 |
C2 |
H7 |
110.962 |
| C2 |
C1 |
H3 |
113.252 |
|
C2 |
C1 |
Cl4 |
106.351 |
| C2 |
C1 |
Cl5 |
111.585 |
|
H3 |
C1 |
Cl4 |
105.701 |
| H3 |
C1 |
Cl5 |
108.492 |
|
Cl4 |
C1 |
Cl5 |
111.323 |
| O6 |
C2 |
H7 |
124.031 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.659 |
|
|
|
| 2 |
C |
0.312 |
|
|
|
| 3 |
H |
0.326 |
|
|
|
| 4 |
Cl |
0.056 |
|
|
|
| 5 |
Cl |
0.107 |
|
|
|
| 6 |
O |
-0.354 |
|
|
|
| 7 |
H |
0.212 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.885 |
-1.141 |
2.339 |
2.748 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-48.349 |
-1.804 |
-0.937 |
| y |
-1.804 |
-42.668 |
-1.583 |
| z |
-0.937 |
-1.583 |
-40.432 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.799 |
-1.804 |
-0.937 |
| y |
-1.804 |
1.723 |
-1.583 |
| z |
-0.937 |
-1.583 |
5.076 |
|
| Polar |
|---|
| 3z2-r2 | 10.153 |
|---|
| x2-y2 | -5.681 |
|---|
| xy | -1.804 |
|---|
| xz | -0.937 |
|---|
| yz | -1.583 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.704 |
-0.938 |
0.144 |
| y |
-0.938 |
5.948 |
-0.455 |
| z |
0.144 |
-0.455 |
3.551 |
<r2> (average value of r
2) Å
2
| <r2> |
194.102 |
| (<r2>)1/2 |
13.932 |
