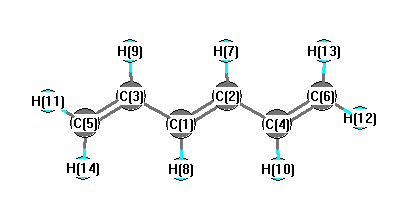
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3244 |
3130 |
0.00 |
|
|
|
| 2 |
Ag |
3161 |
3050 |
0.00 |
|
|
|
| 3 |
Ag |
3154 |
3043 |
0.00 |
|
|
|
| 4 |
Ag |
3144 |
3033 |
0.00 |
|
|
|
| 5 |
Ag |
1689 |
1630 |
0.00 |
|
|
|
| 6 |
Ag |
1652 |
1594 |
0.00 |
|
|
|
| 7 |
Ag |
1487 |
1435 |
0.00 |
|
|
|
| 8 |
Ag |
1379 |
1330 |
0.00 |
|
|
|
| 9 |
Ag |
1351 |
1304 |
0.00 |
|
|
|
| 10 |
Ag |
1240 |
1196 |
0.00 |
|
|
|
| 11 |
Ag |
973 |
938 |
0.00 |
|
|
|
| 12 |
Ag |
456 |
440 |
0.00 |
|
|
|
| 13 |
Ag |
364 |
351 |
0.00 |
|
|
|
| 14 |
Au |
1068 |
1031 |
74.25 |
|
|
|
| 15 |
Au |
983 |
949 |
31.83 |
|
|
|
| 16 |
Au |
949 |
915 |
83.21 |
|
|
|
| 17 |
Au |
717 |
692 |
13.64 |
|
|
|
| 18 |
Au |
252 |
244 |
2.14 |
|
|
|
| 19 |
Au |
100 |
97 |
0.45 |
|
|
|
| 20 |
Bg |
1039 |
1003 |
0.00 |
|
|
|
| 21 |
Bg |
957 |
924 |
0.00 |
|
|
|
| 22 |
Bg |
915 |
882 |
0.00 |
|
|
|
| 23 |
Bg |
619 |
597 |
0.00 |
|
|
|
| 24 |
Bg |
223 |
215 |
0.00 |
|
|
|
| 25 |
Bu |
3244 |
3130 |
28.05 |
|
|
|
| 26 |
Bu |
3163 |
3052 |
37.06 |
|
|
|
| 27 |
Bu |
3161 |
3050 |
14.46 |
|
|
|
| 28 |
Bu |
3144 |
3034 |
1.23 |
|
|
|
| 29 |
Bu |
1693 |
1633 |
14.57 |
|
|
|
| 30 |
Bu |
1518 |
1465 |
3.66 |
|
|
|
| 31 |
Bu |
1368 |
1320 |
3.35 |
|
|
|
| 32 |
Bu |
1336 |
1289 |
0.86 |
|
|
|
| 33 |
Bu |
1172 |
1131 |
9.48 |
|
|
|
| 34 |
Bu |
999 |
964 |
2.90 |
|
|
|
| 35 |
Bu |
566 |
547 |
4.67 |
|
|
|
| 36 |
Bu |
153 |
147 |
1.53 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26315.2 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 25391.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.182 |
|
|
-0.168 |
| 2 |
C |
-0.182 |
|
|
-0.206 |
| 3 |
C |
-0.187 |
|
|
-0.071 |
| 4 |
C |
-0.187 |
|
|
-0.047 |
| 5 |
C |
-0.374 |
|
|
-0.420 |
| 6 |
C |
-0.374 |
|
|
-0.429 |
| 7 |
H |
0.183 |
|
|
0.177 |
| 8 |
H |
0.183 |
|
|
0.171 |
| 9 |
H |
0.186 |
|
|
0.140 |
| 10 |
H |
0.186 |
|
|
0.133 |
| 11 |
H |
0.189 |
|
|
0.177 |
| 12 |
H |
0.189 |
|
|
0.178 |
| 13 |
H |
0.185 |
|
|
0.183 |
| 14 |
H |
0.185 |
|
|
0.181 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.007 |
0.003 |
0.000 |
0.008 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-33.078 |
-0.129 |
0.000 |
| y |
-0.129 |
-33.086 |
0.000 |
| z |
0.000 |
0.000 |
-41.940 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.434 |
-0.129 |
0.000 |
| y |
-0.129 |
4.423 |
0.000 |
| z |
0.000 |
0.000 |
-8.858 |
|
| Polar |
|---|
| 3z2-r2 | -17.715 |
|---|
| x2-y2 | 0.007 |
|---|
| xy | -0.129 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.231 |
4.379 |
0.000 |
| y |
4.379 |
20.621 |
0.000 |
| z |
0.000 |
0.000 |
2.438 |
<r2> (average value of r
2) Å
2
| <r2> |
257.035 |
| (<r2>)1/2 |
16.032 |