Jump to
S1C2
Vibrational Frequencies calculated at B3LYP/3-21G
Geometric Data calculated at B3LYP/3-21G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at B3LYP/3-21G
| | hartrees |
|---|
| Energy at 0K | -115.296075 |
| Energy at 298.15K | -115.297689 |
| HF Energy | -115.296075 |
| Nuclear repulsion energy | 57.470078 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3339 |
3221 |
0.46 |
|
|
|
| 2 |
A' |
3101 |
2992 |
42.13 |
|
|
|
| 3 |
A' |
1631 |
1574 |
0.45 |
|
|
|
| 4 |
A' |
1184 |
1142 |
15.60 |
|
|
|
| 5 |
A' |
995 |
961 |
12.91 |
|
|
|
| 6 |
A' |
935 |
902 |
4.85 |
|
|
|
| 7 |
A' |
664 |
641 |
92.77 |
|
|
|
| 8 |
A" |
3290 |
3174 |
2.46 |
|
|
|
| 9 |
A" |
1071 |
1033 |
18.15 |
|
|
|
| 10 |
A" |
1000 |
965 |
17.33 |
|
|
|
| 11 |
A" |
912 |
880 |
0.53 |
|
|
|
| 12 |
A" |
646 |
623 |
25.74 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9383.0 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 9053.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYP/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.037 |
0.907 |
0.000 |
| H2 |
0.756 |
1.653 |
0.000 |
| C3 |
-0.037 |
-0.426 |
0.660 |
| C4 |
-0.037 |
-0.426 |
-0.660 |
| H5 |
-0.042 |
-0.993 |
1.569 |
| H6 |
-0.042 |
-0.993 |
-1.569 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.0891 | 1.4875 | 1.4875 | 2.4647 | 2.4647 |
H2 | 1.0891 | | 2.3210 | 2.3210 | 3.1783 | 3.1783 | C3 | 1.4875 | 2.3210 | | 1.3190 | 1.0722 | 2.2999 | C4 | 1.4875 | 2.3210 | 1.3190 | | 2.2999 | 1.0722 | H5 | 2.4647 | 3.1783 | 1.0722 | 2.2999 | | 3.1387 | H6 | 2.4647 | 3.1783 | 2.2999 | 1.0722 | 3.1387 | |
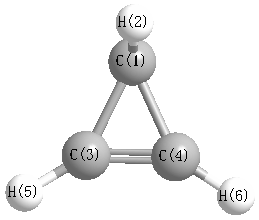 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
C4 |
63.681 |
|
C1 |
C3 |
H5 |
148.257 |
| C1 |
C4 |
C3 |
63.681 |
|
C1 |
C4 |
H6 |
148.257 |
| H2 |
C1 |
C3 |
127.856 |
|
H2 |
C1 |
C4 |
127.856 |
| C3 |
C1 |
C4 |
52.638 |
|
C3 |
C4 |
H6 |
148.061 |
| C4 |
C3 |
H5 |
148.061 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.150 |
|
|
|
| 2 |
H |
0.172 |
|
|
|
| 3 |
C |
-0.205 |
|
|
|
| 4 |
C |
-0.205 |
|
|
|
| 5 |
H |
0.194 |
|
|
|
| 6 |
H |
0.194 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.725 |
-1.293 |
0.000 |
1.482 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.274 |
1.287 |
0.000 |
| y |
1.287 |
-17.971 |
0.000 |
| z |
0.000 |
0.000 |
-15.771 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.402 |
1.287 |
0.000 |
| y |
1.287 |
-0.449 |
0.000 |
| z |
0.000 |
0.000 |
2.851 |
|
| Polar |
|---|
| 3z2-r2 | 5.702 |
|---|
| x2-y2 | -1.302 |
|---|
| xy | 1.287 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
1.815 |
0.323 |
0.000 |
| y |
0.323 |
4.278 |
0.000 |
| z |
0.000 |
0.000 |
4.399 |
<r2> (average value of r
2) Å
2
| <r2> |
33.606 |
| (<r2>)1/2 |
5.797 |
