Jump to
S1C2
S1C3
Energy calculated at MP4=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -168.292364 |
| Energy at 298.15K | -168.296223 |
| HF Energy | -167.937206 |
| Nuclear repulsion energy | 69.077974 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3503 |
3428 |
|
|
|
|
| 2 |
A' |
3344 |
3273 |
|
|
|
|
| 3 |
A' |
2982 |
2919 |
|
|
|
|
| 4 |
A' |
1709 |
1672 |
|
|
|
|
| 5 |
A' |
1423 |
1393 |
|
|
|
|
| 6 |
A' |
1301 |
1273 |
|
|
|
|
| 7 |
A' |
1167 |
1142 |
|
|
|
|
| 8 |
A' |
984 |
963 |
|
|
|
|
| 9 |
A' |
606 |
593 |
|
|
|
|
| 10 |
A" |
1048 |
1025 |
|
|
|
|
| 11 |
A" |
828 |
810 |
|
|
|
|
| 12 |
A" |
300 |
294 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9596.0 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 9392.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.400 |
0.000 |
| O2 |
-1.019 |
-0.569 |
0.000 |
| N3 |
1.218 |
0.005 |
0.000 |
| H4 |
-0.369 |
1.438 |
0.000 |
| H5 |
-1.899 |
-0.109 |
0.000 |
| H6 |
1.894 |
0.787 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.4065 | 1.2807 | 1.1015 | 1.9658 | 1.9333 |
O2 | 1.4065 | | 2.3104 | 2.1098 | 0.9925 | 3.2138 | N3 | 1.2807 | 2.3104 | | 2.1379 | 3.1193 | 1.0332 | H4 | 1.1015 | 2.1098 | 2.1379 | | 2.1757 | 2.3546 | H5 | 1.9658 | 0.9925 | 3.1193 | 2.1757 | | 3.8974 | H6 | 1.9333 | 3.2138 | 1.0332 | 2.3546 | 3.8974 | |
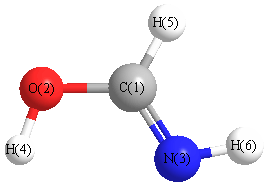 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
108.825 |
|
C1 |
N3 |
H6 |
112.907 |
| O2 |
C1 |
N3 |
118.511 |
|
O2 |
C1 |
H4 |
113.992 |
| N3 |
C1 |
H4 |
127.497 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP4=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -168.305912 |
| Energy at 298.15K | -168.310038 |
| HF Energy | -167.951771 |
| Nuclear repulsion energy | 69.440309 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3443 |
3370 |
|
|
|
|
| 2 |
A' |
3361 |
3289 |
|
|
|
|
| 3 |
A' |
3099 |
3033 |
|
|
|
|
| 4 |
A' |
1658 |
1623 |
|
|
|
|
| 5 |
A' |
1406 |
1376 |
|
|
|
|
| 6 |
A' |
1389 |
1360 |
|
|
|
|
| 7 |
A' |
1163 |
1138 |
|
|
|
|
| 8 |
A' |
997 |
975 |
|
|
|
|
| 9 |
A' |
568 |
556 |
|
|
|
|
| 10 |
A" |
1058 |
1036 |
|
|
|
|
| 11 |
A" |
829 |
811 |
|
|
|
|
| 12 |
A" |
636 |
623 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9802.5 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 9594.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.440 |
0.000 |
| O2 |
-1.135 |
-0.373 |
0.000 |
| N3 |
1.180 |
-0.065 |
0.000 |
| H4 |
-0.315 |
1.487 |
0.000 |
| H5 |
-0.804 |
-1.317 |
0.000 |
| H6 |
1.939 |
0.633 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.3964 | 1.2838 | 1.0933 | 1.9320 | 1.9489 |
O2 | 1.3964 | | 2.3357 | 2.0332 | 1.0000 | 3.2351 | N3 | 1.2838 | 2.3357 | | 2.1552 | 2.3458 | 1.0317 | H4 | 1.0933 | 2.0332 | 2.1552 | | 2.8462 | 2.4103 | H5 | 1.9320 | 1.0000 | 2.3458 | 2.8462 | | 3.3657 | H6 | 1.9489 | 3.2351 | 1.0317 | 2.4103 | 3.3657 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
106.271 |
|
C1 |
N3 |
H6 |
114.198 |
| O2 |
C1 |
N3 |
121.201 |
|
O2 |
C1 |
H4 |
108.889 |
| N3 |
C1 |
H4 |
129.910 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP4=FULL/3-21G
| | hartrees |
|---|
| Energy at 0K | -168.302720 |
| Energy at 298.15K | -168.306750 |
| HF Energy | -167.946159 |
| Nuclear repulsion energy | 68.790381 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4=FULL/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3439 |
3366 |
|
|
|
|
| 2 |
A' |
3264 |
3194 |
|
|
|
|
| 3 |
A' |
3189 |
3121 |
|
|
|
|
| 4 |
A' |
1629 |
1595 |
|
|
|
|
| 5 |
A' |
1439 |
1409 |
|
|
|
|
| 6 |
A' |
1340 |
1312 |
|
|
|
|
| 7 |
A' |
1134 |
1109 |
|
|
|
|
| 8 |
A' |
985 |
964 |
|
|
|
|
| 9 |
A' |
561 |
549 |
|
|
|
|
| 10 |
A" |
1090 |
1067 |
|
|
|
|
| 11 |
A" |
834 |
816 |
|
|
|
|
| 12 |
A" |
495 |
484 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 9699.1 cm
-1
Scaled (by 0.9788) Zero Point Vibrational Energy (zpe) 9493.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP4=FULL/3-21G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.449 |
0.000 |
| O2 |
-1.133 |
-0.394 |
0.000 |
| N3 |
1.254 |
0.167 |
0.000 |
| H4 |
-0.341 |
1.482 |
0.000 |
| H5 |
-0.841 |
-1.348 |
0.000 |
| H6 |
1.472 |
-0.850 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.4127 | 1.2851 | 1.0868 | 1.9847 | 1.9631 |
O2 | 1.4127 | | 2.4522 | 2.0362 | 0.9985 | 2.6446 | N3 | 1.2851 | 2.4522 | | 2.0662 | 2.5853 | 1.0401 | H4 | 1.0868 | 2.0362 | 2.0662 | | 2.8739 | 2.9528 | H5 | 1.9847 | 0.9985 | 2.5853 | 2.8739 | | 2.3654 | H6 | 1.9631 | 2.6446 | 1.0401 | 2.9528 | 2.3654 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
O2 |
H5 |
109.600 |
|
C1 |
N3 |
H6 |
114.777 |
| O2 |
C1 |
N3 |
130.664 |
|
O2 |
C1 |
H4 |
108.393 |
| N3 |
C1 |
H4 |
120.943 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
