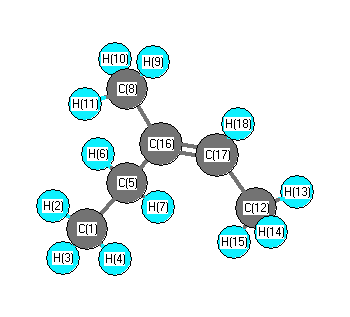
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3143 |
3033 |
51.62 |
|
|
|
| 2 |
A |
3129 |
3019 |
32.44 |
|
|
|
| 3 |
A |
3124 |
3014 |
8.08 |
|
|
|
| 4 |
A |
3117 |
3007 |
8.95 |
|
|
|
| 5 |
A |
3115 |
3006 |
41.08 |
|
|
|
| 6 |
A |
3094 |
2985 |
5.18 |
|
|
|
| 7 |
A |
3075 |
2967 |
28.28 |
|
|
|
| 8 |
A |
3070 |
2962 |
23.19 |
|
|
|
| 9 |
A |
3053 |
2946 |
23.59 |
|
|
|
| 10 |
A |
3046 |
2939 |
22.51 |
|
|
|
| 11 |
A |
3034 |
2927 |
37.09 |
|
|
|
| 12 |
A |
3032 |
2925 |
17.49 |
|
|
|
| 13 |
A |
1742 |
1681 |
0.31 |
|
|
|
| 14 |
A |
1574 |
1519 |
13.87 |
|
|
|
| 15 |
A |
1569 |
1514 |
7.72 |
|
|
|
| 16 |
A |
1565 |
1511 |
3.84 |
|
|
|
| 17 |
A |
1561 |
1506 |
3.49 |
|
|
|
| 18 |
A |
1557 |
1502 |
6.31 |
|
|
|
| 19 |
A |
1550 |
1496 |
9.29 |
|
|
|
| 20 |
A |
1541 |
1487 |
0.37 |
|
|
|
| 21 |
A |
1466 |
1415 |
3.33 |
|
|
|
| 22 |
A |
1462 |
1411 |
5.35 |
|
|
|
| 23 |
A |
1455 |
1404 |
6.84 |
|
|
|
| 24 |
A |
1417 |
1367 |
2.95 |
|
|
|
| 25 |
A |
1370 |
1322 |
1.04 |
|
|
|
| 26 |
A |
1335 |
1288 |
1.74 |
|
|
|
| 27 |
A |
1236 |
1192 |
1.30 |
|
|
|
| 28 |
A |
1157 |
1116 |
2.33 |
|
|
|
| 29 |
A |
1129 |
1090 |
5.81 |
|
|
|
| 30 |
A |
1102 |
1063 |
0.42 |
|
|
|
| 31 |
A |
1093 |
1055 |
4.80 |
|
|
|
| 32 |
A |
1080 |
1042 |
13.83 |
|
|
|
| 33 |
A |
1030 |
994 |
2.08 |
|
|
|
| 34 |
A |
988 |
953 |
1.20 |
|
|
|
| 35 |
A |
932 |
899 |
6.05 |
|
|
|
| 36 |
A |
854 |
824 |
15.44 |
|
|
|
| 37 |
A |
815 |
786 |
3.26 |
|
|
|
| 38 |
A |
738 |
712 |
1.05 |
|
|
|
| 39 |
A |
572 |
552 |
3.71 |
|
|
|
| 40 |
A |
529 |
510 |
2.97 |
|
|
|
| 41 |
A |
397 |
383 |
0.51 |
|
|
|
| 42 |
A |
309 |
298 |
0.39 |
|
|
|
| 43 |
A |
268 |
259 |
1.03 |
|
|
|
| 44 |
A |
243 |
235 |
1.62 |
|
|
|
| 45 |
A |
223 |
215 |
0.13 |
|
|
|
| 46 |
A |
179 |
173 |
1.14 |
|
|
|
| 47 |
A |
125 |
120 |
0.25 |
|
|
|
| 48 |
A |
60 |
57 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36624.5 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 35339.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.543 |
|
|
-0.279 |
| 2 |
H |
0.189 |
|
|
0.083 |
| 3 |
H |
0.184 |
|
|
0.066 |
| 4 |
H |
0.195 |
|
|
0.090 |
| 5 |
C |
-0.440 |
|
|
0.083 |
| 6 |
H |
0.192 |
|
|
0.036 |
| 7 |
H |
0.197 |
|
|
0.021 |
| 8 |
C |
-0.588 |
|
|
-0.438 |
| 9 |
H |
0.192 |
|
|
0.123 |
| 10 |
H |
0.195 |
|
|
0.138 |
| 11 |
H |
0.197 |
|
|
0.144 |
| 12 |
C |
-0.609 |
|
|
-0.392 |
| 13 |
H |
0.197 |
|
|
0.135 |
| 14 |
H |
0.198 |
|
|
0.116 |
| 15 |
H |
0.199 |
|
|
0.127 |
| 16 |
C |
0.046 |
|
|
-0.072 |
| 17 |
C |
-0.169 |
|
|
-0.116 |
| 18 |
H |
0.168 |
|
|
0.135 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.131 |
-0.129 |
-0.005 |
0.184 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.129 |
-0.130 |
-0.024 |
0.185 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.428 |
0.433 |
0.714 |
| y |
0.433 |
-39.014 |
-0.668 |
| z |
0.714 |
-0.668 |
-40.864 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.511 |
0.433 |
0.714 |
| y |
0.433 |
0.632 |
-0.668 |
| z |
0.714 |
-0.668 |
-2.142 |
|
| Polar |
|---|
| 3z2-r2 | -4.285 |
|---|
| x2-y2 | 0.586 |
|---|
| xy | 0.433 |
|---|
| xz | 0.714 |
|---|
| yz | -0.668 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.469 |
0.323 |
0.692 |
| y |
0.323 |
8.825 |
-0.241 |
| z |
0.692 |
-0.241 |
6.813 |
<r2> (average value of r
2) Å
2
| <r2> |
210.575 |
| (<r2>)1/2 |
14.511 |