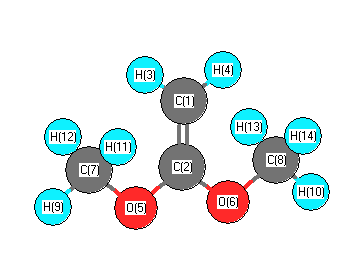
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3232 |
3118 |
0.36 |
|
|
|
| 2 |
A1 |
3162 |
3051 |
20.00 |
|
|
|
| 3 |
A1 |
3033 |
2926 |
5.00 |
|
|
|
| 4 |
A1 |
1693 |
1633 |
169.85 |
|
|
|
| 5 |
A1 |
1576 |
1521 |
6.95 |
|
|
|
| 6 |
A1 |
1529 |
1475 |
4.13 |
|
|
|
| 7 |
A1 |
1469 |
1417 |
20.43 |
|
|
|
| 8 |
A1 |
1209 |
1166 |
2.09 |
|
|
|
| 9 |
A1 |
1118 |
1078 |
1.23 |
|
|
|
| 10 |
A1 |
845 |
815 |
34.13 |
|
|
|
| 11 |
A1 |
488 |
471 |
0.04 |
|
|
|
| 12 |
A1 |
253 |
244 |
6.00 |
|
|
|
| 13 |
A2 |
3086 |
2977 |
0.00 |
|
|
|
| 14 |
A2 |
1552 |
1497 |
0.00 |
|
|
|
| 15 |
A2 |
1164 |
1123 |
0.00 |
|
|
|
| 16 |
A2 |
680 |
656 |
0.00 |
|
|
|
| 17 |
A2 |
233 |
225 |
0.00 |
|
|
|
| 18 |
A2 |
117 |
113 |
0.00 |
|
|
|
| 19 |
B1 |
3086 |
2978 |
76.48 |
|
|
|
| 20 |
B1 |
1551 |
1497 |
16.82 |
|
|
|
| 21 |
B1 |
1169 |
1128 |
0.00 |
|
|
|
| 22 |
B1 |
824 |
795 |
130.27 |
|
|
|
| 23 |
B1 |
685 |
661 |
1.46 |
|
|
|
| 24 |
B1 |
253 |
245 |
0.24 |
|
|
|
| 25 |
B1 |
88 |
85 |
12.68 |
|
|
|
| 26 |
B2 |
3307 |
3191 |
6.78 |
|
|
|
| 27 |
B2 |
3162 |
3051 |
12.43 |
|
|
|
| 28 |
B2 |
3033 |
2926 |
50.01 |
|
|
|
| 29 |
B2 |
1572 |
1517 |
21.26 |
|
|
|
| 30 |
B2 |
1515 |
1462 |
25.79 |
|
|
|
| 31 |
B2 |
1336 |
1290 |
387.59 |
|
|
|
| 32 |
B2 |
1196 |
1154 |
53.80 |
|
|
|
| 33 |
B2 |
1044 |
1007 |
128.84 |
|
|
|
| 34 |
B2 |
926 |
893 |
0.93 |
|
|
|
| 35 |
B2 |
556 |
537 |
0.33 |
|
|
|
| 36 |
B2 |
359 |
347 |
11.65 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26049.6 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 25135.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.502 |
|
|
-0.880 |
| 2 |
C |
0.598 |
|
|
0.735 |
| 3 |
H |
0.181 |
|
|
0.247 |
| 4 |
H |
0.181 |
|
|
0.247 |
| 5 |
O |
-0.520 |
|
|
-0.416 |
| 6 |
O |
-0.520 |
|
|
-0.416 |
| 7 |
C |
-0.327 |
|
|
0.047 |
| 8 |
C |
-0.327 |
|
|
0.047 |
| 9 |
H |
0.221 |
|
|
0.094 |
| 10 |
H |
0.221 |
|
|
0.094 |
| 11 |
H |
0.199 |
|
|
0.049 |
| 12 |
H |
0.199 |
|
|
0.050 |
| 13 |
H |
0.199 |
|
|
0.050 |
| 14 |
H |
0.199 |
|
|
0.049 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
1.566 |
1.566 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
0.006 |
0.000 |
1.590 |
1.590 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.224 |
0.000 |
0.000 |
| y |
0.000 |
-24.727 |
0.000 |
| z |
0.000 |
0.000 |
-39.463 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.129 |
0.000 |
0.000 |
| y |
0.000 |
14.116 |
0.000 |
| z |
0.000 |
0.000 |
-7.987 |
|
| Polar |
|---|
| 3z2-r2 | -15.974 |
|---|
| x2-y2 | -13.497 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.138 |
0.000 |
0.000 |
| y |
0.000 |
8.949 |
0.000 |
| z |
0.000 |
0.000 |
7.122 |
<r2> (average value of r
2) Å
2
| <r2> |
192.358 |
| (<r2>)1/2 |
13.869 |