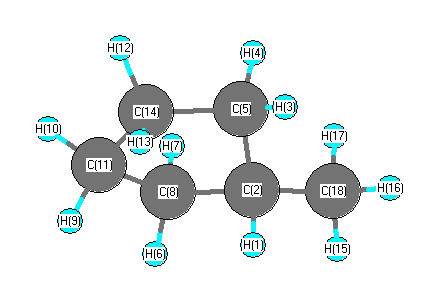
Vibrational Frequencies calculated at B3LYP/3-21G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3131 |
3021 |
74.88 |
|
|
|
| 2 |
A |
3111 |
3002 |
11.78 |
|
|
|
| 3 |
A |
3110 |
3001 |
20.78 |
|
|
|
| 4 |
A |
3102 |
2993 |
31.34 |
|
|
|
| 5 |
A |
3099 |
2990 |
20.07 |
|
|
|
| 6 |
A |
3097 |
2988 |
28.57 |
|
|
|
| 7 |
A |
3080 |
2972 |
34.34 |
|
|
|
| 8 |
A |
3068 |
2960 |
15.22 |
|
|
|
| 9 |
A |
3055 |
2948 |
47.68 |
|
|
|
| 10 |
A |
3047 |
2940 |
19.47 |
|
|
|
| 11 |
A |
3038 |
2931 |
19.32 |
|
|
|
| 12 |
A |
3031 |
2925 |
2.86 |
|
|
|
| 13 |
A |
1572 |
1517 |
3.70 |
|
|
|
| 14 |
A |
1567 |
1512 |
8.08 |
|
|
|
| 15 |
A |
1561 |
1507 |
3.82 |
|
|
|
| 16 |
A |
1550 |
1496 |
4.71 |
|
|
|
| 17 |
A |
1549 |
1494 |
2.61 |
|
|
|
| 18 |
A |
1540 |
1486 |
0.59 |
|
|
|
| 19 |
A |
1460 |
1409 |
5.00 |
|
|
|
| 20 |
A |
1402 |
1353 |
2.43 |
|
|
|
| 21 |
A |
1377 |
1329 |
0.38 |
|
|
|
| 22 |
A |
1364 |
1316 |
0.75 |
|
|
|
| 23 |
A |
1361 |
1313 |
0.65 |
|
|
|
| 24 |
A |
1352 |
1304 |
1.00 |
|
|
|
| 25 |
A |
1342 |
1295 |
0.00 |
|
|
|
| 26 |
A |
1297 |
1251 |
0.53 |
|
|
|
| 27 |
A |
1278 |
1233 |
0.28 |
|
|
|
| 28 |
A |
1233 |
1189 |
0.62 |
|
|
|
| 29 |
A |
1230 |
1187 |
0.43 |
|
|
|
| 30 |
A |
1164 |
1123 |
3.51 |
|
|
|
| 31 |
A |
1103 |
1064 |
0.30 |
|
|
|
| 32 |
A |
1069 |
1032 |
1.98 |
|
|
|
| 33 |
A |
1013 |
978 |
2.95 |
|
|
|
| 34 |
A |
1012 |
977 |
0.47 |
|
|
|
| 35 |
A |
998 |
963 |
0.57 |
|
|
|
| 36 |
A |
946 |
912 |
0.26 |
|
|
|
| 37 |
A |
912 |
880 |
0.56 |
|
|
|
| 38 |
A |
877 |
847 |
2.36 |
|
|
|
| 39 |
A |
852 |
822 |
0.96 |
|
|
|
| 40 |
A |
824 |
795 |
0.37 |
|
|
|
| 41 |
A |
763 |
737 |
1.25 |
|
|
|
| 42 |
A |
641 |
619 |
0.54 |
|
|
|
| 43 |
A |
536 |
517 |
0.77 |
|
|
|
| 44 |
A |
430 |
415 |
0.33 |
|
|
|
| 45 |
A |
308 |
297 |
0.14 |
|
|
|
| 46 |
A |
237 |
228 |
0.00 |
|
|
|
| 47 |
A |
178 |
172 |
0.02 |
|
|
|
| 48 |
A |
43 |
42 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37454.7 cm
-1
Scaled (by 0.9649) Zero Point Vibrational Energy (zpe) 36140.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYP/3-21G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
0.185 |
|
|
0.017 |
| 2 |
C |
-0.224 |
|
|
0.255 |
| 3 |
H |
0.181 |
|
|
0.045 |
| 4 |
H |
0.183 |
|
|
0.067 |
| 5 |
C |
-0.351 |
|
|
-0.155 |
| 6 |
H |
0.181 |
|
|
0.045 |
| 7 |
H |
0.183 |
|
|
0.067 |
| 8 |
C |
-0.351 |
|
|
-0.153 |
| 9 |
H |
0.189 |
|
|
0.055 |
| 10 |
H |
0.187 |
|
|
0.059 |
| 11 |
C |
-0.376 |
|
|
-0.129 |
| 12 |
H |
0.186 |
|
|
0.059 |
| 13 |
H |
0.189 |
|
|
0.055 |
| 14 |
C |
-0.376 |
|
|
-0.126 |
| 15 |
H |
0.184 |
|
|
0.151 |
| 16 |
H |
0.184 |
|
|
0.152 |
| 17 |
H |
0.181 |
|
|
0.152 |
| 18 |
C |
-0.537 |
|
|
-0.615 |
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.010 |
0.000 |
-0.051 |
0.052 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
-0.001 |
0.002 |
-0.029 |
0.029 |
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-40.387 |
0.000 |
-0.148 |
| y |
0.000 |
-40.095 |
-0.000 |
| z |
-0.148 |
-0.000 |
-39.287 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.696 |
0.000 |
-0.148 |
| y |
0.000 |
-0.258 |
-0.000 |
| z |
-0.148 |
-0.000 |
0.954 |
|
| Polar |
|---|
| 3z2-r2 | 1.908 |
|---|
| x2-y2 | -0.292 |
|---|
| xy | 0.000 |
|---|
| xz | -0.148 |
|---|
| yz | -0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.379 |
-0.000 |
-0.029 |
| y |
-0.000 |
8.548 |
0.000 |
| z |
-0.029 |
0.000 |
7.840 |
<r2> (average value of r
2) Å
2
| <r2> |
180.876 |
| (<r2>)1/2 |
13.449 |