return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5OCH3 (Anisole)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-340.758598
Energy at 298.15K-340.767195
HF Energy-340.302125
Nuclear repulsion energy339.265814
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3540 3147 1.94      
2 A' 3530 3138 3.12      
3 A' 3522 3131 3.65      
4 A' 3522 3130 1.24      
5 A' 3513 3122 0.47      
6 A' 3507 3117 0.34      
7 A' 3323 2954 10.72      
8 A' 1749 1555 43.85      
9 A' 1722 1531 9.02      
10 A' 1716 1525 11.61      
11 A' 1643 1461 23.48      
12 A' 1625 1445 77.80      
13 A' 1587 1411 3.66      
14 A' 1442 1282 3.00      
15 A' 1406 1250 121.23      
16 A' 1313 1167 2.73      
17 A' 1274 1133 1.06      
18 A' 1271 1129 5.35      
19 A' 1214 1079 6.29      
20 A' 1168 1038 0.96      
21 A' 1166 1037 1.37      
22 A' 1101 978 0.94      
23 A' 1064 946 0.87      
24 A' 828 736 5.93      
25 A' 655 583 0.00      
26 A' 581 517 2.26      
27 A' 461 410 0.04      
28 A' 276 246 1.18      
29 A" 3484 3097 7.93      
30 A" 1703 1514 1.78      
31 A" 1220 1085 2.29      
32 A" 1071 952 0.06      
33 A" 1052 935 0.01      
34 A" 965 858 2.14      
35 A" 910 809 0.04      
36 A" 798 710 27.60      
37 A" 751 668 2.61      
38 A" 528 469 0.07      
39 A" 442 393 0.01      
40 A" 297 264 0.67      
41 A" 221 196 0.63      
42 A" 91 81 2.88      

Unscaled Zero Point Vibrational Energy (zpe) 31626.2 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 28112.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.16073 0.05070 0.03883

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.543 0.000
C2 0.953 -0.513 0.000
C3 0.508 -1.860 0.000
C4 -0.868 -2.169 0.000
C5 -1.822 -1.117 0.000
C6 -1.397 0.225 0.000
O7 0.275 1.938 0.000
C8 1.709 2.224 0.000
H9 2.030 -0.304 0.000
H10 1.254 -2.669 0.000
H11 -1.201 -3.218 0.000
H12 -2.898 -1.347 0.000
H13 -2.123 1.051 0.000
H14 1.798 3.329 0.000
H15 2.224 1.827 0.902
H16 2.224 1.827 -0.902

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 O7 C8 H9 H10 H11 H12 H13 H14 H15 H16
C11.42282.45612.84842.46461.43301.42192.39672.20033.44843.94843.46032.18333.31542.72182.7218
C21.42281.41852.46202.83972.46312.54352.83941.09752.17683.45793.94053.45133.93372.81152.8115
C32.45611.41851.41052.44512.82353.80534.25672.17681.10102.18283.44423.92395.34674.16554.1655
C42.84842.46201.41051.42032.45174.26395.09353.44712.18071.10002.19003.45666.11075.13315.1331
C52.46462.83972.44511.42031.40683.70524.86043.93703.44562.19111.10082.18885.73275.08425.0842
C61.43302.46312.82352.45171.40682.39423.69373.46813.92453.44812.17341.10054.45474.06144.0614
O71.42192.54353.80534.26393.70522.39421.46212.84814.71025.36344.56762.55712.06232.15092.1509
C82.39672.83944.25675.09354.86043.69371.46212.54864.91396.17105.82894.00761.10861.11201.1120
H92.20031.09752.17683.44713.93703.46812.84812.54862.48874.35125.03784.36953.64062.32222.3222
H103.44842.17681.10102.18073.44563.92454.71024.91392.48872.51614.35785.02486.02244.68684.6868
H113.94843.45792.18281.10002.19113.44815.36346.17104.35122.51612.52584.36777.20106.16426.1642
H123.46033.94053.44422.19001.10082.17344.56765.82895.03784.35782.52582.52056.62716.09336.0933
H132.18333.45133.92393.45662.18881.10052.55714.00764.36955.02484.36772.52054.53474.50764.5076
H143.31543.93375.34676.11075.73274.45472.06231.10863.64066.02247.20106.62714.53471.80311.8031
H152.72182.81154.16555.13315.08424.06142.15091.11202.32224.68686.16426.09334.50761.80311.8033
H162.72182.81154.16555.13315.08424.06142.15091.11202.32224.68686.16426.09334.50761.80311.8033

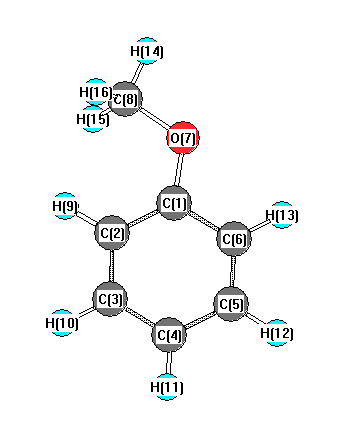
picture of Anisole state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 119.645 C1 C2 H9 121.090
C1 C6 C5 120.426 C1 C6 H13 118.450
C1 O7 C8 112.407 C2 C1 C6 119.202
C2 C1 O7 126.800 C2 C3 C4 120.982
C2 C3 H10 119.001 C3 C2 H9 119.265
C3 C4 C5 119.479 C3 C4 H11 120.289
C4 C3 H10 120.017 C4 C5 C6 120.266
C4 C5 H12 120.072 C5 C4 H11 120.232
C5 C6 H13 121.124 C6 C1 O7 113.998
C6 C5 H12 119.662 O7 C8 H14 105.871
O7 C8 H15 112.648 O7 C8 H16 112.648
H14 C8 H15 108.580 H14 C8 H16 108.580
H15 C8 H16 108.359
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/STO-3G Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.115      
2 C -0.093      
3 C -0.051      
4 C -0.071      
5 C -0.053      
6 C -0.076      
7 O -0.223      
8 C -0.055      
9 H 0.060      
10 H 0.061      
11 H 0.058      
12 H 0.062      
13 H 0.068      
14 H 0.072      
15 H 0.062      
16 H 0.062      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 281.025
(<r2>)1/2 16.764