return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C9H12 (n-propyl benzene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-344.149206
Energy at 298.15K-344.161414
HF Energy-343.626071
Nuclear repulsion energy404.357614
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3553 3158 3.21      
2 A' 3528 3136 1.12      
3 A' 3512 3122 0.84      
4 A' 3501 3112 0.25      
5 A' 3405 3026 7.49      
6 A' 3395 3018 2.88      
7 A' 3373 2998 3.94      
8 A' 1753 1559 2.54      
9 A' 1745 1551 0.34      
10 A' 1723 1532 0.21      
11 A' 1705 1515 1.26      
12 A' 1644 1461 9.35      
13 A' 1624 1443 1.66      
14 A' 1569 1395 1.03      
15 A' 1459 1297 19.45      
16 A' 1330 1183 3.03      
17 A' 1278 1136 0.02      
18 A' 1237 1099 1.82      
19 A' 1167 1037 0.01      
20 A' 1110 987 0.15      
21 A' 1076 957 0.04      
22 A' 1073 954 1.13      
23 A' 1012 900 0.33      
24 A' 991 881 0.89      
25 A' 872 775 0.74      
26 A' 807 717 9.02      
27 A' 763 678 7.49      
28 A' 625 556 1.97      
29 A' 528 469 1.92      
30 A' 325 289 0.03      
31 A' 296 263 0.21      
32 A' 107 95 0.09      
33 A" 3554 3159 4.40      
34 A" 3520 3129 4.55      
35 A" 3518 3127 2.24      
36 A" 3509 3119 0.05      
37 A" 3504 3115 0.12      
38 A" 1736 1543 1.65      
39 A" 1729 1537 0.19      
40 A" 1591 1414 7.37      
41 A" 1474 1310 0.19      
42 A" 1454 1292 0.09      
43 A" 1401 1245 0.06      
44 A" 1309 1164 0.01      
45 A" 1243 1105 0.05      
46 A" 1183 1052 0.45      
47 A" 1148 1021 0.56      
48 A" 1061 944 0.00      
49 A" 972 864 0.06      
50 A" 933 829 0.00      
51 A" 812 722 2.27      
52 A" 662 589 0.00      
53 A" 438 389 0.00      
54 A" 347 309 0.02      
55 A" 238 212 0.00      
56 A" 85 76 0.00      
57 A" 34 30 0.00      

Unscaled Zero Point Vibrational Energy (zpe) 44770.5 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 39796.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.14102 0.02752 0.02463

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.395 -0.764 1.226
C2 0.395 -2.184 1.228
C3 0.395 -2.898 0.000
C4 0.395 -2.184 -1.228
C5 0.395 -0.764 -1.226
C6 0.396 -0.034 0.000
C7 0.362 1.518 0.000
C8 -1.105 2.087 0.000
C9 -1.138 3.649 0.000
H10 0.400 -0.217 2.186
H11 0.398 -2.732 2.187
H12 0.396 -4.003 0.000
H13 0.398 -2.732 -2.187
H14 0.400 -0.217 -2.186
H15 0.898 1.904 0.893
H16 0.898 1.904 -0.893
H17 -1.646 1.704 -0.891
H18 -1.646 1.704 0.891
H19 -2.181 4.020 0.000
H20 -0.630 4.055 0.895
H21 -0.630 4.055 -0.895

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 C8 C9 H10 H11 H12 H13 H14 H15 H16 H17 H18 H19 H20 H21
C11.41952.46092.83492.45171.42702.59133.44714.83031.10512.18973.46253.93953.45522.73603.44413.83913.22045.57034.93835.3641
C21.41951.42072.45602.83492.47613.90074.69026.15482.18801.10472.19453.45893.93994.13264.63284.87564.40416.82876.33146.6694
C32.46091.42071.42072.46092.86454.41675.20576.72423.45952.19361.10442.19363.45954.91064.91065.11255.11257.38207.08507.0850
C42.83492.45601.42071.41952.47613.90074.69026.15483.93993.45892.19451.10472.18804.63284.13264.40414.87566.82876.66946.3314
C52.45172.83492.46091.41951.42702.59133.44714.83033.45523.93953.46252.18971.10513.44412.73603.22043.83915.57035.36414.9383
C61.42702.47612.86452.47611.42701.55252.59803.98962.19333.47313.96883.47312.19332.19202.19202.82532.82534.80344.30954.3095
C72.59133.90074.41673.90072.59131.55251.57352.60612.79084.78005.52114.78002.79081.11021.11022.20452.20453.56732.86712.8671
C83.44714.69025.20574.69023.44712.59801.57351.56243.51385.50106.27185.50103.51382.20052.20051.11001.11002.21232.21342.2134
C94.83036.15486.72426.15484.83033.98962.60611.56244.69956.91797.80396.91794.69952.82642.82642.19822.19821.10631.10621.1062
H101.10512.18803.45953.93993.45522.19332.79083.51384.69952.51514.37165.04464.37132.53353.77124.16383.09045.42084.57985.3662
H112.18971.10472.19363.45893.93953.47314.78005.50106.91792.51512.52974.37465.04464.83925.58815.77305.05327.55116.98487.5243
H123.46252.19451.10442.19453.46253.96885.52116.27187.80394.37162.52972.52974.37165.99505.99506.12626.12628.42638.17188.1718
H133.93953.45892.19361.10472.18973.47314.78005.50106.91795.04464.37462.52972.51515.58814.83925.05325.77307.55117.52436.9848
H143.45523.93993.45952.18801.10512.19332.79083.51384.69954.37135.04464.37162.51513.77122.53353.09044.16385.42085.36624.5798
H152.73604.13264.91064.63283.44412.19201.11022.20052.82642.53354.83925.99505.58813.77121.78513.11302.55173.84092.63853.1868
H163.44414.63284.91064.13262.73602.19201.11022.20052.82643.77125.58815.99504.83922.53351.78512.55173.11303.84093.18682.6385
H173.83914.87565.11254.40413.22042.82532.20451.11002.19824.16385.77306.12625.05323.09043.11302.55171.78132.53823.12162.5607
H183.22044.40415.11254.87563.83912.82532.20451.11002.19823.09045.05326.12625.77304.16382.55173.11301.78132.53822.56073.1216
H195.57036.82877.38206.82875.57034.80343.56732.21231.10635.42087.55118.42637.55115.42083.84093.84092.53822.53821.79041.7904
H204.93836.33147.08506.66945.36414.30952.86712.21341.10624.57986.98488.17187.52435.36622.63853.18683.12162.56071.79041.7892
H215.36416.66947.08506.33144.93834.30952.86712.21341.10625.36627.52438.17186.98484.57983.18682.63852.56073.12161.79041.7892

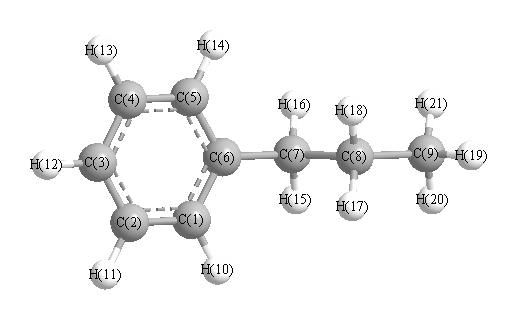
picture of n-propyl benzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 120.101 C1 C2 H11 119.814
C1 C6 C5 118.414 C1 C6 C7 120.784
C2 C1 C6 120.879 C2 C1 H10 119.627
C2 C3 C4 119.625 C2 C3 H12 120.187
C3 C2 H11 120.085 C3 C4 C5 120.101
C3 C4 H13 120.085 C4 C3 H12 120.187
C4 C5 C6 120.879 C4 C5 H14 119.627
C5 C4 H13 119.814 C5 C6 C7 120.784
C6 C1 H10 119.493 C6 C5 H14 119.493
C6 C7 C8 112.420 C6 C7 H15 109.697
C6 C7 H16 109.697 C7 C8 C9 112.416
C7 C8 H17 109.252 C7 C8 H18 109.252
C8 C7 H15 108.931 C8 C7 H16 108.931
C8 C9 H19 110.834 C8 C9 H20 110.923
C8 C9 H21 110.923 C9 C8 H17 109.520
C9 C8 H18 109.520 H15 C7 H16 107.011
H17 C8 H18 106.716 H19 C9 H20 108.044
H19 C9 H21 108.044 H20 C9 H21 107.950
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability