return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H4Cl2 (1,4-dichlorobenzene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes D2H 1AG
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-1136.273510
Energy at 298.15K-1136.277326
HF Energy-1135.889449
Nuclear repulsion energy438.410222
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 Ag 3534 3141 0.00      
2 Ag 1716 1525 0.00      
3 Ag 1256 1116 0.00      
4 Ag 1214 1079 0.00      
5 Ag 805 715 0.00      
6 Ag 340 302 0.00      
7 Au 1048 931 0.00      
8 Au 437 388 0.00      
9 B1g 903 802 0.00      
10 B1u 3521 3129 15.35      
11 B1u 1612 1433 79.16      
12 B1u 1219 1084 107.76      
13 B1u 1082 962 13.05      
14 B1u 589 523 9.54      
15 B2g 1024 911 0.00      
16 B2g 747 664 0.00      
17 B2g 309 275 0.00      
18 B2u 3531 3139 34.41      
19 B2u 1495 1329 5.51      
20 B2u 1237 1099 0.07      
21 B2u 1170 1040 0.03      
22 B2u 214 190 4.10      
23 B3g 3522 3130 0.00      
24 B3g 1687 1500 0.00      
25 B3g 1394 1239 0.00      
26 B3g 664 590 0.00      
27 B3g 343 305 0.00      
28 B3u 876 778 31.08      
29 B3u 507 451 1.20      
30 B3u 108 96 2.43      

Unscaled Zero Point Vibrational Energy (zpe) 19048.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 16932.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.18164 0.02114 0.01894

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is D2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.404
C2 0.000 0.000 -1.404
C3 0.000 1.238 0.708
C4 0.000 -1.238 0.708
C5 0.000 -1.238 -0.708
C6 0.000 1.238 -0.708
Cl7 0.000 0.000 3.208
Cl8 0.000 0.000 -3.208
H9 0.000 2.187 1.266
H10 0.000 -2.187 1.266
H11 0.000 -2.187 -1.266
H12 0.000 2.187 -1.266

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 Cl7 Cl8 H9 H10 H11 H12
C12.80871.42031.42032.44802.44801.80324.61192.19132.19133.45143.4514
C22.80872.44802.44801.42031.42034.61191.80323.45143.45142.19132.1913
C31.42032.44802.47542.85151.41542.78954.10631.10113.46983.95252.1898
C41.42032.44802.47541.41542.85152.78954.10633.46981.10112.18983.9525
C52.44801.42032.85151.41542.47544.10632.78953.95252.18981.10113.4698
C62.44801.42031.41542.85152.47544.10632.78952.18983.95253.46981.1011
Cl71.80324.61192.78952.78954.10634.10636.41512.92472.92474.97924.9792
Cl84.61191.80324.10634.10632.78952.78956.41514.97924.97922.92472.9247
H92.19133.45141.10113.46983.95252.18982.92474.97924.37395.05362.5314
H102.19133.45143.46981.10112.18983.95252.92474.97924.37392.53145.0536
H113.45142.19133.95252.18981.10113.46984.97922.92475.05362.53144.3739
H123.45142.19132.18983.95253.46981.10114.97922.92472.53145.05364.3739

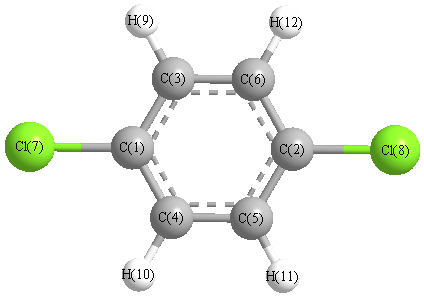
picture of 1,4-dichlorobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C3 C6 119.374 C1 C3 H9 120.180
C1 C4 C5 119.374 C1 C4 H10 120.180
C2 C5 C4 119.374 C2 C5 H11 120.180
C2 C6 C3 119.374 C2 C6 H12 120.180
C3 C1 C4 121.253 C3 C1 Cl7 119.374
C3 C6 H12 120.446 C4 C1 Cl7 119.374
C4 C5 H11 120.446 C5 C2 C6 121.253
C5 C2 Cl8 119.374 C5 C4 H10 120.446
C6 C2 Cl8 119.374 C6 C3 H9 120.446
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability