return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CH2CH2Br (n-propyl bromide)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-2661.145542
Energy at 298.15K-2661.155916
HF Energy-2660.972699
Nuclear repulsion energy233.915140
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3556 3161 2.23      
2 A' 3412 3033 3.41      
3 A' 3390 3013 2.53      
4 A' 3376 3001 2.89      
5 A' 1741 1548 0.56      
6 A' 1717 1526 0.03      
7 A' 1699 1510 1.67      
8 A' 1622 1442 0.97      
9 A' 1553 1381 1.15      
10 A' 1421 1263 39.39      
11 A' 1229 1092 0.15      
12 A' 1167 1037 0.24      
13 A' 1011 898 4.55      
14 A' 811 721 7.60      
15 A' 342 304 0.37      
16 A' 228 203 0.94      
17 A" 3556 3161 3.96      
18 A" 3525 3134 3.06      
19 A" 3505 3116 1.36      
20 A" 1734 1541 1.93      
21 A" 1463 1301 0.01      
22 A" 1397 1242 0.14      
23 A" 1199 1066 0.02      
24 A" 951 846 0.72      
25 A" 803 714 2.46      
26 A" 236 209 0.01      
27 A" 109 97 0.61      

Unscaled Zero Point Vibrational Energy (zpe) 23376.0 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 20778.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.82645 0.05325 0.05150

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.813 0.000
C2 1.554 0.625 0.000
C3 2.290 2.000 0.000
Br4 -0.959 -0.872 0.000
H5 -0.309 1.389 0.895
H6 -0.309 1.389 -0.895
H7 1.852 0.042 0.892
H8 1.852 0.042 -0.892
H9 3.383 1.850 0.000
H10 2.021 2.587 -0.894
H11 2.021 2.587 0.894

Atom - Atom Distances (Å)
  C1 C2 C3 Br4 H5 H6 H7 H8 H9 H10 H11
C11.56492.57931.93851.10821.10822.19572.19573.53842.83442.8344
C21.56491.55972.92472.20322.20321.10641.10642.20172.20692.2069
C32.57931.55974.33622.81572.81572.19522.19521.10331.10341.1034
Br41.93852.92474.33622.51692.51693.08813.08815.12474.65284.6528
H51.10822.20322.81572.51691.78942.54693.11103.82673.17272.6202
H61.10822.20322.81572.51691.78943.11102.54693.82672.62023.1727
H72.19571.10642.19523.08812.54693.11101.78362.53103.11402.5509
H82.19571.10642.19523.08813.11102.54691.78362.53102.55093.1140
H93.53842.20171.10335.12473.82673.82672.53102.53101.78841.7884
H102.83442.20691.10344.65283.17272.62023.11402.55091.78841.7886
H112.83442.20691.10344.65282.62023.17272.55093.11401.78841.7886

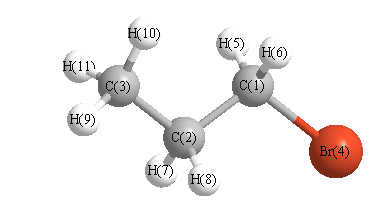
picture of n-propyl bromide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 111.271 C1 C2 H7 109.358
C1 C2 H8 109.358 C2 C1 Br4 112.758
C2 C1 H5 109.835 C2 C1 H6 109.835
C2 C3 H9 110.366 C2 C3 H10 110.763
C2 C3 H11 110.763 C3 C2 H7 109.676
C3 C2 H8 109.676 Br4 C1 H5 108.297
Br4 C1 H6 108.297 H5 C1 H6 107.671
H7 C2 H8 107.419 H9 C3 H10 108.286
H9 C3 H11 108.286 H10 C3 H11 108.290
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability