Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -155.679298 |
| Energy at 298.15K | -155.689796 |
| HF Energy | -155.463420 |
| Nuclear repulsion energy | 128.556097 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
|
Au |
119 |
106 |
0.00 |
|
|
|
|
Au |
219 |
195 |
0.00 |
|
|
|
|
Bg |
254 |
226 |
0.00 |
|
|
|
|
Bu |
274 |
244 |
0.12 |
|
|
|
|
Ag |
446 |
397 |
0.00 |
|
|
|
|
Au |
800 |
711 |
3.79 |
|
|
|
|
Bg |
901 |
801 |
0.00 |
|
|
|
|
Ag |
944 |
839 |
0.00 |
|
|
|
|
Au |
1078 |
958 |
0.95 |
|
|
|
|
Bu |
1093 |
971 |
3.35 |
|
|
|
|
Bu |
1149 |
1021 |
0.58 |
|
|
|
|
Ag |
1189 |
1057 |
0.00 |
|
|
|
|
Ag |
1298 |
1154 |
0.00 |
|
|
|
|
Bg |
1359 |
1208 |
0.00 |
|
|
|
|
Au |
1441 |
1281 |
0.01 |
|
|
|
|
Bg |
1478 |
1313 |
0.00 |
|
|
|
|
Bu |
1481 |
1317 |
16.45 |
|
|
|
|
Ag |
1591 |
1415 |
0.00 |
|
|
|
|
Bu |
1623 |
1443 |
1.00 |
|
|
|
|
Ag |
1630 |
1449 |
0.00 |
|
|
|
|
Ag |
1715 |
1524 |
0.00 |
|
|
|
|
Bu |
1728 |
1536 |
0.63 |
|
|
|
|
Au |
1737 |
1544 |
2.77 |
|
|
|
|
Bg |
1738 |
1545 |
0.00 |
|
|
|
|
Ag |
1746 |
1552 |
0.00 |
|
|
|
|
Bu |
1748 |
1553 |
0.41 |
|
|
|
|
Ag |
3374 |
2999 |
0.00 |
|
|
|
|
Bu |
3374 |
2999 |
7.55 |
|
|
|
|
Ag |
3402 |
3024 |
0.00 |
|
|
|
|
Bu |
3406 |
3028 |
11.62 |
|
|
|
|
Bg |
3510 |
3120 |
0.00 |
|
|
|
|
Au |
3521 |
3130 |
8.03 |
|
|
|
|
Ag |
3554 |
3159 |
0.00 |
|
|
|
|
Bu |
3554 |
3159 |
7.32 |
|
|
|
|
Bg |
3555 |
3160 |
0.00 |
|
|
|
|
Au |
3555 |
3160 |
10.62 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32792.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29149.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.435 |
0.650 |
0.000 |
| C2 |
0.435 |
-0.650 |
0.000 |
| C3 |
0.435 |
1.944 |
0.000 |
| C4 |
-0.435 |
-1.944 |
0.000 |
| H5 |
-1.092 |
0.649 |
0.890 |
| H6 |
-1.092 |
0.649 |
-0.890 |
| H7 |
1.092 |
-0.649 |
0.890 |
| H8 |
1.092 |
-0.649 |
-0.890 |
| H9 |
-0.204 |
2.844 |
0.000 |
| H10 |
1.081 |
1.980 |
0.893 |
| H11 |
1.081 |
1.980 |
-0.893 |
| H12 |
0.204 |
-2.844 |
0.000 |
| H13 |
-1.081 |
-1.980 |
0.893 |
| H14 |
-1.081 |
-1.980 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
| C1 | | 1.5635 | 1.5592 | 2.5936 | 1.1065 | 1.1065 | 2.1931 | 2.1931 | 2.2063 | 2.2058 | 2.2058 | 3.5514 | 2.8514 | 2.8514 |
C2 | 1.5635 | | 2.5936 | 1.5592 | 2.1931 | 2.1931 | 1.1065 | 1.1065 | 3.5514 | 2.8514 | 2.8514 | 2.2063 | 2.2058 | 2.2058 | C3 | 1.5592 | 2.5936 | | 3.9839 | 2.1910 | 2.1910 | 2.8191 | 2.8191 | 1.1034 | 1.1033 | 1.1033 | 4.7933 | 4.3003 | 4.3003 | C4 | 2.5936 | 1.5592 | 3.9839 | | 2.8191 | 2.8191 | 2.1910 | 2.1910 | 4.7933 | 4.3003 | 4.3003 | 1.1034 | 1.1033 | 1.1033 | H5 | 1.1065 | 2.1931 | 2.1910 | 2.8191 | | 1.7806 | 2.5403 | 3.1022 | 2.5297 | 2.5483 | 3.1105 | 3.8302 | 2.6288 | 3.1767 | H6 | 1.1065 | 2.1931 | 2.1910 | 2.8191 | 1.7806 | | 3.1022 | 2.5403 | 2.5297 | 3.1105 | 2.5483 | 3.8302 | 3.1767 | 2.6288 | H7 | 2.1931 | 1.1065 | 2.8191 | 2.1910 | 2.5403 | 3.1022 | | 1.7806 | 3.8302 | 2.6288 | 3.1767 | 2.5297 | 2.5483 | 3.1105 | H8 | 2.1931 | 1.1065 | 2.8191 | 2.1910 | 3.1022 | 2.5403 | 1.7806 | | 3.8302 | 3.1767 | 2.6288 | 2.5297 | 3.1105 | 2.5483 | H9 | 2.2063 | 3.5514 | 1.1034 | 4.7933 | 2.5297 | 2.5297 | 3.8302 | 3.8302 | | 1.7876 | 1.7876 | 5.7022 | 4.9836 | 4.9836 | H10 | 2.2058 | 2.8514 | 1.1033 | 4.3003 | 2.5483 | 3.1105 | 2.6288 | 3.1767 | 1.7876 | | 1.7867 | 4.9836 | 4.5117 | 4.8526 | H11 | 2.2058 | 2.8514 | 1.1033 | 4.3003 | 3.1105 | 2.5483 | 3.1767 | 2.6288 | 1.7876 | 1.7867 | | 4.9836 | 4.8526 | 4.5117 | H12 | 3.5514 | 2.2063 | 4.7933 | 1.1034 | 3.8302 | 3.8302 | 2.5297 | 2.5297 | 5.7022 | 4.9836 | 4.9836 | | 1.7876 | 1.7876 | H13 | 2.8514 | 2.2058 | 4.3003 | 1.1033 | 2.6288 | 3.1767 | 2.5483 | 3.1105 | 4.9836 | 4.5117 | 4.8526 | 1.7876 | | 1.7867 | H14 | 2.8514 | 2.2058 | 4.3003 | 1.1033 | 3.1767 | 2.6288 | 3.1105 | 2.5483 | 4.9836 | 4.8526 | 4.5117 | 1.7876 | 1.7867 | |
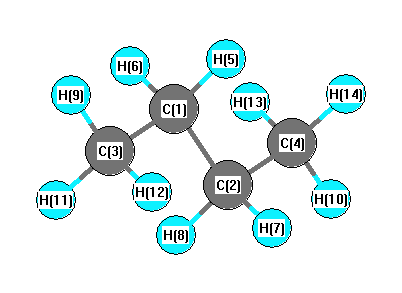 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.316 |
|
C1 |
C2 |
H7 |
109.249 |
| C1 |
C2 |
H8 |
109.249 |
|
C1 |
C3 |
H9 |
110.749 |
| C1 |
C3 |
H11 |
110.720 |
|
C1 |
C3 |
H12 |
31.132 |
| C2 |
C1 |
C3 |
112.316 |
|
C2 |
C1 |
H5 |
109.249 |
| C2 |
C1 |
H6 |
109.249 |
|
C2 |
C4 |
H10 |
17.448 |
| C2 |
C4 |
H13 |
110.720 |
|
C2 |
C4 |
H14 |
110.720 |
| C3 |
C1 |
H5 |
109.377 |
|
C3 |
C1 |
H6 |
109.377 |
| C4 |
C2 |
H7 |
109.377 |
|
C4 |
C2 |
H8 |
109.377 |
| H5 |
C1 |
H6 |
107.136 |
|
H7 |
C2 |
H8 |
107.136 |
| H9 |
C3 |
H11 |
108.210 |
|
H9 |
C3 |
H12 |
141.881 |
| H10 |
C4 |
H13 |
93.903 |
|
H10 |
C4 |
H14 |
113.859 |
| H11 |
C3 |
H12 |
93.484 |
|
H13 |
C4 |
H14 |
108.132 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -155.678149 |
| Energy at 298.15K | |
| Nuclear repulsion energy | 130.327572 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3561 |
3165 |
3.48 |
|
|
|
| 2 |
A |
3553 |
3158 |
0.05 |
|
|
|
| 3 |
A |
3510 |
3120 |
2.74 |
|
|
|
| 4 |
A |
3403 |
3025 |
6.45 |
|
|
|
| 5 |
A |
3377 |
3002 |
3.20 |
|
|
|
| 6 |
A |
1749 |
1555 |
0.03 |
|
|
|
| 7 |
A |
1743 |
1549 |
0.76 |
|
|
|
| 8 |
A |
1723 |
1532 |
0.27 |
|
|
|
| 9 |
A |
1628 |
1447 |
0.06 |
|
|
|
| 10 |
A |
1575 |
1400 |
1.45 |
|
|
|
| 11 |
A |
1458 |
1296 |
0.18 |
|
|
|
| 12 |
A |
1341 |
1192 |
0.08 |
|
|
|
| 13 |
A |
1213 |
1078 |
0.05 |
|
|
|
| 14 |
A |
1117 |
993 |
0.30 |
|
|
|
| 15 |
A |
934 |
830 |
0.07 |
|
|
|
| 16 |
A |
879 |
781 |
1.52 |
|
|
|
| 17 |
A |
330 |
294 |
0.07 |
|
|
|
| 18 |
A |
257 |
229 |
0.01 |
|
|
|
| 19 |
A |
110 |
97 |
0.00 |
|
|
|
| 20 |
B |
3558 |
3162 |
5.98 |
|
|
|
| 21 |
B |
3553 |
3158 |
7.52 |
|
|
|
| 22 |
B |
3516 |
3125 |
6.30 |
|
|
|
| 23 |
B |
3401 |
3023 |
5.78 |
|
|
|
| 24 |
B |
3375 |
3000 |
3.30 |
|
|
|
| 25 |
B |
1744 |
1551 |
0.93 |
|
|
|
| 26 |
B |
1737 |
1544 |
1.71 |
|
|
|
| 27 |
B |
1716 |
1525 |
0.05 |
|
|
|
| 28 |
B |
1629 |
1448 |
0.27 |
|
|
|
| 29 |
B |
1545 |
1373 |
17.47 |
|
|
|
| 30 |
B |
1437 |
1278 |
0.60 |
|
|
|
| 31 |
B |
1288 |
1145 |
2.33 |
|
|
|
| 32 |
B |
1092 |
971 |
0.70 |
|
|
|
| 33 |
B |
1080 |
960 |
1.83 |
|
|
|
| 34 |
B |
834 |
741 |
3.45 |
|
|
|
| 35 |
B |
459 |
408 |
0.33 |
|
|
|
| 36 |
B |
213 |
190 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 32817.3 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29171.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2h
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
