Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -153.258987 |
| Energy at 298.15K | -153.264042 |
| HF Energy | -153.011107 |
| Nuclear repulsion energy | 103.273040 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3614 |
3213 |
0.10 |
|
|
|
| 2 |
A |
3514 |
3124 |
0.06 |
|
|
|
| 3 |
A |
3461 |
3076 |
3.95 |
|
|
|
| 4 |
A |
1794 |
1594 |
0.00 |
|
|
|
| 5 |
A |
1614 |
1435 |
0.86 |
|
|
|
| 6 |
A |
1456 |
1294 |
0.05 |
|
|
|
| 7 |
A |
1133 |
1007 |
0.00 |
|
|
|
| 8 |
A |
1124 |
999 |
0.14 |
|
|
|
| 9 |
A |
984 |
874 |
0.00 |
|
|
|
| 10 |
A |
940 |
835 |
0.00 |
|
|
|
| 11 |
A |
805 |
716 |
0.00 |
|
|
|
| 12 |
A |
311 |
276 |
0.00 |
|
|
|
| 13 |
A |
73 |
65 |
0.00 |
|
|
|
| 14 |
B |
3613 |
3212 |
0.08 |
|
|
|
| 15 |
B |
3508 |
3118 |
4.07 |
|
|
|
| 16 |
B |
3457 |
3073 |
13.29 |
|
|
|
| 17 |
B |
1804 |
1604 |
0.17 |
|
|
|
| 18 |
B |
1581 |
1405 |
0.81 |
|
|
|
| 19 |
B |
1413 |
1256 |
4.79 |
|
|
|
| 20 |
B |
1195 |
1062 |
2.42 |
|
|
|
| 21 |
B |
1134 |
1008 |
8.63 |
|
|
|
| 22 |
B |
989 |
879 |
22.85 |
|
|
|
| 23 |
B |
592 |
526 |
0.65 |
|
|
|
| 24 |
B |
533 |
474 |
3.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20320.3 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 18062.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.000 |
0.755 |
0.582 |
| C2 |
0.000 |
-0.755 |
0.582 |
| C3 |
0.000 |
1.544 |
-0.517 |
| C4 |
-0.000 |
-1.544 |
-0.517 |
| H5 |
0.000 |
1.229 |
1.577 |
| H6 |
-0.000 |
-1.229 |
1.577 |
| H7 |
0.000 |
2.639 |
-0.432 |
| H8 |
-0.000 |
1.129 |
-1.534 |
| H9 |
-0.000 |
-2.639 |
-0.432 |
| H10 |
0.000 |
-1.129 |
-1.534 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.5095 | 1.3534 | 2.5481 | 1.1018 | 2.2192 | 2.1401 | 2.1495 | 3.5422 | 2.8336 |
C2 | 1.5095 | | 2.5481 | 1.3534 | 2.2192 | 1.1018 | 3.5422 | 2.8336 | 2.1401 | 2.1495 | C3 | 1.3534 | 2.5481 | | 3.0881 | 2.1172 | 3.4748 | 1.0983 | 1.0986 | 4.1840 | 2.8604 | C4 | 2.5481 | 1.3534 | 3.0881 | | 3.4748 | 2.1172 | 4.1840 | 2.8604 | 1.0983 | 1.0986 | H5 | 1.1018 | 2.2192 | 2.1172 | 3.4748 | | 2.4584 | 2.4542 | 3.1125 | 4.3588 | 3.9039 | H6 | 2.2192 | 1.1018 | 3.4748 | 2.1172 | 2.4584 | | 4.3588 | 3.9039 | 2.4542 | 3.1125 | H7 | 2.1401 | 3.5422 | 1.0983 | 4.1840 | 2.4542 | 4.3588 | | 1.8692 | 5.2781 | 3.9262 | H8 | 2.1495 | 2.8336 | 1.0986 | 2.8604 | 3.1125 | 3.9039 | 1.8692 | | 3.9262 | 2.2586 | H9 | 3.5422 | 2.1401 | 4.1840 | 1.0983 | 4.3588 | 2.4542 | 5.2781 | 3.9262 | | 1.8692 | H10 | 2.8336 | 2.1495 | 2.8604 | 1.0986 | 3.9039 | 3.1125 | 3.9262 | 2.2586 | 1.8692 | |
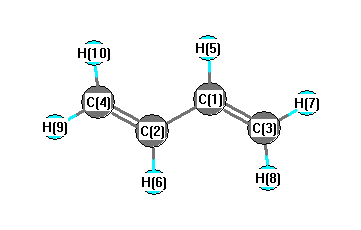 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
125.680 |
|
C1 |
C2 |
H6 |
115.509 |
| C1 |
C3 |
H7 |
121.253 |
|
C1 |
C3 |
H8 |
122.139 |
| C2 |
C1 |
C3 |
125.680 |
|
C2 |
C1 |
H5 |
115.509 |
| C2 |
C4 |
H9 |
121.253 |
|
C2 |
C4 |
H10 |
122.139 |
| C3 |
C1 |
H5 |
118.811 |
|
C4 |
C2 |
H6 |
118.811 |
| H7 |
C3 |
H8 |
116.609 |
|
H9 |
C4 |
H10 |
116.609 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
