Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -3076.519723 |
| Energy at 298.15K | |
| HF Energy | -3076.390870 |
| Nuclear repulsion energy | 284.396416 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3410 |
3031 |
0.71 |
17.08 |
0.15 |
0.26 |
| 2 |
A' |
3402 |
3024 |
0.30 |
45.35 |
0.14 |
0.25 |
| 3 |
A' |
1690 |
1502 |
1.15 |
6.35 |
0.75 |
0.86 |
| 4 |
A' |
1671 |
1485 |
0.13 |
17.52 |
0.75 |
0.86 |
| 5 |
A' |
1488 |
1323 |
0.39 |
11.05 |
0.27 |
0.42 |
| 6 |
A' |
1373 |
1221 |
45.04 |
0.02 |
0.72 |
0.84 |
| 7 |
A' |
1138 |
1011 |
0.77 |
10.44 |
0.75 |
0.86 |
| 8 |
A' |
898 |
798 |
15.38 |
24.36 |
0.32 |
0.48 |
| 9 |
A' |
810 |
720 |
18.93 |
5.50 |
0.37 |
0.55 |
| 10 |
A' |
266 |
237 |
0.69 |
4.34 |
0.40 |
0.58 |
| 11 |
A' |
202 |
180 |
6.84 |
0.04 |
0.47 |
0.64 |
| 12 |
A" |
3534 |
3141 |
0.12 |
8.84 |
0.75 |
0.86 |
| 13 |
A" |
3518 |
3127 |
0.15 |
41.12 |
0.75 |
0.86 |
| 14 |
A" |
1425 |
1267 |
0.01 |
18.37 |
0.75 |
0.86 |
| 15 |
A" |
1257 |
1117 |
1.10 |
0.10 |
0.75 |
0.86 |
| 16 |
A" |
1068 |
949 |
0.06 |
8.70 |
0.75 |
0.86 |
| 17 |
A" |
814 |
724 |
0.98 |
0.02 |
0.75 |
0.86 |
| 18 |
A" |
99 |
88 |
5.87 |
0.00 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 14031.1 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 12472.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.628 |
0.000 |
| C2 |
1.245 |
-0.320 |
0.000 |
| Br3 |
-1.645 |
-0.392 |
0.000 |
| Cl4 |
2.797 |
0.661 |
0.000 |
| H5 |
0.025 |
1.277 |
0.897 |
| H6 |
0.025 |
1.277 |
-0.897 |
| H7 |
1.258 |
-0.963 |
0.903 |
| H8 |
1.258 |
-0.963 |
-0.903 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5643 | 1.9354 | 2.7970 | 1.1079 | 1.1079 | 2.2202 | 2.2202 |
C2 | 1.5643 | | 2.8906 | 1.8358 | 2.2004 | 2.2004 | 1.1088 | 1.1088 | Br3 | 1.9354 | 2.8906 | | 4.5649 | 2.5261 | 2.5261 | 3.0937 | 3.0937 | Cl4 | 2.7970 | 1.8358 | 4.5649 | | 2.9775 | 2.9775 | 2.4122 | 2.4122 | H5 | 1.1079 | 2.2004 | 2.5261 | 2.9775 | | 1.7946 | 2.5568 | 3.1271 | H6 | 1.1079 | 2.2004 | 2.5261 | 2.9775 | 1.7946 | | 3.1271 | 2.5568 | H7 | 2.2202 | 1.1088 | 3.0937 | 2.4122 | 2.5568 | 3.1271 | | 1.8063 | H8 | 2.2202 | 1.1088 | 3.0937 | 2.4122 | 3.1271 | 2.5568 | 1.8063 | |
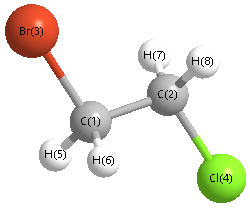 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
110.439 |
|
C1 |
C2 |
H7 |
111.164 |
| C1 |
C2 |
H8 |
111.164 |
|
C2 |
C1 |
Br3 |
110.927 |
| C2 |
C1 |
Cl4 |
37.955 |
|
C2 |
C1 |
H6 |
109.680 |
| Br3 |
C1 |
H5 |
109.163 |
|
Br3 |
C1 |
H6 |
109.163 |
| Cl4 |
C2 |
H7 |
107.422 |
|
Cl4 |
C2 |
H8 |
107.422 |
| H5 |
C1 |
H6 |
108.177 |
|
H7 |
C2 |
H8 |
109.083 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -3076.518134 |
| Energy at 298.15K | |
| HF Energy | -3076.388815 |
| Nuclear repulsion energy | 301.168916 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3525 |
3133 |
0.11 |
27.67 |
0.74 |
0.85 |
| 2 |
A |
3508 |
3119 |
0.16 |
37.05 |
0.74 |
0.85 |
| 3 |
A |
3402 |
3024 |
0.20 |
46.32 |
0.12 |
0.21 |
| 4 |
A |
3391 |
3015 |
0.55 |
36.51 |
0.18 |
0.30 |
| 5 |
A |
1678 |
1491 |
1.24 |
10.47 |
0.75 |
0.86 |
| 6 |
A |
1661 |
1476 |
1.06 |
17.47 |
0.75 |
0.86 |
| 7 |
A |
1484 |
1319 |
13.88 |
5.78 |
0.50 |
0.67 |
| 8 |
A |
1462 |
1299 |
53.20 |
0.72 |
0.70 |
0.83 |
| 9 |
A |
1337 |
1188 |
0.21 |
16.88 |
0.75 |
0.86 |
| 10 |
A |
1269 |
1128 |
1.08 |
6.84 |
0.75 |
0.86 |
| 11 |
A |
1136 |
1009 |
1.83 |
4.98 |
0.75 |
0.86 |
| 12 |
A |
1041 |
925 |
3.67 |
5.80 |
0.51 |
0.68 |
| 13 |
A |
967 |
859 |
6.57 |
2.56 |
0.63 |
0.78 |
| 14 |
A |
795 |
707 |
8.30 |
11.04 |
0.46 |
0.63 |
| 15 |
A |
707 |
629 |
4.29 |
12.38 |
0.20 |
0.34 |
| 16 |
A |
406 |
361 |
5.73 |
2.54 |
0.75 |
0.86 |
| 17 |
A |
235 |
209 |
0.76 |
0.95 |
0.48 |
0.65 |
| 18 |
A |
90 |
80 |
1.10 |
1.32 |
0.74 |
0.85 |
Unscaled Zero Point Vibrational Energy (zpe) 14045.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 12485.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.042 |
1.116 |
-0.382 |
| C2 |
1.305 |
0.940 |
0.396 |
| Br3 |
-1.360 |
-0.234 |
0.038 |
| Cl4 |
2.190 |
-0.591 |
-0.087 |
| H5 |
-0.455 |
2.112 |
-0.122 |
| H6 |
0.156 |
1.101 |
-1.472 |
| H7 |
1.979 |
1.793 |
0.172 |
| H8 |
1.133 |
0.892 |
1.491 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5655 | 1.9326 | 2.8252 | 1.1092 | 1.1081 | 2.2028 | 2.2221 |
C2 | 1.5655 | | 2.9341 | 1.8334 | 2.1762 | 2.1986 | 1.1096 | 1.1090 | Br3 | 1.9326 | 2.9341 | | 3.5700 | 2.5198 | 2.5227 | 3.9086 | 3.0971 | Cl4 | 2.8252 | 1.8334 | 3.5700 | | 3.7814 | 2.9858 | 2.4067 | 2.4094 | H5 | 1.1092 | 2.1762 | 2.5198 | 3.7814 | | 1.7941 | 2.4723 | 2.5705 | H6 | 1.1081 | 2.1986 | 2.5227 | 2.9858 | 1.7941 | | 2.5503 | 3.1263 | H7 | 2.2028 | 1.1096 | 3.9086 | 2.4067 | 2.4723 | 2.5503 | | 1.8070 | H8 | 2.2221 | 1.1090 | 3.0971 | 2.4094 | 2.5705 | 3.1263 | 1.8070 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.204 |
|
C1 |
C2 |
H7 |
109.686 |
| C1 |
C2 |
H8 |
111.226 |
|
C2 |
C1 |
Br3 |
113.608 |
| C2 |
C1 |
Cl4 |
36.930 |
|
C2 |
C1 |
H6 |
109.452 |
| Br3 |
C1 |
H5 |
108.833 |
|
Br3 |
C1 |
H6 |
109.098 |
| Cl4 |
C2 |
H7 |
107.142 |
|
Cl4 |
C2 |
H8 |
107.367 |
| H5 |
C1 |
H6 |
108.021 |
|
H7 |
C2 |
H8 |
109.077 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
