return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C3H6O (2-Propen-1-ol)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C1 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-189.695462
Energy at 298.15K-189.701550
HF Energy-189.479545
Nuclear repulsion energy113.993447
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3863 3434 47.99      
2 A 3614 3212 0.31      
3 A 3515 3125 1.93      
4 A 3458 3074 6.13      
5 A 3424 3044 15.44      
6 A 3316 2947 11.42      
7 A 1824 1621 0.59      
8 A 1725 1533 0.95      
9 A 1666 1481 8.56      
10 A 1590 1413 3.54      
11 A 1430 1271 12.77      
12 A 1412 1255 23.70      
13 A 1352 1201 5.25      
14 A 1233 1096 3.87      
15 A 1207 1073 2.97      
16 A 1130 1004 5.55      
17 A 1042 926 1.91      
18 A 1001 889 11.72      
19 A 985 876 2.65      
20 A 682 607 1.19      
21 A 446 397 3.08      
22 A 343 305 33.60      
23 A 329 293 34.69      
24 A 87 78 0.61      

Unscaled Zero Point Vibrational Energy (zpe) 20336.6 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 18077.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.86389 0.13794 0.13290

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C1

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.608 0.454 0.308
C2 -0.699 -0.368 0.276
C3 -1.858 0.088 -0.238
O4 1.658 -0.330 -0.349
H5 0.441 1.434 -0.200
H6 0.879 0.677 1.370
H7 -0.642 -1.379 0.709
H8 -2.771 -0.524 -0.234
H9 -1.939 1.092 -0.677
H10 2.457 0.293 -0.253

Atom - Atom Distances (Å)
  C1 C2 C3 O4 H5 H6 H7 H8 H9 H10
C11.54412.55271.46631.11611.11812.25473.55902.80461.9383
C21.54411.34782.43862.18392.18571.10162.13962.13943.2666
C32.55271.34783.54342.66433.22882.12801.09821.09884.3199
O41.46632.43863.54342.14892.13912.73994.43483.88251.0174
H51.11612.18392.66432.14891.79703.14793.76102.45112.3168
H61.11812.18573.22882.13911.79702.64154.16343.50762.2953
H72.25471.10162.12802.73993.14792.64152.48063.11603.6495
H83.55902.13961.09824.43483.76104.16342.48061.87025.2907
H92.80462.13941.09883.88252.45113.50763.11601.87024.4878
H101.93833.26664.31991.01742.31682.29533.64955.29074.4878

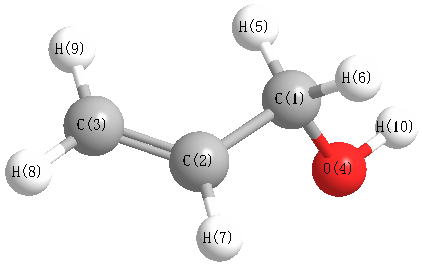
picture of 2-Propen-1-ol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 123.801 C1 C2 H7 115.895
C1 O4 H10 101.045 C2 C1 O4 108.172
C2 C1 H5 109.305 C2 C1 H6 109.329
C2 C3 H8 121.686 C2 C3 H9 121.624
C3 C2 H7 120.303 O4 C1 H5 111.918
O4 C1 H6 111.005 H5 C1 H6 107.080
H8 C3 H9 116.690
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability