Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -205.460345 |
| Energy at 298.15K | -205.466071 |
| HF Energy | -205.231394 |
| Nuclear repulsion energy | 116.357534 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3853 |
3425 |
15.50 |
|
|
|
| 2 |
A' |
3565 |
3169 |
0.01 |
|
|
|
| 3 |
A' |
3493 |
3105 |
4.83 |
|
|
|
| 4 |
A' |
3371 |
2996 |
0.94 |
|
|
|
| 5 |
A' |
1729 |
1537 |
0.11 |
|
|
|
| 6 |
A' |
1695 |
1507 |
3.99 |
|
|
|
| 7 |
A' |
1591 |
1414 |
3.12 |
|
|
|
| 8 |
A' |
1531 |
1361 |
16.17 |
|
|
|
| 9 |
A' |
1434 |
1275 |
27.68 |
|
|
|
| 10 |
A' |
1238 |
1101 |
4.83 |
|
|
|
| 11 |
A' |
1094 |
972 |
32.45 |
|
|
|
| 12 |
A' |
983 |
874 |
1.04 |
|
|
|
| 13 |
A' |
673 |
599 |
4.76 |
|
|
|
| 14 |
A' |
294 |
261 |
0.54 |
|
|
|
| 15 |
A" |
3537 |
3144 |
0.94 |
|
|
|
| 16 |
A" |
1715 |
1524 |
3.41 |
|
|
|
| 17 |
A" |
1165 |
1035 |
0.00 |
|
|
|
| 18 |
A" |
875 |
777 |
3.35 |
|
|
|
| 19 |
A" |
480 |
427 |
17.30 |
|
|
|
| 20 |
A" |
376 |
334 |
47.11 |
|
|
|
| 21 |
A" |
47 |
42 |
0.18 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17368.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15438.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.482 |
0.484 |
0.000 |
| C2 |
0.000 |
0.896 |
0.000 |
| N3 |
1.077 |
0.094 |
0.000 |
| O4 |
0.640 |
-1.292 |
0.000 |
| H5 |
1.559 |
-1.736 |
0.000 |
| H6 |
-1.575 |
-0.615 |
0.000 |
| H7 |
-1.993 |
0.885 |
0.894 |
| H8 |
-1.993 |
0.885 |
-0.894 |
| H9 |
0.243 |
1.974 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5388 | 2.5887 | 2.7673 | 3.7651 | 1.1024 | 1.1048 | 1.1048 | 2.2795 |
C2 | 1.5388 | | 1.3427 | 2.2803 | 3.0591 | 2.1823 | 2.1843 | 2.1843 | 1.1044 | N3 | 2.5887 | 1.3427 | | 1.4539 | 1.8925 | 2.7445 | 3.2937 | 3.2937 | 2.0563 | O4 | 2.7673 | 2.2803 | 1.4539 | | 1.0206 | 2.3155 | 3.5316 | 3.5316 | 3.2902 | H5 | 3.7651 | 3.0591 | 1.8925 | 1.0206 | | 3.3279 | 4.5039 | 4.5039 | 3.9362 | H6 | 1.1024 | 2.1823 | 2.7445 | 2.3155 | 3.3279 | | 1.7956 | 1.7956 | 3.1627 | H7 | 1.1048 | 2.1843 | 3.2937 | 3.5316 | 4.5039 | 1.7956 | | 1.7873 | 2.6423 | H8 | 1.1048 | 2.1843 | 3.2937 | 3.5316 | 4.5039 | 1.7956 | 1.7873 | | 2.6423 | H9 | 2.2795 | 1.1044 | 2.0563 | 3.2902 | 3.9362 | 3.1627 | 2.6423 | 2.6423 | |
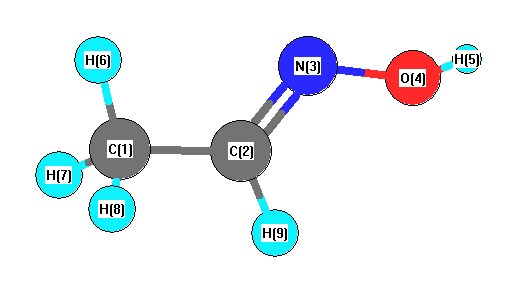 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
127.766 |
|
C1 |
C2 |
H9 |
118.242 |
| C2 |
C1 |
H6 |
110.341 |
|
C2 |
C1 |
H7 |
110.357 |
| C2 |
C1 |
H8 |
110.357 |
|
C2 |
N3 |
O4 |
109.183 |
| N3 |
C2 |
H9 |
113.992 |
|
N3 |
O4 |
H5 |
98.251 |
| H6 |
C1 |
H7 |
108.878 |
|
H6 |
C1 |
H8 |
108.878 |
| H7 |
C1 |
H8 |
107.973 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
