return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H5Br (bromobenzene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-2772.353488
Energy at 298.15K-2772.361264
Nuclear repulsion energy424.995483
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3531 3139 1.39      
2 A1 3517 3127 4.42      
3 A1 3507 3117 1.04      
4 A1 1729 1537 16.71      
5 A1 1620 1440 29.19      
6 A1 1274 1132 0.12      
7 A1 1202 1069 38.45      
8 A1 1102 979 1.44      
9 A1 1067 948 2.06      
10 A1 732 651 7.51      
11 A1 349 310 0.37      
12 A2 1058 941 0.00      
13 A2 924 821 0.00      
14 A2 436 388 0.00      
15 B1 1075 955 0.04      
16 B1 990 880 1.06      
17 B1 796 707 19.72      
18 B1 754 670 4.17      
19 B1 494 439 0.64      
20 B1 184 164 0.00      
21 B2 3522 3130 7.68      
22 B2 3510 3120 0.45      
23 B2 1710 1520 2.32      
24 B2 1570 1395 5.48      
25 B2 1429 1270 0.25      
26 B2 1285 1142 0.00      
27 B2 1205 1071 0.22      
28 B2 1157 1028 0.05      
29 B2 656 583 0.02      
30 B2 258 229 0.36      

Unscaled Zero Point Vibrational Energy (zpe) 21320.2 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 18951.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.18421 0.03209 0.02733

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Br1 0.000 0.000 1.828
C2 0.000 0.000 -0.077
C3 0.000 1.230 -0.792
C4 0.000 -1.230 -0.792
C5 0.000 1.224 -2.208
C6 0.000 -1.224 -2.208
C7 0.000 0.000 -2.918
H8 0.000 2.185 -0.243
H9 0.000 -2.185 -0.243
H10 0.000 2.181 -2.754
H11 0.000 -2.181 -2.754
H12 0.000 0.000 -4.019

Atom - Atom Distances (Å)
  Br1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12
Br11.90482.89492.89494.21734.21734.74653.01063.01065.07445.07445.8471
C21.90481.42331.42332.45762.45762.84172.19112.19113.45293.45293.9423
C32.89491.42332.46051.41522.83342.45631.10123.45892.17963.93463.4532
C42.89491.42332.46052.83341.41522.45633.45891.10123.93462.17963.4532
C54.21732.45761.41522.83342.44901.41592.18643.93461.10113.44862.1865
C64.21732.45762.83341.41522.44901.41593.93462.18643.44861.10112.1865
C74.74652.84172.45632.45631.41591.41593.45393.45392.18672.18671.1006
H83.01062.19111.10123.45892.18643.93463.45394.36952.51075.03584.3623
H93.01062.19113.45891.10123.93462.18643.45394.36955.03582.51074.3623
H105.07443.45292.17963.93461.10113.44862.18672.51075.03584.36102.5209
H115.07443.45293.93462.17963.44861.10112.18675.03582.51074.36102.5209
H125.84713.94233.45323.45322.18652.18651.10064.36234.36232.52092.5209

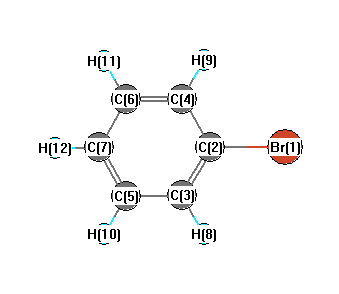
picture of bromobenzene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Br1 C2 C3 120.190 Br1 C2 C4 120.190
C2 C3 C5 119.957 C2 C3 H8 119.898
C2 C4 C6 119.957 C2 C4 H9 119.898
C3 C2 C4 119.620 C3 C5 C7 120.368
C3 C5 H10 119.518 C4 C6 C7 120.368
C4 C6 H11 119.518 C5 C3 H8 120.144
C5 C7 C6 119.730 C5 C7 H12 120.135
C6 C4 H9 120.144 C6 C7 H12 120.135
C7 C5 H10 120.115 C7 C6 H11 120.115
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability