Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -193.099926 |
| Energy at 298.15K | -193.109898 |
| HF Energy | -192.814962 |
| Nuclear repulsion energy | 167.205767 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3613 |
3211 |
0.59 |
|
|
|
| 2 |
A |
3555 |
3160 |
4.86 |
|
|
|
| 3 |
A |
3554 |
3159 |
3.36 |
|
|
|
| 4 |
A |
3520 |
3129 |
6.29 |
|
|
|
| 5 |
A |
3508 |
3118 |
1.42 |
|
|
|
| 6 |
A |
3501 |
3112 |
2.15 |
|
|
|
| 7 |
A |
3456 |
3072 |
5.87 |
|
|
|
| 8 |
A |
3405 |
3027 |
6.76 |
|
|
|
| 9 |
A |
3392 |
3015 |
2.89 |
|
|
|
| 10 |
A |
3373 |
2999 |
3.71 |
|
|
|
| 11 |
A |
1826 |
1623 |
0.16 |
|
|
|
| 12 |
A |
1745 |
1551 |
0.31 |
|
|
|
| 13 |
A |
1737 |
1544 |
1.48 |
|
|
|
| 14 |
A |
1725 |
1533 |
0.12 |
|
|
|
| 15 |
A |
1707 |
1517 |
0.70 |
|
|
|
| 16 |
A |
1625 |
1445 |
0.26 |
|
|
|
| 17 |
A |
1614 |
1435 |
1.97 |
|
|
|
| 18 |
A |
1568 |
1393 |
0.80 |
|
|
|
| 19 |
A |
1471 |
1307 |
0.12 |
|
|
|
| 20 |
A |
1456 |
1294 |
9.95 |
|
|
|
| 21 |
A |
1424 |
1266 |
9.80 |
|
|
|
| 22 |
A |
1412 |
1255 |
0.13 |
|
|
|
| 23 |
A |
1316 |
1170 |
0.68 |
|
|
|
| 24 |
A |
1240 |
1103 |
0.38 |
|
|
|
| 25 |
A |
1175 |
1045 |
0.19 |
|
|
|
| 26 |
A |
1142 |
1015 |
2.41 |
|
|
|
| 27 |
A |
1132 |
1006 |
1.54 |
|
|
|
| 28 |
A |
1044 |
928 |
0.90 |
|
|
|
| 29 |
A |
995 |
885 |
11.77 |
|
|
|
| 30 |
A |
986 |
876 |
1.87 |
|
|
|
| 31 |
A |
967 |
860 |
0.09 |
|
|
|
| 32 |
A |
816 |
725 |
2.72 |
|
|
|
| 33 |
A |
677 |
602 |
2.91 |
|
|
|
| 34 |
A |
450 |
400 |
0.50 |
|
|
|
| 35 |
A |
397 |
353 |
0.04 |
|
|
|
| 36 |
A |
247 |
219 |
0.04 |
|
|
|
| 37 |
A |
238 |
212 |
0.03 |
|
|
|
| 38 |
A |
102 |
91 |
0.01 |
|
|
|
| 39 |
A |
86 |
77 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33597.2 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29864.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
2.501 |
-0.236 |
-0.318 |
| C2 |
1.447 |
0.360 |
0.274 |
| C3 |
0.071 |
-0.305 |
0.470 |
| C4 |
-1.064 |
0.454 |
-0.299 |
| C5 |
-2.460 |
-0.205 |
-0.084 |
| H6 |
3.463 |
0.281 |
-0.441 |
| H7 |
2.443 |
-1.265 |
-0.697 |
| H8 |
1.542 |
1.395 |
0.641 |
| H9 |
0.116 |
-1.354 |
0.120 |
| H10 |
-0.180 |
-0.324 |
1.549 |
| H11 |
-1.097 |
1.506 |
0.044 |
| H12 |
-0.827 |
0.469 |
-1.380 |
| H13 |
-3.241 |
0.347 |
-0.634 |
| H14 |
-2.459 |
-1.249 |
-0.442 |
| H15 |
-2.729 |
-0.210 |
0.986 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3481 | 2.5554 | 3.6318 | 4.9675 | 1.0981 | 1.0985 | 2.1215 | 2.6707 | 3.2686 | 4.0137 | 3.5640 | 5.7808 | 5.0641 | 5.3908 |
C2 | 1.3481 | | 1.5406 | 2.5773 | 3.9642 | 2.1403 | 2.1394 | 1.1023 | 2.1759 | 2.1777 | 2.7989 | 2.8136 | 4.7751 | 4.2844 | 4.2745 | C3 | 2.5554 | 1.5406 | | 1.5678 | 2.5936 | 3.5600 | 2.8121 | 2.2545 | 1.1066 | 1.1084 | 2.1967 | 2.1974 | 3.5521 | 2.8502 | 2.8494 | C4 | 3.6318 | 2.5773 | 1.5678 | | 1.5592 | 4.5323 | 3.9261 | 2.9258 | 2.1999 | 2.1919 | 1.1067 | 1.1066 | 2.2051 | 2.2059 | 2.2058 | C5 | 4.9675 | 3.9642 | 2.5936 | 1.5592 | | 5.9537 | 5.0540 | 4.3708 | 2.8285 | 2.8072 | 2.1919 | 2.1920 | 1.1035 | 1.1034 | 1.1034 | H6 | 1.0981 | 2.1403 | 3.5600 | 4.5323 | 5.9537 | | 1.8693 | 2.4705 | 3.7667 | 4.1949 | 4.7457 | 4.3948 | 6.7068 | 6.1157 | 6.3731 | H7 | 1.0985 | 2.1394 | 2.8121 | 3.9261 | 5.0540 | 1.8693 | | 3.1114 | 2.4680 | 3.5795 | 4.5558 | 3.7634 | 5.9087 | 4.9084 | 5.5408 | H8 | 2.1215 | 1.1023 | 2.2545 | 2.9258 | 4.3708 | 2.4705 | 3.1114 | | 3.1407 | 2.5974 | 2.7071 | 3.2482 | 5.0595 | 4.9161 | 4.5754 | H9 | 2.6707 | 2.1759 | 1.1066 | 2.1999 | 2.8285 | 3.7667 | 2.4680 | 3.1407 | | 1.7861 | 3.1075 | 2.5426 | 3.8387 | 2.6376 | 3.1869 | H10 | 3.2686 | 2.1777 | 1.1084 | 2.1919 | 2.8072 | 4.1949 | 3.5795 | 2.5974 | 1.7861 | | 2.5404 | 3.1029 | 3.8192 | 3.1641 | 2.6129 | H11 | 4.0137 | 2.7989 | 2.1967 | 1.1067 | 2.1919 | 4.7457 | 4.5558 | 2.7071 | 3.1075 | 2.5404 | | 1.7821 | 2.5300 | 3.1113 | 2.5485 | H12 | 3.5640 | 2.8136 | 2.1974 | 1.1066 | 2.1920 | 4.3948 | 3.7634 | 3.2482 | 2.5426 | 3.1029 | 1.7821 | | 2.5301 | 2.5486 | 3.1113 | H13 | 5.7808 | 4.7751 | 3.5521 | 2.2051 | 1.1035 | 6.7068 | 5.9087 | 5.0595 | 3.8387 | 3.8192 | 2.5300 | 2.5301 | | 1.7880 | 1.7882 | H14 | 5.0641 | 4.2844 | 2.8502 | 2.2059 | 1.1034 | 6.1157 | 4.9084 | 4.9161 | 2.6376 | 3.1641 | 3.1113 | 2.5486 | 1.7880 | | 1.7873 | H15 | 5.3908 | 4.2745 | 2.8494 | 2.2058 | 1.1034 | 6.3731 | 5.5408 | 4.5754 | 3.1869 | 2.6129 | 2.5485 | 3.1113 | 1.7882 | 1.7873 | |
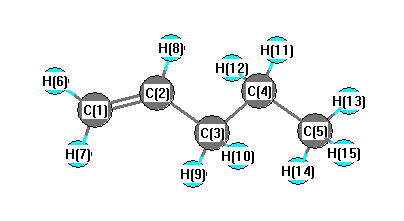 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.277 |
|
C1 |
C2 |
H8 |
119.611 |
| C2 |
C1 |
H6 |
121.737 |
|
C2 |
C1 |
H7 |
121.623 |
| C2 |
C3 |
C4 |
112.022 |
|
C2 |
C3 |
H9 |
109.474 |
| C2 |
C3 |
H10 |
109.510 |
|
C3 |
C2 |
H8 |
116.108 |
| C3 |
C4 |
C5 |
112.080 |
|
C3 |
C4 |
H11 |
109.223 |
| C3 |
C4 |
H12 |
109.282 |
|
C4 |
C3 |
H9 |
109.481 |
| C4 |
C3 |
H10 |
108.760 |
|
C4 |
C5 |
H13 |
110.657 |
| C4 |
C5 |
H14 |
110.719 |
|
C4 |
C5 |
H15 |
110.717 |
| C5 |
C4 |
H11 |
109.437 |
|
C5 |
C4 |
H12 |
109.449 |
| H6 |
C1 |
H7 |
116.640 |
|
H9 |
C3 |
H10 |
107.484 |
| H11 |
C4 |
H12 |
107.249 |
|
H13 |
C5 |
H14 |
108.233 |
| H13 |
C5 |
H15 |
108.248 |
|
H14 |
C5 |
H15 |
108.172 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -193.099583 |
| Energy at 298.15K | -193.109625 |
| HF Energy | -192.814504 |
| Nuclear repulsion energy | 170.703233 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3612 |
3211 |
0.56 |
|
|
|
| 2 |
A |
3557 |
3162 |
3.70 |
|
|
|
| 3 |
A |
3554 |
3159 |
3.76 |
|
|
|
| 4 |
A |
3515 |
3124 |
5.85 |
|
|
|
| 5 |
A |
3507 |
3118 |
1.70 |
|
|
|
| 6 |
A |
3500 |
3111 |
2.78 |
|
|
|
| 7 |
A |
3456 |
3072 |
5.67 |
|
|
|
| 8 |
A |
3402 |
3024 |
6.04 |
|
|
|
| 9 |
A |
3390 |
3013 |
4.27 |
|
|
|
| 10 |
A |
3375 |
3000 |
3.20 |
|
|
|
| 11 |
A |
1827 |
1624 |
0.11 |
|
|
|
| 12 |
A |
1744 |
1550 |
0.21 |
|
|
|
| 13 |
A |
1735 |
1543 |
1.38 |
|
|
|
| 14 |
A |
1719 |
1528 |
0.59 |
|
|
|
| 15 |
A |
1707 |
1517 |
0.69 |
|
|
|
| 16 |
A |
1626 |
1445 |
0.16 |
|
|
|
| 17 |
A |
1615 |
1436 |
0.36 |
|
|
|
| 18 |
A |
1555 |
1382 |
8.38 |
|
|
|
| 19 |
A |
1519 |
1351 |
11.69 |
|
|
|
| 20 |
A |
1440 |
1280 |
0.54 |
|
|
|
| 21 |
A |
1426 |
1267 |
3.50 |
|
|
|
| 22 |
A |
1395 |
1240 |
1.10 |
|
|
|
| 23 |
A |
1314 |
1168 |
0.78 |
|
|
|
| 24 |
A |
1222 |
1086 |
1.59 |
|
|
|
| 25 |
A |
1191 |
1058 |
0.94 |
|
|
|
| 26 |
A |
1139 |
1013 |
3.30 |
|
|
|
| 27 |
A |
1119 |
995 |
0.28 |
|
|
|
| 28 |
A |
1029 |
915 |
1.03 |
|
|
|
| 29 |
A |
995 |
885 |
11.98 |
|
|
|
| 30 |
A |
979 |
870 |
0.56 |
|
|
|
| 31 |
A |
951 |
846 |
0.71 |
|
|
|
| 32 |
A |
852 |
757 |
2.40 |
|
|
|
| 33 |
A |
679 |
604 |
2.91 |
|
|
|
| 34 |
A |
446 |
396 |
0.20 |
|
|
|
| 35 |
A |
412 |
366 |
0.73 |
|
|
|
| 36 |
A |
301 |
267 |
0.03 |
|
|
|
| 37 |
A |
219 |
195 |
0.03 |
|
|
|
| 38 |
A |
118 |
105 |
0.00 |
|
|
|
| 39 |
A |
81 |
72 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 33611.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29876.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-2.122 |
0.574 |
0.129 |
| C2 |
-1.203 |
-0.246 |
-0.418 |
| C3 |
-0.066 |
-0.926 |
0.369 |
| C4 |
1.356 |
-0.530 |
-0.161 |
| C5 |
1.670 |
0.985 |
0.027 |
| H6 |
-2.917 |
1.035 |
-0.473 |
| H7 |
-2.115 |
0.813 |
1.201 |
| H8 |
-1.240 |
-0.462 |
-1.499 |
| H9 |
-0.176 |
-2.027 |
0.289 |
| H10 |
-0.149 |
-0.660 |
1.440 |
| H11 |
2.115 |
-1.131 |
0.375 |
| H12 |
1.429 |
-0.792 |
-1.234 |
| H13 |
2.679 |
1.221 |
-0.353 |
| H14 |
0.940 |
1.607 |
-0.517 |
| H15 |
1.630 |
1.261 |
1.095 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
| C1 | | 1.3482 | 2.5565 | 3.6605 | 3.8163 | 1.0982 | 1.0986 | 2.1215 | 3.2524 | 2.6710 | 4.5739 | 4.0421 | 4.8687 | 3.2960 | 3.9348 |
C2 | 1.3482 | | 1.5406 | 2.5871 | 3.1577 | 2.1404 | 2.1395 | 1.1024 | 2.1738 | 2.1757 | 3.5243 | 2.8095 | 4.1507 | 2.8353 | 3.5477 | C3 | 2.5565 | 1.5406 | | 1.5684 | 2.6049 | 3.5608 | 2.8137 | 2.2541 | 1.1086 | 1.1067 | 2.1908 | 2.1967 | 3.5594 | 2.8663 | 2.8615 | C4 | 3.6605 | 2.5871 | 1.5684 | | 1.5587 | 4.5605 | 3.9634 | 2.9210 | 2.1882 | 2.2013 | 1.1069 | 1.1069 | 2.2034 | 2.2061 | 2.2046 | C5 | 3.8163 | 3.1577 | 2.6049 | 1.5587 | | 4.6144 | 3.9675 | 3.5908 | 3.5421 | 2.8309 | 2.1896 | 2.1919 | 1.1034 | 1.1031 | 1.1034 | H6 | 1.0982 | 2.1404 | 3.5608 | 4.5605 | 4.6144 | | 1.8694 | 2.4704 | 4.1794 | 3.7671 | 5.5431 | 4.7753 | 5.5999 | 3.8991 | 4.8144 | H7 | 1.0986 | 2.1395 | 2.8137 | 3.9634 | 3.9675 | 1.8694 | | 3.1115 | 3.5577 | 2.4683 | 4.7282 | 4.5906 | 5.0566 | 3.5945 | 3.7735 | H8 | 2.1215 | 1.1024 | 2.2541 | 2.9210 | 3.5908 | 2.4704 | 3.1115 | | 2.6036 | 3.1407 | 3.9009 | 2.7030 | 4.4167 | 3.1622 | 4.2346 | H9 | 3.2524 | 2.1738 | 1.1086 | 2.1882 | 3.5421 | 4.1794 | 3.5577 | 2.6036 | | 1.7863 | 2.4612 | 2.5347 | 4.3717 | 3.8859 | 3.8363 | H10 | 2.6710 | 2.1757 | 1.1067 | 2.2013 | 2.8309 | 3.7671 | 2.4683 | 3.1407 | 1.7863 | | 2.5458 | 3.1082 | 3.8411 | 3.1871 | 2.6412 | H11 | 4.5739 | 3.5243 | 2.1908 | 1.1069 | 2.1896 | 5.5431 | 4.7282 | 3.9009 | 2.4612 | 2.5458 | | 1.7821 | 2.5260 | 3.1100 | 2.5441 | H12 | 4.0421 | 2.8095 | 2.1967 | 1.1069 | 2.1919 | 4.7753 | 4.5906 | 2.7030 | 2.5347 | 3.1082 | 1.7821 | | 2.5277 | 2.5510 | 3.1107 | H13 | 4.8687 | 4.1507 | 3.5594 | 2.2034 | 1.1034 | 5.5999 | 5.0566 | 4.4167 | 4.3717 | 3.8411 | 2.5260 | 2.5277 | | 1.7887 | 1.7882 | H14 | 3.2960 | 2.8353 | 2.8663 | 2.2061 | 1.1031 | 3.8991 | 3.5945 | 3.1622 | 3.8859 | 3.1871 | 3.1100 | 2.5510 | 1.7887 | | 1.7869 | H15 | 3.9348 | 3.5477 | 2.8615 | 2.2046 | 1.1034 | 4.8144 | 3.7735 | 4.2346 | 3.8363 | 2.6412 | 2.5441 | 3.1107 | 1.7882 | 1.7869 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.356 |
|
C1 |
C2 |
H8 |
119.589 |
| C2 |
C1 |
H6 |
121.741 |
|
C2 |
C1 |
H7 |
121.618 |
| C2 |
C3 |
C4 |
112.633 |
|
C2 |
C3 |
H9 |
109.187 |
| C2 |
C3 |
H10 |
109.442 |
|
C3 |
C2 |
H8 |
116.054 |
| C3 |
C4 |
C5 |
112.817 |
|
C3 |
C4 |
H11 |
108.717 |
| C3 |
C4 |
H12 |
109.174 |
|
C4 |
C3 |
H9 |
108.428 |
| C4 |
C3 |
H10 |
109.536 |
|
C4 |
C5 |
H13 |
110.563 |
| C4 |
C5 |
H14 |
110.792 |
|
C4 |
C5 |
H15 |
110.655 |
| C5 |
C4 |
H11 |
109.284 |
|
C5 |
C4 |
H12 |
109.468 |
| H6 |
C1 |
H7 |
116.641 |
|
H9 |
C3 |
H10 |
107.477 |
| H11 |
C4 |
H12 |
107.224 |
|
H13 |
C5 |
H14 |
108.320 |
| H13 |
C5 |
H15 |
108.254 |
|
H14 |
C5 |
H15 |
108.164 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
