return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H9SH (1-Butanethiol)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-548.874164
Energy at 298.15K 
HF Energy-548.632153
Nuclear repulsion energy220.038374
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3554 3159 3.43      
2 A' 3405 3026 10.15      
3 A' 3400 3022 0.48      
4 A' 3374 2999 3.76      
5 A' 3369 2995 5.63      
6 A' 3086 2744 25.43      
7 A' 1746 1552 0.14      
8 A' 1740 1546 1.20      
9 A' 1726 1535 0.25      
10 A' 1714 1524 0.69      
11 A' 1626 1445 0.35      
12 A' 1595 1418 3.94      
13 A' 1515 1347 1.63      
14 A' 1415 1257 34.73      
15 A' 1263 1123 0.48      
16 A' 1191 1059 1.21      
17 A' 1165 1035 1.58      
18 A' 1076 956 5.14      
19 A' 988 878 0.92      
20 A' 915 813 0.48      
21 A' 411 365 0.59      
22 A' 345 307 0.33      
23 A' 162 144 0.50      
24 A" 3555 3160 4.98      
25 A" 3521 3130 10.61      
26 A" 3510 3120 0.23      
27 A" 3480 3093 7.23      
28 A" 1737 1544 1.45      
29 A" 1477 1313 0.00      
30 A" 1456 1295 0.01      
31 A" 1385 1231 0.03      
32 A" 1213 1078 0.30      
33 A" 1033 918 0.14      
34 A" 875 778 0.32      
35 A" 798 709 3.09      
36 A" 241 215 0.00      
37 A" 197 175 9.10      
38 A" 113 101 1.44      
39 A" 98 87 0.18      

Unscaled Zero Point Vibrational Energy (zpe) 32733.8 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 29097.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.52688 0.04292 0.04092

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
S1 1.463 -1.839 0.000
C2 -0.169 -1.014 0.000
C3 0.000 0.541 0.000
C4 -1.378 1.281 0.000
C5 -1.214 2.831 0.000
H6 0.944 -3.085 0.000
H7 -0.759 -1.312 0.891
H8 -0.759 -1.312 -0.891
H9 0.581 0.848 -0.891
H10 0.581 0.848 0.891
H11 -1.958 0.973 0.891
H12 -1.958 0.973 -0.891
H13 -2.199 3.328 0.000
H14 -0.659 3.165 0.894
H15 -0.659 3.165 -0.894

Atom - Atom Distances (Å)
  S1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13 H14 H15
S11.82902.79384.21945.38311.34982.45132.45132.96562.96564.51694.51696.33315.50855.5085
C21.82901.56462.59383.98502.35111.10861.10862.19692.19692.81792.81794.79314.30194.3019
C32.79381.56461.56372.59213.74692.19122.19121.10691.10692.19382.19383.55002.84932.8493
C44.21942.59381.56371.55924.94482.81012.81012.19452.19451.10671.10672.20562.20572.2057
C55.38313.98502.59211.55926.29764.26204.26202.81882.81882.19122.19121.10341.10331.1033
H61.34982.35113.74694.94486.29762.61502.61504.04944.04945.06765.06767.14176.51416.5141
H72.45131.10862.19122.81014.26202.61501.78103.10392.54192.58003.13514.93884.47794.8202
H82.45131.10862.19122.81014.26202.61501.78102.54193.10393.13512.58004.93884.82024.4779
H92.96562.19691.10692.19452.81884.04943.10392.54191.78143.10372.54163.82993.17612.6276
H102.96562.19691.10692.19452.81884.04942.54193.10391.78142.54163.10373.82992.62763.1761
H114.51692.81792.19381.10672.19125.06762.58003.13513.10372.54161.78132.52922.54813.1106
H124.51692.81792.19381.10672.19125.06763.13512.58002.54163.10371.78132.52923.11062.5481
H136.33314.79313.55002.20561.10347.14174.93884.93883.82993.82992.52922.52921.78801.7880
H145.50854.30192.84932.20571.10336.51414.47794.82023.17612.62762.54813.11061.78801.7871
H155.50854.30192.84932.20571.10336.51414.82024.47792.62763.17613.11062.54811.78801.7871

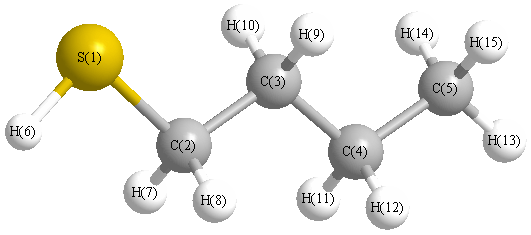
picture of 1-Butanethiol state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
S1 C2 C3 110.584 S1 C2 H7 110.715
S1 C2 H8 110.715 C2 S1 H6 94.186
C2 C3 C4 112.019 C2 C3 H9 109.447
C2 C3 H10 109.447 C3 C2 H7 108.919
C3 C2 H8 108.919 C3 C4 C5 112.200
C3 C4 H11 109.279 C3 C4 H12 109.279
C4 C3 H9 109.321 C4 C3 H10 109.321
C4 C5 H13 110.694 C4 C5 H14 110.710
C4 C5 H15 110.710 C5 C4 H11 109.383
C5 C4 H12 109.383 H7 C2 H8 106.890
H9 C3 H10 107.162 H11 C4 H12 107.187
H13 C5 H14 108.234 H13 C5 H15 108.234
H14 C5 H15 108.162
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability