Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -227.139667 |
| Energy at 298.15K | -227.145414 |
| HF Energy | -226.850261 |
| Nuclear repulsion energy | 157.929838 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3637 |
3233 |
0.10 |
|
|
|
| 2 |
A |
3636 |
3232 |
0.28 |
|
|
|
| 3 |
A |
3507 |
3117 |
0.14 |
|
|
|
| 4 |
A |
3489 |
3101 |
0.85 |
|
|
|
| 5 |
A |
3475 |
3089 |
25.64 |
|
|
|
| 6 |
A |
3464 |
3079 |
28.74 |
|
|
|
| 7 |
A |
1812 |
1610 |
116.57 |
|
|
|
| 8 |
A |
1799 |
1599 |
7.70 |
|
|
|
| 9 |
A |
1597 |
1420 |
10.57 |
|
|
|
| 10 |
A |
1575 |
1400 |
47.47 |
|
|
|
| 11 |
A |
1435 |
1275 |
2.68 |
|
|
|
| 12 |
A |
1416 |
1259 |
11.70 |
|
|
|
| 13 |
A |
1325 |
1178 |
105.05 |
|
|
|
| 14 |
A |
1195 |
1062 |
4.39 |
|
|
|
| 15 |
A |
1097 |
976 |
11.59 |
|
|
|
| 16 |
A |
1087 |
966 |
16.07 |
|
|
|
| 17 |
A |
1076 |
956 |
8.97 |
|
|
|
| 18 |
A |
932 |
829 |
3.10 |
|
|
|
| 19 |
A |
922 |
820 |
16.32 |
|
|
|
| 20 |
A |
903 |
802 |
20.61 |
|
|
|
| 21 |
A |
747 |
664 |
0.91 |
|
|
|
| 22 |
A |
706 |
627 |
1.42 |
|
|
|
| 23 |
A |
613 |
545 |
1.04 |
|
|
|
| 24 |
A |
449 |
399 |
0.94 |
|
|
|
| 25 |
A |
307 |
273 |
1.23 |
|
|
|
| 26 |
A |
186 |
166 |
4.71 |
|
|
|
| 27 |
A |
87 |
77 |
0.41 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21235.7 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 18876.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.061 |
-0.858 |
0.049 |
| C2 |
1.309 |
-0.497 |
0.010 |
| C3 |
-0.925 |
0.253 |
0.268 |
| C4 |
1.869 |
0.726 |
-0.142 |
| C5 |
-2.202 |
0.230 |
-0.175 |
| H6 |
1.935 |
-1.404 |
0.088 |
| H7 |
-0.530 |
1.088 |
0.876 |
| H8 |
2.961 |
0.815 |
-0.160 |
| H9 |
1.286 |
1.645 |
-0.255 |
| H10 |
-2.887 |
1.049 |
0.068 |
| H11 |
-2.584 |
-0.607 |
-0.772 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.4172 | 1.4241 | 2.5042 | 2.4112 | 2.0703 | 2.1655 | 3.4601 | 2.8586 | 3.4091 | 2.6650 |
C2 | 1.4172 | | 2.3703 | 1.3538 | 3.5895 | 1.1051 | 2.5770 | 2.1161 | 2.1586 | 4.4720 | 3.9723 | C3 | 1.4241 | 2.3703 | | 2.8633 | 1.3517 | 3.3105 | 1.1058 | 3.9492 | 2.6641 | 2.1269 | 2.1389 | C4 | 2.5042 | 1.3538 | 2.8633 | | 4.1011 | 2.1431 | 2.6308 | 1.0953 | 1.0946 | 4.7720 | 4.6912 | C5 | 2.4112 | 3.5895 | 1.3517 | 4.1011 | | 4.4556 | 2.1536 | 5.1954 | 3.7646 | 1.0957 | 1.0966 | H6 | 2.0703 | 1.1051 | 3.3105 | 2.1431 | 4.4556 | | 3.5925 | 2.4564 | 3.1362 | 5.4105 | 4.6691 | H7 | 2.1655 | 2.5770 | 1.1058 | 2.6308 | 2.1536 | 3.5925 | | 3.6509 | 2.2100 | 2.4925 | 3.1322 | H8 | 3.4601 | 2.1161 | 3.9492 | 1.0953 | 5.1954 | 2.4564 | 3.6509 | | 1.8721 | 5.8570 | 5.7570 | H9 | 2.8586 | 2.1586 | 2.6641 | 1.0946 | 3.7646 | 3.1362 | 2.2100 | 1.8721 | | 4.2276 | 4.5075 | H10 | 3.4091 | 4.4720 | 2.1269 | 4.7720 | 1.0957 | 5.4105 | 2.4925 | 5.8570 | 4.2276 | | 1.8816 | H11 | 2.6650 | 3.9723 | 2.1389 | 4.6912 | 1.0966 | 4.6691 | 3.1322 | 5.7570 | 4.5075 | 1.8816 | |
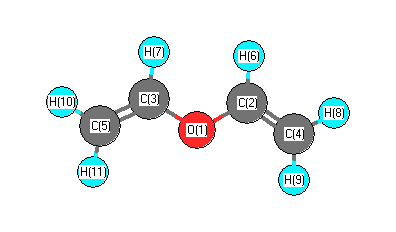 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
129.293 |
|
O1 |
C2 |
H6 |
109.711 |
| O1 |
C3 |
C5 |
120.579 |
|
O1 |
C3 |
H7 |
117.174 |
| C2 |
O1 |
C3 |
113.065 |
|
C2 |
C4 |
H8 |
119.177 |
| C2 |
C4 |
H9 |
123.332 |
|
C3 |
C5 |
H10 |
120.332 |
| C3 |
C5 |
H11 |
121.421 |
|
C4 |
C2 |
H6 |
120.959 |
| C5 |
C3 |
H7 |
122.084 |
|
H8 |
C4 |
H9 |
117.491 |
| H10 |
C5 |
H11 |
118.242 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
