Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -247.522077 |
| Energy at 298.15K | -247.536233 |
| Nuclear repulsion energy | 253.355857 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3637 |
3233 |
15.25 |
|
|
|
| 2 |
A' |
3520 |
3129 |
11.95 |
|
|
|
| 3 |
A' |
3512 |
3122 |
3.87 |
|
|
|
| 4 |
A' |
3509 |
3119 |
0.37 |
|
|
|
| 5 |
A' |
3402 |
3024 |
4.82 |
|
|
|
| 6 |
A' |
3398 |
3020 |
3.37 |
|
|
|
| 7 |
A' |
3394 |
3017 |
9.32 |
|
|
|
| 8 |
A' |
1727 |
1535 |
0.03 |
|
|
|
| 9 |
A' |
1710 |
1520 |
0.80 |
|
|
|
| 10 |
A' |
1698 |
1510 |
0.93 |
|
|
|
| 11 |
A' |
1561 |
1388 |
0.56 |
|
|
|
| 12 |
A' |
1552 |
1379 |
12.13 |
|
|
|
| 13 |
A' |
1461 |
1298 |
6.23 |
|
|
|
| 14 |
A' |
1418 |
1261 |
6.87 |
|
|
|
| 15 |
A' |
1346 |
1196 |
5.08 |
|
|
|
| 16 |
A' |
1161 |
1032 |
1.25 |
|
|
|
| 17 |
A' |
1110 |
987 |
7.72 |
|
|
|
| 18 |
A' |
1082 |
962 |
21.38 |
|
|
|
| 19 |
A' |
954 |
848 |
12.08 |
|
|
|
| 20 |
A' |
920 |
818 |
10.05 |
|
|
|
| 21 |
A' |
873 |
776 |
13.11 |
|
|
|
| 22 |
A' |
587 |
522 |
0.65 |
|
|
|
| 23 |
A' |
466 |
415 |
0.86 |
|
|
|
| 24 |
A' |
383 |
340 |
4.13 |
|
|
|
| 25 |
A' |
243 |
216 |
3.74 |
|
|
|
| 26 |
A" |
3515 |
3125 |
0.38 |
|
|
|
| 27 |
A" |
3508 |
3118 |
3.45 |
|
|
|
| 28 |
A" |
3399 |
3021 |
7.14 |
|
|
|
| 29 |
A" |
3393 |
3016 |
6.22 |
|
|
|
| 30 |
A" |
1714 |
1523 |
0.31 |
|
|
|
| 31 |
A" |
1697 |
1509 |
0.01 |
|
|
|
| 32 |
A" |
1653 |
1470 |
2.68 |
|
|
|
| 33 |
A" |
1571 |
1397 |
1.20 |
|
|
|
| 34 |
A" |
1537 |
1366 |
7.49 |
|
|
|
| 35 |
A" |
1493 |
1327 |
1.82 |
|
|
|
| 36 |
A" |
1431 |
1272 |
1.12 |
|
|
|
| 37 |
A" |
1331 |
1183 |
1.82 |
|
|
|
| 38 |
A" |
1248 |
1109 |
2.52 |
|
|
|
| 39 |
A" |
1232 |
1095 |
1.27 |
|
|
|
| 40 |
A" |
1178 |
1047 |
0.87 |
|
|
|
| 41 |
A" |
1042 |
927 |
4.04 |
|
|
|
| 42 |
A" |
975 |
866 |
0.07 |
|
|
|
| 43 |
A" |
892 |
793 |
0.10 |
|
|
|
| 44 |
A" |
474 |
421 |
1.63 |
|
|
|
| 45 |
A" |
232 |
207 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 39069.0 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 34728.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.671 |
1.349 |
0.000 |
| H2 |
0.596 |
2.453 |
0.000 |
| H3 |
1.746 |
1.087 |
0.000 |
| C4 |
-0.003 |
0.762 |
1.281 |
| C5 |
-0.003 |
0.762 |
-1.281 |
| C6 |
-0.003 |
-0.806 |
-1.234 |
| C7 |
-0.003 |
-0.806 |
1.234 |
| N8 |
-0.662 |
-1.402 |
0.000 |
| H9 |
-1.639 |
-0.986 |
0.000 |
| H10 |
0.529 |
1.105 |
2.187 |
| H11 |
0.529 |
1.105 |
-2.187 |
| H12 |
-1.047 |
1.125 |
1.348 |
| H13 |
-1.047 |
1.125 |
-1.348 |
| H14 |
1.042 |
-1.174 |
-1.264 |
| H15 |
1.042 |
-1.174 |
1.264 |
| H16 |
-0.532 |
-1.208 |
-2.122 |
| H17 |
-0.532 |
-1.208 |
2.122 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
H3 |
C4 |
C5 |
C6 |
C7 |
N8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
| C1 | | 1.1064 | 1.1066 | 1.5618 | 1.5618 | 2.5734 | 2.5734 | 3.0570 | 3.2845 | 2.2057 | 2.2057 | 2.1952 | 2.1952 | 2.8467 | 2.8467 | 3.5339 | 3.5339 |
H2 | 1.1064 | | 1.7855 | 2.2041 | 2.2041 | 3.5360 | 3.5360 | 4.0551 | 4.1013 | 2.5706 | 2.5706 | 2.5058 | 2.5058 | 3.8672 | 3.8672 | 4.3791 | 4.3791 | H3 | 1.1066 | 1.7855 | | 2.1924 | 2.1924 | 2.8582 | 2.8582 | 3.4635 | 3.9695 | 2.5032 | 2.5032 | 3.1017 | 3.1017 | 2.6851 | 2.6851 | 3.8679 | 3.8679 | C4 | 1.5618 | 2.2041 | 2.1924 | | 2.5610 | 2.9634 | 1.5689 | 2.5994 | 2.7147 | 1.1062 | 3.5253 | 1.1070 | 2.8514 | 3.3642 | 2.2007 | 3.9671 | 2.2063 | C5 | 1.5618 | 2.2041 | 2.1924 | 2.5610 | | 1.5689 | 2.9634 | 2.5994 | 2.7147 | 3.5253 | 1.1062 | 2.8514 | 1.1070 | 2.2007 | 3.3642 | 2.2063 | 3.9671 | C6 | 2.5734 | 3.5360 | 2.8582 | 2.9634 | 1.5689 | | 2.4679 | 1.5203 | 2.0565 | 3.9548 | 2.2008 | 3.3891 | 2.1982 | 1.1088 | 2.7329 | 1.1089 | 3.4212 | C7 | 2.5734 | 3.5360 | 2.8582 | 1.5689 | 2.9634 | 2.4679 | | 1.5203 | 2.0565 | 2.2008 | 3.9548 | 2.1982 | 3.3891 | 2.7329 | 1.1088 | 3.4212 | 1.1089 | N8 | 3.0570 | 4.0551 | 3.4635 | 2.5994 | 2.5994 | 1.5203 | 1.5203 | | 1.0619 | 3.5337 | 3.5337 | 2.8902 | 2.8902 | 2.1336 | 2.1336 | 2.1350 | 2.1350 | H9 | 3.2845 | 4.1013 | 3.9695 | 2.7147 | 2.7147 | 2.0565 | 2.0565 | 1.0619 | | 3.7223 | 3.7223 | 2.5739 | 2.5739 | 2.9697 | 2.9697 | 2.4037 | 2.4037 | H10 | 2.2057 | 2.5706 | 2.5032 | 1.1062 | 3.5253 | 3.9548 | 2.2008 | 3.5337 | 3.7223 | | 4.3749 | 1.7859 | 3.8711 | 4.1677 | 2.5118 | 5.0046 | 2.5450 | H11 | 2.2057 | 2.5706 | 2.5032 | 3.5253 | 1.1062 | 2.2008 | 3.9548 | 3.5337 | 3.7223 | 4.3749 | | 3.8711 | 1.7859 | 2.5118 | 4.1677 | 2.5450 | 5.0046 | H12 | 2.1952 | 2.5058 | 3.1017 | 1.1070 | 2.8514 | 3.3891 | 2.1982 | 2.8902 | 2.5739 | 1.7859 | 3.8711 | | 2.6962 | 4.0590 | 3.1079 | 4.2132 | 2.5114 | H13 | 2.1952 | 2.5058 | 3.1017 | 2.8514 | 1.1070 | 2.1982 | 3.3891 | 2.8902 | 2.5739 | 3.8711 | 1.7859 | 2.6962 | | 3.1079 | 4.0590 | 2.5114 | 4.2132 | H14 | 2.8467 | 3.8672 | 2.6851 | 3.3642 | 2.2007 | 1.1088 | 2.7329 | 2.1336 | 2.9697 | 4.1677 | 2.5118 | 4.0590 | 3.1079 | | 2.5282 | 1.7929 | 3.7343 | H15 | 2.8467 | 3.8672 | 2.6851 | 2.2007 | 3.3642 | 2.7329 | 1.1088 | 2.1336 | 2.9697 | 2.5118 | 4.1677 | 3.1079 | 4.0590 | 2.5282 | | 3.7343 | 1.7929 | H16 | 3.5339 | 4.3791 | 3.8679 | 3.9671 | 2.2063 | 1.1089 | 3.4212 | 2.1350 | 2.4037 | 5.0046 | 2.5450 | 4.2132 | 2.5114 | 1.7929 | 3.7343 | | 4.2444 | H17 | 3.5339 | 4.3791 | 3.8679 | 2.2063 | 3.9671 | 3.4212 | 1.1089 | 2.1350 | 2.4037 | 2.5450 | 5.0046 | 2.5114 | 4.2132 | 3.7343 | 1.7929 | 4.2444 | |
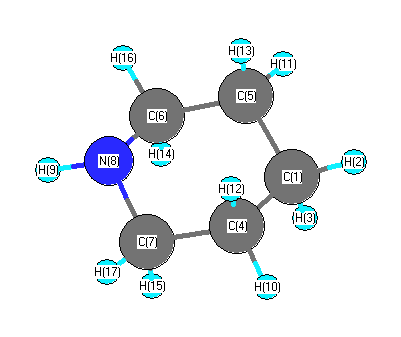 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
C7 |
110.572 |
|
C1 |
C4 |
H10 |
110.359 |
| C1 |
C4 |
H12 |
109.502 |
|
C1 |
C5 |
C6 |
110.572 |
| C1 |
C5 |
H11 |
110.359 |
|
C1 |
C5 |
H13 |
109.502 |
| H2 |
C1 |
H3 |
107.573 |
|
H2 |
C1 |
C4 |
110.224 |
| H2 |
C1 |
C5 |
110.224 |
|
H3 |
C1 |
C4 |
109.311 |
| H3 |
C1 |
C5 |
109.311 |
|
C4 |
C1 |
C5 |
110.149 |
| C4 |
C7 |
N8 |
114.580 |
|
C4 |
C7 |
H15 |
109.346 |
| C4 |
C7 |
H17 |
109.772 |
|
C5 |
C6 |
N8 |
114.580 |
| C5 |
C6 |
H14 |
109.346 |
|
C5 |
C6 |
H16 |
109.772 |
| C6 |
C5 |
H11 |
109.498 |
|
C6 |
C5 |
H13 |
109.253 |
| C6 |
N8 |
C7 |
108.519 |
|
C6 |
N8 |
H9 |
104.167 |
| C7 |
C4 |
H10 |
109.498 |
|
C7 |
C4 |
H12 |
109.253 |
| C7 |
N8 |
H9 |
104.167 |
|
N8 |
C6 |
H14 |
107.460 |
| N8 |
C6 |
H16 |
107.567 |
|
N8 |
C7 |
H15 |
107.460 |
| N8 |
C7 |
H17 |
107.567 |
|
H10 |
C4 |
H12 |
107.598 |
| H11 |
C5 |
H13 |
107.598 |
|
H14 |
C6 |
H16 |
107.891 |
| H15 |
C7 |
H17 |
107.891 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
