Jump to
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -306.815159 |
| Energy at 298.15K | -306.830417 |
| HF Energy | -306.455111 |
| Nuclear repulsion energy | 308.313843 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3555 |
3160 |
4.36 |
|
|
|
| 2 |
A1 |
3413 |
3034 |
6.91 |
|
|
|
| 3 |
A1 |
3374 |
2999 |
1.88 |
|
|
|
| 4 |
A1 |
3318 |
2949 |
21.67 |
|
|
|
| 5 |
A1 |
1746 |
1552 |
0.07 |
|
|
|
| 6 |
A1 |
1741 |
1547 |
1.42 |
|
|
|
| 7 |
A1 |
1714 |
1524 |
0.02 |
|
|
|
| 8 |
A1 |
1626 |
1445 |
0.10 |
|
|
|
| 9 |
A1 |
1608 |
1430 |
0.79 |
|
|
|
| 10 |
A1 |
1487 |
1322 |
2.81 |
|
|
|
| 11 |
A1 |
1287 |
1144 |
0.00 |
|
|
|
| 12 |
A1 |
1179 |
1048 |
0.25 |
|
|
|
| 13 |
A1 |
1117 |
992 |
2.68 |
|
|
|
| 14 |
A1 |
1030 |
915 |
1.93 |
|
|
|
| 15 |
A1 |
482 |
428 |
0.03 |
|
|
|
| 16 |
A1 |
328 |
292 |
0.00 |
|
|
|
| 17 |
A1 |
111 |
99 |
0.11 |
|
|
|
| 18 |
A2 |
3554 |
3159 |
0.00 |
|
|
|
| 19 |
A2 |
3525 |
3134 |
0.00 |
|
|
|
| 20 |
A2 |
3426 |
3045 |
0.00 |
|
|
|
| 21 |
A2 |
1737 |
1544 |
0.00 |
|
|
|
| 22 |
A2 |
1454 |
1292 |
0.00 |
|
|
|
| 23 |
A2 |
1399 |
1243 |
0.00 |
|
|
|
| 24 |
A2 |
1268 |
1128 |
0.00 |
|
|
|
| 25 |
A2 |
990 |
880 |
0.00 |
|
|
|
| 26 |
A2 |
825 |
733 |
0.00 |
|
|
|
| 27 |
A2 |
240 |
214 |
0.00 |
|
|
|
| 28 |
A2 |
136 |
121 |
0.00 |
|
|
|
| 29 |
A2 |
77 |
68 |
0.00 |
|
|
|
| 30 |
B1 |
3554 |
3159 |
9.62 |
|
|
|
| 31 |
B1 |
3525 |
3134 |
5.09 |
|
|
|
| 32 |
B1 |
3422 |
3042 |
33.45 |
|
|
|
| 33 |
B1 |
1737 |
1544 |
3.30 |
|
|
|
| 34 |
B1 |
1453 |
1291 |
0.01 |
|
|
|
| 35 |
B1 |
1397 |
1242 |
2.28 |
|
|
|
| 36 |
B1 |
1283 |
1140 |
7.48 |
|
|
|
| 37 |
B1 |
1003 |
892 |
3.27 |
|
|
|
| 38 |
B1 |
829 |
737 |
2.70 |
|
|
|
| 39 |
B1 |
239 |
213 |
0.09 |
|
|
|
| 40 |
B1 |
153 |
136 |
4.16 |
|
|
|
| 41 |
B1 |
56 |
50 |
0.02 |
|
|
|
| 42 |
B2 |
3555 |
3160 |
1.55 |
|
|
|
| 43 |
B2 |
3413 |
3034 |
0.11 |
|
|
|
| 44 |
B2 |
3374 |
2999 |
4.90 |
|
|
|
| 45 |
B2 |
3319 |
2950 |
5.92 |
|
|
|
| 46 |
B2 |
1745 |
1551 |
1.17 |
|
|
|
| 47 |
B2 |
1736 |
1543 |
2.73 |
|
|
|
| 48 |
B2 |
1713 |
1523 |
1.23 |
|
|
|
| 49 |
B2 |
1624 |
1444 |
4.02 |
|
|
|
| 50 |
B2 |
1580 |
1405 |
14.45 |
|
|
|
| 51 |
B2 |
1459 |
1297 |
80.35 |
|
|
|
| 52 |
B2 |
1279 |
1137 |
13.59 |
|
|
|
| 53 |
B2 |
1231 |
1094 |
1.31 |
|
|
|
| 54 |
B2 |
1166 |
1036 |
0.01 |
|
|
|
| 55 |
B2 |
996 |
885 |
2.17 |
|
|
|
| 56 |
B2 |
515 |
458 |
1.20 |
|
|
|
| 57 |
B2 |
262 |
233 |
0.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 47681.2 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 42383.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
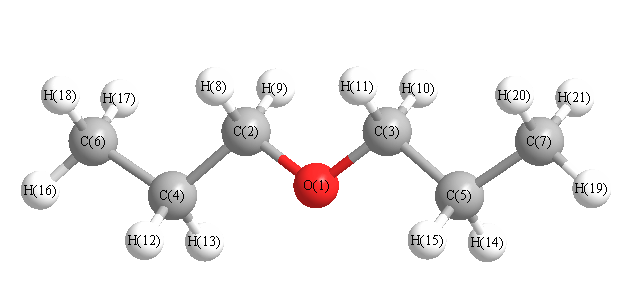
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.495 |
| C2 |
0.000 |
1.178 |
-0.373 |
| C3 |
0.000 |
-1.178 |
-0.373 |
| C4 |
0.000 |
2.449 |
0.540 |
| C5 |
0.000 |
-2.449 |
0.540 |
| C6 |
0.000 |
3.764 |
-0.297 |
| C7 |
0.000 |
-3.764 |
-0.297 |
| H8 |
0.897 |
1.192 |
-1.037 |
| H9 |
-0.897 |
1.192 |
-1.037 |
| H10 |
-0.897 |
-1.192 |
-1.037 |
| H11 |
0.897 |
-1.192 |
-1.037 |
| H12 |
0.892 |
2.420 |
1.194 |
| H13 |
-0.892 |
2.420 |
1.194 |
| H14 |
-0.892 |
-2.420 |
1.194 |
| H15 |
0.892 |
-2.420 |
1.194 |
| H16 |
0.000 |
4.646 |
0.366 |
| H17 |
-0.894 |
3.816 |
-0.942 |
| H18 |
0.894 |
3.816 |
-0.942 |
| H19 |
0.000 |
-4.646 |
0.366 |
| H20 |
0.894 |
-3.816 |
-0.942 |
| H21 |
-0.894 |
-3.816 |
-0.942 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4634 | 1.4634 | 2.4495 | 2.4495 | 3.8460 | 3.8460 | 2.1388 | 2.1388 | 2.1388 | 2.1388 | 2.6725 | 2.6725 | 2.6725 | 2.6725 | 4.6473 | 4.1748 | 4.1748 | 4.6473 | 4.1748 | 4.1748 |
C2 | 1.4634 | | 2.3567 | 1.5647 | 3.7405 | 2.5863 | 4.9425 | 1.1167 | 1.1167 | 2.6202 | 2.6202 | 2.1893 | 2.1893 | 4.0251 | 4.0251 | 3.5450 | 2.8430 | 2.8430 | 5.8705 | 5.1059 | 5.1059 | C3 | 1.4634 | 2.3567 | | 3.7405 | 1.5647 | 4.9425 | 2.5863 | 2.6202 | 2.6202 | 1.1167 | 1.1167 | 4.0251 | 4.0251 | 2.1893 | 2.1893 | 5.8705 | 5.1059 | 5.1059 | 3.5450 | 2.8430 | 2.8430 | C4 | 2.4495 | 1.5647 | 3.7405 | | 4.8981 | 1.5585 | 6.2688 | 2.2077 | 2.2077 | 4.0683 | 4.0683 | 1.1061 | 1.1061 | 4.9935 | 4.9935 | 2.2034 | 2.2058 | 2.2058 | 7.0967 | 6.5002 | 6.5002 | C5 | 2.4495 | 3.7405 | 1.5647 | 4.8981 | | 6.2688 | 1.5585 | 4.0683 | 4.0683 | 2.2077 | 2.2077 | 4.9935 | 4.9935 | 1.1061 | 1.1061 | 7.0967 | 6.5002 | 6.5002 | 2.2034 | 2.2058 | 2.2058 | C6 | 3.8460 | 2.5863 | 4.9425 | 1.5585 | 6.2688 | | 7.5271 | 2.8224 | 2.8224 | 5.0902 | 5.0902 | 2.1960 | 2.1960 | 6.4234 | 6.4234 | 1.1033 | 1.1035 | 1.1035 | 8.4351 | 7.6597 | 7.6597 | C7 | 3.8460 | 4.9425 | 2.5863 | 6.2688 | 1.5585 | 7.5271 | | 5.0902 | 5.0902 | 2.8224 | 2.8224 | 6.4234 | 6.4234 | 2.1960 | 2.1960 | 8.4351 | 7.6597 | 7.6597 | 1.1033 | 1.1035 | 1.1035 | H8 | 2.1388 | 1.1167 | 2.6202 | 2.2077 | 4.0683 | 2.8224 | 5.0902 | | 1.7943 | 2.9838 | 2.3840 | 2.5473 | 3.1127 | 4.6076 | 4.2462 | 3.8342 | 3.1787 | 2.6261 | 6.0705 | 5.0094 | 5.3199 | H9 | 2.1388 | 1.1167 | 2.6202 | 2.2077 | 4.0683 | 2.8224 | 5.0902 | 1.7943 | | 2.3840 | 2.9838 | 3.1127 | 2.5473 | 4.2462 | 4.6076 | 3.8342 | 2.6261 | 3.1787 | 6.0705 | 5.3199 | 5.0094 | H10 | 2.1388 | 2.6202 | 1.1167 | 4.0683 | 2.2077 | 5.0902 | 2.8224 | 2.9838 | 2.3840 | | 1.7943 | 4.6076 | 4.2462 | 2.5473 | 3.1127 | 6.0705 | 5.0094 | 5.3199 | 3.8342 | 3.1787 | 2.6261 | H11 | 2.1388 | 2.6202 | 1.1167 | 4.0683 | 2.2077 | 5.0902 | 2.8224 | 2.3840 | 2.9838 | 1.7943 | | 4.2462 | 4.6076 | 3.1127 | 2.5473 | 6.0705 | 5.3199 | 5.0094 | 3.8342 | 2.6261 | 3.1787 | H12 | 2.6725 | 2.1893 | 4.0251 | 1.1061 | 4.9935 | 2.1960 | 6.4234 | 2.5473 | 3.1127 | 4.6076 | 4.2462 | | 1.7833 | 5.1590 | 4.8409 | 2.5361 | 3.1144 | 2.5518 | 7.1700 | 6.5925 | 6.8301 | H13 | 2.6725 | 2.1893 | 4.0251 | 1.1061 | 4.9935 | 2.1960 | 6.4234 | 3.1127 | 2.5473 | 4.2462 | 4.6076 | 1.7833 | | 4.8409 | 5.1590 | 2.5361 | 2.5518 | 3.1144 | 7.1700 | 6.8301 | 6.5925 | H14 | 2.6725 | 4.0251 | 2.1893 | 4.9935 | 1.1061 | 6.4234 | 2.1960 | 4.6076 | 4.2462 | 2.5473 | 3.1127 | 5.1590 | 4.8409 | | 1.7833 | 7.1700 | 6.5925 | 6.8301 | 2.5361 | 3.1144 | 2.5518 | H15 | 2.6725 | 4.0251 | 2.1893 | 4.9935 | 1.1061 | 6.4234 | 2.1960 | 4.2462 | 4.6076 | 3.1127 | 2.5473 | 4.8409 | 5.1590 | 1.7833 | | 7.1700 | 6.8301 | 6.5925 | 2.5361 | 2.5518 | 3.1144 | H16 | 4.6473 | 3.5450 | 5.8705 | 2.2034 | 7.0967 | 1.1033 | 8.4351 | 3.8342 | 3.8342 | 6.0705 | 6.0705 | 2.5361 | 2.5361 | 7.1700 | 7.1700 | | 1.7880 | 1.7880 | 9.2910 | 8.6089 | 8.6089 | H17 | 4.1748 | 2.8430 | 5.1059 | 2.2058 | 6.5002 | 1.1035 | 7.6597 | 3.1787 | 2.6261 | 5.0094 | 5.3199 | 3.1144 | 2.5518 | 6.5925 | 6.8301 | 1.7880 | | 1.7877 | 8.6089 | 7.8394 | 7.6329 | H18 | 4.1748 | 2.8430 | 5.1059 | 2.2058 | 6.5002 | 1.1035 | 7.6597 | 2.6261 | 3.1787 | 5.3199 | 5.0094 | 2.5518 | 3.1144 | 6.8301 | 6.5925 | 1.7880 | 1.7877 | | 8.6089 | 7.6329 | 7.8394 | H19 | 4.6473 | 5.8705 | 3.5450 | 7.0967 | 2.2034 | 8.4351 | 1.1033 | 6.0705 | 6.0705 | 3.8342 | 3.8342 | 7.1700 | 7.1700 | 2.5361 | 2.5361 | 9.2910 | 8.6089 | 8.6089 | | 1.7880 | 1.7880 | H20 | 4.1748 | 5.1059 | 2.8430 | 6.5002 | 2.2058 | 7.6597 | 1.1035 | 5.0094 | 5.3199 | 3.1787 | 2.6261 | 6.5925 | 6.8301 | 3.1144 | 2.5518 | 8.6089 | 7.8394 | 7.6329 | 1.7880 | | 1.7877 | H21 | 4.1748 | 5.1059 | 2.8430 | 6.5002 | 2.2058 | 7.6597 | 1.1035 | 5.3199 | 5.0094 | 2.6261 | 3.1787 | 6.8301 | 6.5925 | 2.5518 | 3.1144 | 8.6089 | 7.6329 | 7.8394 | 1.7880 | 1.7877 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
107.935 |
|
O1 |
C2 |
H8 |
111.276 |
| O1 |
C2 |
H9 |
111.276 |
|
O1 |
C3 |
C5 |
107.935 |
| O1 |
C3 |
H10 |
111.276 |
|
O1 |
C3 |
H11 |
111.276 |
| C2 |
O1 |
C3 |
107.260 |
|
C2 |
C4 |
C6 |
111.809 |
| C2 |
C4 |
H12 |
108.902 |
|
C2 |
C4 |
H13 |
108.902 |
| C3 |
C5 |
C7 |
111.809 |
|
C3 |
C5 |
H14 |
108.902 |
| C3 |
C5 |
H15 |
108.902 |
|
C4 |
C2 |
H8 |
109.718 |
| C4 |
C2 |
H9 |
109.718 |
|
C4 |
C6 |
H16 |
110.573 |
| C4 |
C6 |
H17 |
110.756 |
|
C4 |
C6 |
H18 |
110.756 |
| C5 |
C3 |
H10 |
109.718 |
|
C5 |
C3 |
H11 |
109.718 |
| C5 |
C7 |
H19 |
110.573 |
|
C5 |
C7 |
H20 |
110.756 |
| C5 |
C7 |
H21 |
110.756 |
|
C6 |
C4 |
H12 |
109.839 |
| C6 |
C4 |
H13 |
109.839 |
|
C7 |
C5 |
H14 |
109.839 |
| C7 |
C5 |
H15 |
109.839 |
|
H8 |
C2 |
H9 |
106.916 |
| H10 |
C3 |
H11 |
106.916 |
|
H12 |
C4 |
H13 |
107.442 |
| H14 |
C5 |
H15 |
107.442 |
|
H16 |
C6 |
H17 |
108.231 |
| H16 |
C6 |
H18 |
108.231 |
|
H17 |
C6 |
H18 |
108.199 |
| H19 |
C7 |
H20 |
108.231 |
|
H19 |
C7 |
H21 |
108.231 |
| H20 |
C7 |
H21 |
108.199 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -306.815113 |
| Energy at 298.15K | -306.830499 |
| HF Energy | -306.455027 |
| Nuclear repulsion energy | 318.544600 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3561 |
3165 |
0.72 |
|
|
|
| 2 |
A |
3554 |
3159 |
4.27 |
|
|
|
| 3 |
A |
3522 |
3130 |
2.76 |
|
|
|
| 4 |
A |
3426 |
3045 |
0.01 |
|
|
|
| 5 |
A |
3408 |
3029 |
0.17 |
|
|
|
| 6 |
A |
3376 |
3001 |
1.31 |
|
|
|
| 7 |
A |
3316 |
2948 |
22.73 |
|
|
|
| 8 |
A |
1743 |
1549 |
0.48 |
|
|
|
| 9 |
A |
1737 |
1544 |
0.01 |
|
|
|
| 10 |
A |
1732 |
1540 |
2.64 |
|
|
|
| 11 |
A |
1706 |
1517 |
0.01 |
|
|
|
| 12 |
A |
1622 |
1442 |
0.05 |
|
|
|
| 13 |
A |
1594 |
1417 |
1.15 |
|
|
|
| 14 |
A |
1546 |
1374 |
8.86 |
|
|
|
| 15 |
A |
1445 |
1285 |
0.75 |
|
|
|
| 16 |
A |
1384 |
1230 |
0.01 |
|
|
|
| 17 |
A |
1294 |
1150 |
2.64 |
|
|
|
| 18 |
A |
1246 |
1108 |
1.35 |
|
|
|
| 19 |
A |
1198 |
1064 |
0.90 |
|
|
|
| 20 |
A |
1061 |
943 |
0.49 |
|
|
|
| 21 |
A |
1020 |
906 |
2.32 |
|
|
|
| 22 |
A |
996 |
885 |
0.00 |
|
|
|
| 23 |
A |
831 |
739 |
0.76 |
|
|
|
| 24 |
A |
515 |
458 |
0.00 |
|
|
|
| 25 |
A |
347 |
308 |
0.15 |
|
|
|
| 26 |
A |
261 |
232 |
0.15 |
|
|
|
| 27 |
A |
166 |
148 |
0.05 |
|
|
|
| 28 |
A |
110 |
98 |
0.02 |
|
|
|
| 29 |
A |
51 |
45 |
0.00 |
|
|
|
| 30 |
B |
3561 |
3165 |
4.69 |
|
|
|
| 31 |
B |
3554 |
3159 |
3.90 |
|
|
|
| 32 |
B |
3522 |
3130 |
3.14 |
|
|
|
| 33 |
B |
3422 |
3042 |
38.35 |
|
|
|
| 34 |
B |
3407 |
3029 |
3.87 |
|
|
|
| 35 |
B |
3376 |
3001 |
5.31 |
|
|
|
| 36 |
B |
3318 |
2950 |
5.89 |
|
|
|
| 37 |
B |
1742 |
1549 |
0.58 |
|
|
|
| 38 |
B |
1733 |
1540 |
1.17 |
|
|
|
| 39 |
B |
1730 |
1538 |
6.59 |
|
|
|
| 40 |
B |
1706 |
1517 |
0.21 |
|
|
|
| 41 |
B |
1621 |
1441 |
0.87 |
|
|
|
| 42 |
B |
1560 |
1387 |
10.75 |
|
|
|
| 43 |
B |
1528 |
1358 |
85.99 |
|
|
|
| 44 |
B |
1436 |
1276 |
1.51 |
|
|
|
| 45 |
B |
1379 |
1225 |
5.54 |
|
|
|
| 46 |
B |
1289 |
1146 |
5.29 |
|
|
|
| 47 |
B |
1271 |
1130 |
5.59 |
|
|
|
| 48 |
B |
1218 |
1083 |
9.77 |
|
|
|
| 49 |
B |
1132 |
1006 |
5.89 |
|
|
|
| 50 |
B |
1032 |
917 |
1.76 |
|
|
|
| 51 |
B |
966 |
859 |
0.53 |
|
|
|
| 52 |
B |
862 |
767 |
2.64 |
|
|
|
| 53 |
B |
508 |
452 |
0.49 |
|
|
|
| 54 |
B |
321 |
285 |
0.49 |
|
|
|
| 55 |
B |
209 |
186 |
0.51 |
|
|
|
| 56 |
B |
151 |
135 |
4.13 |
|
|
|
| 57 |
B |
79 |
70 |
0.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 47698.5 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 42399.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.035 |
| C2 |
0.003 |
1.178 |
0.902 |
| C3 |
-0.003 |
-1.178 |
0.902 |
| C4 |
0.000 |
2.452 |
-0.008 |
| C5 |
0.000 |
-2.452 |
-0.008 |
| C6 |
-1.284 |
2.544 |
-0.886 |
| C7 |
1.284 |
-2.544 |
-0.886 |
| H8 |
-0.892 |
1.191 |
1.571 |
| H9 |
0.904 |
1.191 |
1.562 |
| H10 |
0.892 |
-1.191 |
1.571 |
| H11 |
-0.904 |
-1.191 |
1.562 |
| H12 |
0.894 |
2.427 |
-0.658 |
| H13 |
0.081 |
3.347 |
0.638 |
| H14 |
-0.894 |
-2.427 |
-0.658 |
| H15 |
-0.081 |
-3.347 |
0.638 |
| H16 |
-1.246 |
3.434 |
-1.537 |
| H17 |
-1.377 |
1.648 |
-1.522 |
| H18 |
-2.185 |
2.615 |
-0.253 |
| H19 |
1.246 |
-3.434 |
-1.537 |
| H20 |
1.377 |
-1.648 |
-1.522 |
| H21 |
2.185 |
-2.615 |
-0.253 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
C7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
H17 |
H18 |
H19 |
H20 |
H21 |
| O1 | | 1.4630 | 1.4630 | 2.4522 | 2.4522 | 2.9947 | 2.9947 | 2.1380 | 2.1372 | 2.1380 | 2.1372 | 2.6783 | 3.4015 | 2.6783 | 3.4015 | 3.9771 | 2.6530 | 3.4199 | 3.9771 | 2.6530 | 3.4199 |
C2 | 1.4630 | | 2.3567 | 1.5653 | 3.7425 | 2.5919 | 4.3236 | 1.1168 | 1.1168 | 2.6171 | 2.6215 | 2.1886 | 2.1857 | 4.0301 | 4.5335 | 3.5495 | 2.8291 | 2.8610 | 5.3637 | 3.9693 | 4.5261 | C3 | 1.4630 | 2.3567 | | 3.7425 | 1.5653 | 4.3236 | 2.5919 | 2.6171 | 2.6215 | 1.1168 | 1.1168 | 4.0301 | 4.5335 | 2.1886 | 2.1857 | 5.3637 | 3.9693 | 4.5261 | 3.5495 | 2.8291 | 2.8610 | C4 | 2.4522 | 1.5653 | 3.7425 | | 4.9037 | 1.5579 | 5.2324 | 2.2084 | 2.2070 | 4.0689 | 4.0686 | 1.1062 | 1.1066 | 5.0030 | 5.8349 | 2.2038 | 2.1991 | 2.2045 | 6.2076 | 4.5826 | 5.5234 | C5 | 2.4522 | 3.7425 | 1.5653 | 4.9037 | | 5.2324 | 1.5579 | 4.0689 | 4.0686 | 2.2084 | 2.2070 | 5.0030 | 5.8349 | 1.1062 | 1.1066 | 6.2076 | 4.5826 | 5.5234 | 2.2038 | 2.1991 | 2.2045 | C6 | 2.9947 | 2.5919 | 4.3236 | 1.5579 | 5.2324 | | 5.6992 | 2.8319 | 3.5508 | 4.9715 | 4.4820 | 2.1932 | 2.1973 | 4.9919 | 6.2024 | 1.1033 | 1.1029 | 1.1034 | 6.5240 | 5.0061 | 6.2489 | C7 | 2.9947 | 4.3236 | 2.5919 | 5.2324 | 1.5579 | 5.6992 | | 4.9715 | 4.4820 | 2.8319 | 3.5508 | 4.9919 | 6.2024 | 2.1932 | 2.1973 | 6.5240 | 5.0061 | 6.2489 | 1.1033 | 1.1029 | 1.1034 | H8 | 2.1380 | 1.1168 | 2.6171 | 2.2084 | 4.0689 | 2.8319 | 4.9715 | | 1.7955 | 2.9752 | 2.3820 | 3.1124 | 2.5422 | 4.2496 | 4.7027 | 3.8491 | 3.1640 | 2.6508 | 5.9680 | 4.7721 | 5.2226 | H9 | 2.1372 | 1.1168 | 2.6215 | 2.2070 | 4.0686 | 3.5508 | 4.4820 | 1.7955 | | 2.3820 | 2.9906 | 2.5413 | 2.4854 | 4.6107 | 4.7345 | 4.3885 | 3.8633 | 3.8551 | 5.5780 | 4.2188 | 4.4072 | H10 | 2.1380 | 2.6171 | 1.1168 | 4.0689 | 2.2084 | 4.9715 | 2.8319 | 2.9752 | 2.3820 | | 1.7955 | 4.2496 | 4.7027 | 3.1124 | 2.5422 | 5.9680 | 4.7721 | 5.2226 | 3.8491 | 3.1640 | 2.6508 | H11 | 2.1372 | 2.6215 | 1.1168 | 4.0686 | 2.2070 | 4.4820 | 3.5508 | 2.3820 | 2.9906 | 1.7955 | | 4.6107 | 4.7345 | 2.5413 | 2.4854 | 5.5780 | 4.2188 | 4.4072 | 4.3885 | 3.8633 | 3.8551 | H12 | 2.6783 | 2.1886 | 4.0301 | 1.1062 | 5.0030 | 2.1932 | 4.9919 | 3.1124 | 2.5413 | 4.2496 | 4.6107 | | 1.7854 | 5.1740 | 5.9976 | 2.5237 | 2.5524 | 3.1115 | 5.9373 | 4.1940 | 5.2209 | H13 | 3.4015 | 2.1857 | 4.5335 | 1.1066 | 5.8349 | 2.1973 | 6.2024 | 2.5422 | 2.4854 | 4.7027 | 4.7345 | 1.7854 | | 5.9976 | 6.6952 | 2.5497 | 3.1109 | 2.5420 | 7.2157 | 5.5943 | 6.3847 | H14 | 2.6783 | 4.0301 | 2.1886 | 5.0030 | 1.1062 | 4.9919 | 2.1932 | 4.2496 | 4.6107 | 3.1124 | 2.5413 | 5.1740 | 5.9976 | | 1.7854 | 5.9373 | 4.1940 | 5.2209 | 2.5237 | 2.5524 | 3.1115 | H15 | 3.4015 | 4.5335 | 2.1857 | 5.8349 | 1.1066 | 6.2024 | 2.1973 | 4.7027 | 4.7345 | 2.5422 | 2.4854 | 5.9976 | 6.6952 | 1.7854 | | 7.2157 | 5.5943 | 6.3847 | 2.5497 | 3.1109 | 2.5420 | H16 | 3.9771 | 3.5495 | 5.3637 | 2.2038 | 6.2076 | 1.1033 | 6.5240 | 3.8491 | 4.3885 | 5.9680 | 5.5780 | 2.5237 | 2.5497 | 5.9373 | 7.2157 | | 1.7906 | 1.7888 | 7.3062 | 5.7194 | 7.0721 | H17 | 2.6530 | 2.8291 | 3.9693 | 2.1991 | 4.5826 | 1.1029 | 5.0061 | 3.1640 | 3.8633 | 4.7721 | 4.2188 | 2.5524 | 3.1109 | 4.1940 | 5.5943 | 1.7906 | | 1.7883 | 5.7194 | 4.2958 | 5.6988 | H18 | 3.4199 | 2.8610 | 4.5261 | 2.2045 | 5.5234 | 1.1034 | 6.2489 | 2.6508 | 3.8551 | 5.2226 | 4.4072 | 3.1115 | 2.5420 | 5.2209 | 6.3847 | 1.7888 | 1.7883 | | 7.0721 | 5.6988 | 6.8154 | H19 | 3.9771 | 5.3637 | 3.5495 | 6.2076 | 2.2038 | 6.5240 | 1.1033 | 5.9680 | 5.5780 | 3.8491 | 4.3885 | 5.9373 | 7.2157 | 2.5237 | 2.5497 | 7.3062 | 5.7194 | 7.0721 | | 1.7906 | 1.7888 | H20 | 2.6530 | 3.9693 | 2.8291 | 4.5826 | 2.1991 | 5.0061 | 1.1029 | 4.7721 | 4.2188 | 3.1640 | 3.8633 | 4.1940 | 5.5943 | 2.5524 | 3.1109 | 5.7194 | 4.2958 | 5.6988 | 1.7906 | | 1.7883 | H21 | 3.4199 | 4.5261 | 2.8610 | 5.5234 | 2.2045 | 6.2489 | 1.1034 | 5.2226 | 4.4072 | 2.6508 | 3.8551 | 5.2209 | 6.3847 | 3.1115 | 2.5420 | 7.0721 | 5.6988 | 6.8154 | 1.7888 | 1.7883 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C4 |
108.100 |
|
O1 |
C2 |
H8 |
111.232 |
| O1 |
C2 |
H9 |
111.167 |
|
O1 |
C3 |
C5 |
108.100 |
| O1 |
C3 |
H10 |
111.232 |
|
O1 |
C3 |
H11 |
111.167 |
| C2 |
O1 |
C3 |
107.302 |
|
C2 |
C4 |
C6 |
112.171 |
| C2 |
C4 |
H12 |
108.804 |
|
C2 |
C4 |
H13 |
108.563 |
| C3 |
C5 |
C7 |
112.171 |
|
C3 |
C5 |
H14 |
108.804 |
| C3 |
C5 |
H15 |
108.563 |
|
C4 |
C2 |
H8 |
109.719 |
| C4 |
C2 |
H9 |
109.614 |
|
C4 |
C6 |
H16 |
110.646 |
| C4 |
C6 |
H17 |
110.303 |
|
C4 |
C6 |
H18 |
110.701 |
| C5 |
C3 |
H10 |
109.719 |
|
C5 |
C3 |
H11 |
109.614 |
| C5 |
C7 |
H19 |
110.646 |
|
C5 |
C7 |
H20 |
110.303 |
| C5 |
C7 |
H21 |
110.701 |
|
C6 |
C4 |
H12 |
109.652 |
| C6 |
C4 |
H13 |
109.953 |
|
C7 |
C5 |
H14 |
109.652 |
| C7 |
C5 |
H15 |
109.953 |
|
H8 |
C2 |
H9 |
107.001 |
| H10 |
C3 |
H11 |
107.001 |
|
H12 |
C4 |
H13 |
107.579 |
| H14 |
C5 |
H15 |
107.579 |
|
H16 |
C6 |
H17 |
108.503 |
| H16 |
C6 |
H18 |
108.309 |
|
H17 |
C6 |
H18 |
108.302 |
| H19 |
C7 |
H20 |
108.503 |
|
H19 |
C7 |
H21 |
108.309 |
| H20 |
C7 |
H21 |
108.302 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
