Jump to
S2C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -1795.910004 |
| Energy at 298.15K | -1795.911094 |
| Nuclear repulsion energy | 74.173467 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3377 |
3002 |
1.04 |
|
|
|
| 2 |
A1 |
1663 |
1478 |
5.42 |
|
|
|
| 3 |
A1 |
724 |
644 |
95.77 |
|
|
|
| 4 |
B1 |
476 |
423 |
7.74 |
|
|
|
| 5 |
B2 |
3505 |
3116 |
1.11 |
|
|
|
| 6 |
B2 |
364 |
324 |
6.70 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5054.6 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 4493.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.395 |
| C2 |
0.000 |
0.000 |
-1.320 |
| H3 |
0.000 |
0.890 |
-1.969 |
| H4 |
0.000 |
-0.890 |
-1.969 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.7148 | 2.5266 | 2.5266 |
C2 | 1.7148 | | 1.1021 | 1.1021 | H3 | 2.5266 | 1.1021 | | 1.7805 | H4 | 2.5266 | 1.1021 | 1.7805 | |
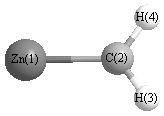 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Zn1 |
C2 |
H3 |
126.123 |
|
Zn1 |
C2 |
H4 |
126.123 |
| H3 |
C2 |
H4 |
107.753 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -1795.884625 |
| Energy at 298.15K | -1795.885991 |
| Nuclear repulsion energy | 73.814943 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3360 |
2987 |
18.63 |
|
|
|
| 2 |
A1 |
1579 |
1403 |
3.20 |
|
|
|
| 3 |
A1 |
793 |
705 |
14.08 |
|
|
|
| 4 |
B1 |
866 |
770 |
34.84 |
|
|
|
| 5 |
B2 |
3490 |
3102 |
0.01 |
|
|
|
| 6 |
B2 |
403 |
358 |
2.54 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5245.6 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 4662.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Zn1 |
0.000 |
0.000 |
0.396 |
| C2 |
0.000 |
0.000 |
-1.330 |
| H3 |
0.000 |
0.907 |
-1.958 |
| H4 |
0.000 |
-0.907 |
-1.958 |
Atom - Atom Distances (Å)
| |
Zn1 |
C2 |
H3 |
H4 |
| Zn1 | | 1.7260 | 2.5234 | 2.5234 |
C2 | 1.7260 | | 1.1036 | 1.1036 | H3 | 2.5234 | 1.1036 | | 1.8140 | H4 | 2.5234 | 1.1036 | 1.8140 | |
More geometry information
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
