return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C7H8 (Norbornadiene)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-266.826704
Energy at 298.15K-266.835891
Nuclear repulsion energy283.483295
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3558 3163 0.02      
2 A1 3483 3096 0.94      
3 A1 3404 3026 3.31      
4 A1 1728 1536 0.09      
5 A1 1694 1506 1.36      
6 A1 1413 1256 0.91      
7 A1 1214 1079 0.02      
8 A1 1030 915 1.17      
9 A1 1007 896 0.09      
10 A1 847 753 0.39      
11 A1 802 713 18.87      
12 A1 454 404 0.39      
13 A2 3526 3134 0.00      
14 A2 1469 1306 0.00      
15 A2 1382 1228 0.00      
16 A2 1240 1103 0.00      
17 A2 1029 915 0.00      
18 A2 990 880 0.00      
19 A2 762 677 0.00      
20 A2 474 421 0.00      
21 B1 3556 3161 0.00      
22 B1 3536 3143 6.17      
23 B1 1690 1503 1.56      
24 B1 1376 1223 2.39      
25 B1 1172 1042 0.58      
26 B1 1127 1002 0.45      
27 B1 1022 909 0.01      
28 B1 737 655 11.53      
29 B1 517 459 1.75      
30 B2 3526 3134 8.45      
31 B2 3480 3093 4.99      
32 B2 1496 1330 3.43      
33 B2 1406 1250 0.02      
34 B2 1341 1192 0.02      
35 B2 1082 961 0.19      
36 B2 998 887 1.39      
37 B2 981 872 0.01      
38 B2 896 796 0.43      
39 B2 594 528 0.57      

Unscaled Zero Point Vibrational Energy (zpe) 31018.6 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 27572.5 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.13856 0.11690 0.10339

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 1.376
C2 0.000 1.138 0.282
C3 0.000 -1.138 0.282
C4 1.256 0.678 -0.532
C5 1.256 -0.678 -0.532
C6 -1.256 0.678 -0.532
C7 -1.256 -0.678 -0.532
H8 -0.907 0.000 2.009
H9 0.907 0.000 2.009
H10 0.000 2.190 0.625
H11 0.000 -2.190 0.625
H12 1.954 1.359 -1.036
H13 1.954 -1.359 -1.036
H14 -1.954 1.359 -1.036
H15 -1.954 -1.359 -1.036

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 C7 H8 H9 H10 H11 H12 H13 H14 H15
C11.57881.57882.38212.38212.38212.38211.10641.10642.31492.31493.38843.38843.38843.3884
C21.57882.27701.56582.35321.56582.35322.25882.25881.10583.34592.36733.43392.36733.4339
C31.57882.27702.35321.56582.35321.56582.25882.25883.34591.10583.43392.36733.43392.3673
C42.38211.56582.35321.35522.51222.85443.40492.65232.28073.33721.09812.21103.32043.8351
C52.38212.35321.56581.35522.85442.51223.40492.65233.33722.28072.21101.09813.83513.3204
C62.38211.56582.35322.51222.85441.35522.65233.40492.28073.33723.32043.83511.09812.2110
C72.38212.35321.56582.85442.51221.35522.65233.40493.33722.28073.83513.32042.21101.0981
H81.10642.25882.25883.40493.40492.65232.65231.81472.74482.74484.39384.39383.49473.4947
H91.10642.25882.25882.65232.65233.40493.40491.81472.74482.74483.49473.49474.39384.3938
H102.31491.10583.34592.28073.33722.28073.33722.74482.74484.37962.69594.37822.69594.3782
H112.31493.34591.10583.33722.28073.33722.28072.74482.74484.37964.37822.69594.37822.6959
H123.38842.36733.43391.09812.21103.32043.83514.39383.49472.69594.37822.71743.90874.7605
H133.38843.43392.36732.21101.09813.83513.32044.39383.49474.37822.69592.71744.76053.9087
H143.38842.36733.43393.32043.83511.09812.21103.49474.39382.69594.37823.90874.76052.7174
H153.38843.43392.36733.83513.32042.21101.09813.49474.39384.37822.69594.76053.90872.7174

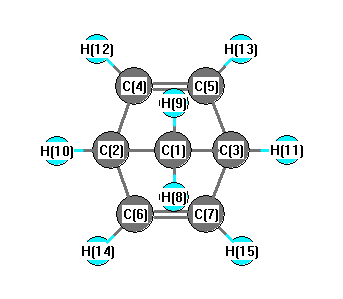
picture of Norbornadiene state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 98.491 C1 C2 C6 98.491
C1 C2 H10 118.076 C1 C3 C5 98.491
C1 C3 C7 98.491 C1 C3 H11 118.076
C2 C1 C3 92.290 C2 C1 H8 113.360
C2 C1 H9 113.360 C2 C4 C5 107.118
C2 C4 H12 124.479 C2 C6 C7 107.118
C2 C6 H14 124.479 C3 C1 H8 113.360
C3 C1 H9 113.360 C3 C5 C4 107.118
C3 C5 H13 124.479 C3 C7 C6 107.118
C3 C7 H15 124.479 C4 C2 C6 106.681
C4 C2 H10 116.164 C4 C5 H13 128.336
C5 C3 C7 106.681 C5 C3 H11 116.164
C5 C4 H12 128.336 C6 C2 H10 116.164
C6 C7 H15 128.336 C7 C3 H11 116.164
C7 C6 H14 128.336 H8 C1 H9 110.179
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability