Jump to
S1C2
S1C3
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -205.488698 |
| Energy at 298.15K | |
| HF Energy | -205.262859 |
| Nuclear repulsion energy | 115.677899 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3868 |
3439 |
15.72 |
|
|
|
| 2 |
A' |
3535 |
3143 |
1.86 |
|
|
|
| 3 |
A' |
3338 |
2967 |
17.59 |
|
|
|
| 4 |
A' |
3301 |
2934 |
31.63 |
|
|
|
| 5 |
A' |
1879 |
1670 |
152.75 |
|
|
|
| 6 |
A' |
1737 |
1544 |
6.59 |
|
|
|
| 7 |
A' |
1638 |
1456 |
24.12 |
|
|
|
| 8 |
A' |
1590 |
1414 |
1.45 |
|
|
|
| 9 |
A' |
1493 |
1327 |
0.38 |
|
|
|
| 10 |
A' |
1337 |
1189 |
99.90 |
|
|
|
| 11 |
A' |
1265 |
1125 |
17.25 |
|
|
|
| 12 |
A' |
1076 |
957 |
40.84 |
|
|
|
| 13 |
A' |
603 |
536 |
4.81 |
|
|
|
| 14 |
A' |
335 |
298 |
3.30 |
|
|
|
| 15 |
A" |
3491 |
3103 |
7.33 |
|
|
|
| 16 |
A" |
1701 |
1512 |
2.78 |
|
|
|
| 17 |
A" |
1242 |
1104 |
4.26 |
|
|
|
| 18 |
A" |
994 |
884 |
0.93 |
|
|
|
| 19 |
A" |
409 |
364 |
89.41 |
|
|
|
| 20 |
A" |
170 |
151 |
3.24 |
|
|
|
| 21 |
A" |
239i |
212i |
43.85 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17381.9 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15450.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.283 |
-0.795 |
0.000 |
| O2 |
1.440 |
-1.287 |
0.000 |
| N3 |
0.000 |
0.600 |
0.000 |
| C4 |
-1.360 |
1.174 |
0.000 |
| H5 |
-0.676 |
-1.381 |
0.000 |
| H6 |
0.804 |
1.256 |
0.000 |
| H7 |
-2.091 |
0.343 |
0.000 |
| H8 |
-1.547 |
1.800 |
0.898 |
| H9 |
-1.547 |
1.800 |
-0.898 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2576 | 1.4230 | 2.5636 | 1.1232 | 2.1162 | 2.6318 | 3.2992 | 3.2992 |
O2 | 1.2576 | | 2.3735 | 3.7273 | 2.1177 | 2.6214 | 3.8885 | 4.3879 | 4.3879 | N3 | 1.4230 | 2.3735 | | 1.4760 | 2.0927 | 1.0378 | 2.1063 | 2.1537 | 2.1537 | C4 | 2.5636 | 3.7273 | 1.4760 | | 2.6443 | 2.1655 | 1.1062 | 1.1109 | 1.1109 | H5 | 1.1232 | 2.1177 | 2.0927 | 2.6443 | | 3.0237 | 2.2300 | 3.4178 | 3.4178 | H6 | 2.1162 | 2.6214 | 1.0378 | 2.1655 | 3.0237 | | 3.0352 | 2.5744 | 2.5744 | H7 | 2.6318 | 3.8885 | 2.1063 | 1.1062 | 2.2300 | 3.0352 | | 1.7960 | 1.7960 | H8 | 3.2992 | 4.3879 | 2.1537 | 1.1109 | 3.4178 | 2.5744 | 1.7960 | | 1.7967 | H9 | 3.2992 | 4.3879 | 2.1537 | 1.1109 | 3.4178 | 2.5744 | 1.7960 | 1.7967 | |
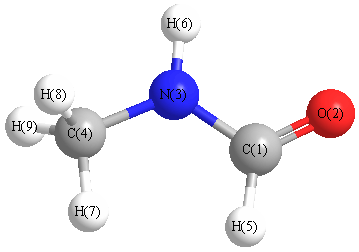 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.322 |
|
C1 |
N3 |
H6 |
117.770 |
| O2 |
C1 |
N3 |
124.489 |
|
O2 |
C1 |
H5 |
125.520 |
| N3 |
C1 |
H5 |
109.991 |
|
N3 |
C4 |
H7 |
108.467 |
| N3 |
C4 |
H8 |
111.952 |
|
N3 |
C4 |
H9 |
111.952 |
| C4 |
N3 |
H6 |
117.908 |
|
H7 |
C4 |
H8 |
108.204 |
| H7 |
C4 |
H9 |
108.204 |
|
H8 |
C4 |
H9 |
107.933 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -205.488698 |
| Energy at 298.15K | |
| HF Energy | -205.262859 |
| Nuclear repulsion energy | 115.677899 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at MP2=FULL/STO-3G
| | hartrees |
|---|
| Energy at 0K | -205.493235 |
| Energy at 298.15K | -205.499161 |
| HF Energy | -205.264685 |
| Nuclear repulsion energy | 117.566352 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3737 |
3322 |
1.94 |
|
|
|
| 2 |
A |
3555 |
3160 |
0.34 |
|
|
|
| 3 |
A |
3513 |
3123 |
5.76 |
|
|
|
| 4 |
A |
3348 |
2976 |
8.50 |
|
|
|
| 5 |
A |
3311 |
2943 |
34.06 |
|
|
|
| 6 |
A |
1820 |
1618 |
64.35 |
|
|
|
| 7 |
A |
1723 |
1531 |
0.77 |
|
|
|
| 8 |
A |
1706 |
1516 |
8.23 |
|
|
|
| 9 |
A |
1615 |
1436 |
21.44 |
|
|
|
| 10 |
A |
1604 |
1426 |
9.70 |
|
|
|
| 11 |
A |
1494 |
1328 |
7.15 |
|
|
|
| 12 |
A |
1273 |
1132 |
58.79 |
|
|
|
| 13 |
A |
1228 |
1092 |
21.64 |
|
|
|
| 14 |
A |
1209 |
1075 |
3.42 |
|
|
|
| 15 |
A |
1031 |
916 |
12.60 |
|
|
|
| 16 |
A |
950 |
844 |
2.13 |
|
|
|
| 17 |
A |
810 |
720 |
64.37 |
|
|
|
| 18 |
A |
591 |
525 |
89.52 |
|
|
|
| 19 |
A |
309 |
275 |
8.33 |
|
|
|
| 20 |
A |
243 |
216 |
7.41 |
|
|
|
| 21 |
A |
122 |
109 |
0.39 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 17595.2 cm
-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15640.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/STO-3G
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.883 |
0.456 |
0.048 |
| O2 |
1.422 |
-0.678 |
-0.016 |
| N3 |
-0.540 |
0.715 |
-0.186 |
| C4 |
-1.423 |
-0.480 |
0.044 |
| H5 |
1.436 |
1.422 |
0.197 |
| H6 |
-0.850 |
1.511 |
0.426 |
| H7 |
-2.453 |
-0.238 |
-0.278 |
| H8 |
-1.043 |
-1.319 |
-0.569 |
| H9 |
-1.451 |
-0.812 |
1.104 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2569 | 1.4656 | 2.4890 | 1.1230 | 2.0640 | 3.4229 | 2.6909 | 2.8585 |
O2 | 1.2569 | | 2.4123 | 2.8526 | 2.1103 | 3.1857 | 3.9088 | 2.6070 | 3.0865 | N3 | 1.4656 | 2.4123 | | 1.5034 | 2.1339 | 1.0518 | 2.1384 | 2.1292 | 2.1963 | C4 | 2.4890 | 2.8526 | 1.5034 | | 3.4375 | 2.1076 | 1.1060 | 1.1055 | 1.1109 | H5 | 1.1230 | 2.1103 | 2.1339 | 3.4375 | | 2.2991 | 4.2548 | 3.7742 | 3.7612 | H6 | 2.0640 | 3.1857 | 1.0518 | 2.1076 | 2.2991 | | 2.4748 | 3.0061 | 2.4933 | H7 | 3.4229 | 3.9088 | 2.1384 | 1.1060 | 4.2548 | 2.4748 | | 1.8001 | 1.8005 | H8 | 2.6909 | 2.6070 | 2.1292 | 1.1055 | 3.7742 | 3.0061 | 1.8001 | | 1.7948 | H9 | 2.8585 | 3.0865 | 2.1963 | 1.1109 | 3.7612 | 2.4933 | 1.8005 | 1.7948 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
113.921 |
|
C1 |
N3 |
H6 |
109.043 |
| O2 |
C1 |
N3 |
124.590 |
|
O2 |
C1 |
H5 |
124.841 |
| N3 |
C1 |
H5 |
110.348 |
|
N3 |
C4 |
H7 |
109.126 |
| N3 |
C4 |
H8 |
108.438 |
|
N3 |
C4 |
H9 |
113.452 |
| C4 |
N3 |
H6 |
109.887 |
|
H7 |
C4 |
H8 |
108.974 |
| H7 |
C4 |
H9 |
108.618 |
|
H8 |
C4 |
H9 |
108.151 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
