return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CClCHCH2 (1,3-Butadiene, 2-chloro-)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-607.275899
Energy at 298.15K 
HF Energy-607.017996
Nuclear repulsion energy192.910810
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3624 3221 0.74 28.21 0.71 0.83
2 A' 3621 3219 2.39 39.59 0.75 0.86
3 A' 3510 3120 0.70 48.96 0.24 0.38
4 A' 3464 3079 8.35 43.83 0.14 0.25
5 A' 3460 3076 32.62 31.92 0.13 0.23
6 A' 1805 1605 3.10 18.16 0.60 0.75
7 A' 1739 1545 3.98 2.27 0.65 0.79
8 A' 1583 1407 0.39 33.64 0.49 0.66
9 A' 1524 1355 1.56 2.51 0.72 0.84
10 A' 1412 1255 3.67 10.06 0.64 0.78
11 A' 1357 1206 37.79 8.37 0.30 0.46
12 A' 1112 989 5.80 3.05 0.75 0.86
13 A' 969 862 3.70 2.68 0.47 0.64
14 A' 691 615 10.70 8.33 0.21 0.35
15 A' 536 476 0.83 2.80 0.69 0.82
16 A' 378 336 1.89 1.77 0.75 0.86
17 A' 236 210 0.64 1.42 0.70 0.82
18 A" 1117 993 6.35 0.10 0.75 0.86
19 A" 991 881 12.33 0.01 0.75 0.86
20 A" 947 842 19.00 0.03 0.75 0.86
21 A" 824 733 0.06 9.55 0.75 0.86
22 A" 714 635 2.63 2.11 0.75 0.86
23 A" 405 360 0.16 2.33 0.75 0.86
24 A" 119 106 0.15 1.93 0.75 0.86

Unscaled Zero Point Vibrational Energy (zpe) 18069.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 16062.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.17236 0.11852 0.07023

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.336 1.921 0.000
C2 0.000 0.605 0.000
C3 1.420 0.096 0.000
C4 1.787 -1.206 0.000
Cl5 -1.316 -0.636 0.000
H6 0.454 2.685 0.000
H7 -1.381 2.257 0.000
H8 2.183 0.891 0.000
H9 2.848 -1.494 0.000
H10 1.043 -2.014 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 Cl5 H6 H7 H8 H9 H10
C11.35742.53273.77922.73851.09891.09812.72084.66814.1691
C21.35741.50902.54461.80962.12812.15332.20143.53762.8194
C32.53271.50901.35232.83282.76343.53851.10212.13612.1430
C43.77922.54461.35233.15534.11254.69402.13431.09861.0984
Cl52.73851.80962.83283.15533.76332.89463.81814.25112.7321
H61.09892.12812.76344.11253.76331.88412.49084.81524.7352
H71.09812.15333.53854.69402.89461.88413.81675.65264.9114
H82.72082.20141.10212.13433.81812.49083.81672.47603.1208
H94.66813.53762.13611.09864.25114.81525.65262.47601.8779
H104.16912.81942.14301.09842.73214.73524.91143.12081.8779

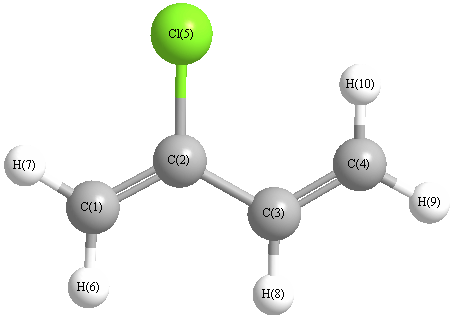
picture of 1,3-Butadiene, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.073 C1 C2 Cl5 119.007
C2 C1 H6 119.716 C2 C1 H7 122.195
C2 C3 C4 125.490 C2 C3 H8 114.031
C3 C2 Cl5 116.921 C3 C4 H9 120.930
C3 C4 H10 121.612 C4 C3 H8 120.479
H6 C1 H7 118.089 H9 C4 H10 117.458
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability