Jump to
S1C2
Energy calculated at M06-2X/STO-3G
| | hartrees |
|---|
| Energy at 0K | -394.526391 |
| Energy at 298.15K | |
| HF Energy | -394.526391 |
| Nuclear repulsion energy | 70.264868 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at M06-2X/STO-3G
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.000 |
| Na2 |
0.000 |
0.000 |
1.781 |
| Na3 |
0.000 |
0.000 |
-1.781 |
Atom - Atom Distances (Å)
| |
O1 |
Na2 |
Na3 |
| O1 | | 1.7811 | 1.7811 |
Na2 | 1.7811 | | 3.5622 | Na3 | 1.7811 | 3.5622 | |
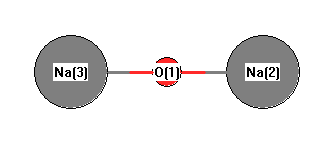 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
O1 |
Na3 |
180.000 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.752 |
|
|
|
| 2 |
Na |
0.376 |
|
|
|
| 3 |
Na |
0.376 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.656 |
0.000 |
0.000 |
| y |
0.000 |
-13.656 |
0.000 |
| z |
0.000 |
0.000 |
5.943 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.799 |
0.000 |
0.000 |
| y |
0.000 |
-9.799 |
0.000 |
| z |
0.000 |
0.000 |
19.599 |
|
| Polar |
|---|
| 3z2-r2 | 39.197 |
|---|
| x2-y2 | 0.000 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.903 |
0.000 |
0.000 |
| y |
0.000 |
0.903 |
0.000 |
| z |
0.000 |
0.000 |
7.528 |
<r2> (average value of r
2) Å
2
| <r2> |
74.242 |
| (<r2>)1/2 |
8.616 |
Jump to
S1C1
Energy calculated at M06-2X/STO-3G
| | hartrees |
|---|
| Energy at 0K | -394.548531 |
| Energy at 298.15K | -394.550534 |
| HF Energy | -394.548531 |
| Nuclear repulsion energy | 74.411092 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Geometric Data calculated at M06-2X/STO-3G
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
0.915 |
| Na2 |
0.000 |
1.349 |
-0.333 |
| Na3 |
0.000 |
-1.349 |
-0.333 |
Atom - Atom Distances (Å)
| |
O1 |
Na2 |
Na3 |
| O1 | | 1.8379 | 1.8379 |
Na2 | 1.8379 | | 2.6977 | Na3 | 1.8379 | 2.6977 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Na2 |
O1 |
Na3 |
94.433 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at M06-2X/STO-3G
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.645 |
|
|
|
| 2 |
Na |
0.322 |
|
|
|
| 3 |
Na |
0.322 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-5.487 |
5.487 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-13.907 |
0.000 |
0.000 |
| y |
0.000 |
-2.685 |
0.000 |
| z |
0.000 |
0.000 |
-16.012 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.558 |
0.000 |
0.000 |
| y |
0.000 |
12.274 |
0.000 |
| z |
0.000 |
0.000 |
-7.716 |
|
| Polar |
|---|
| 3z2-r2 | -15.432 |
|---|
| x2-y2 | -11.221 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.974 |
0.000 |
0.000 |
| y |
0.000 |
4.939 |
0.000 |
| z |
0.000 |
0.000 |
2.271 |
<r2> (average value of r
2) Å
2
| <r2> |
55.957 |
| (<r2>)1/2 |
7.480 |
