return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H3F3 (Benzene trifluoride 123)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-520.662209
Energy at 298.15K-520.665628
HF Energy-520.253155
Nuclear repulsion energy421.460873
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3533 3140 7.42      
2 A1 3515 3124 0.01      
3 A1 1731 1539 4.69      
4 A1 1615 1436 71.78      
5 A1 1469 1305 2.02      
6 A1 1371 1219 11.76      
7 A1 1141 1015 0.38      
8 A1 884 786 0.72      
9 A1 715 636 2.74      
10 A1 485 431 0.04      
11 A1 290 258 1.44      
12 A2 964 857 0.00      
13 A2 593 527 0.00      
14 A2 260 231 0.00      
15 B1 1052 935 0.86      
16 B1 838 745 25.40      
17 B1 739 656 0.00      
18 B1 559 497 0.03      
19 B1 313 278 0.38      
20 B1 155 137 0.06      
21 B2 3524 3132 7.90      
22 B2 1719 1528 44.51      
23 B2 1613 1434 47.11      
24 B2 1384 1231 16.50      
25 B2 1268 1127 0.03      
26 B2 1201 1067 0.22      
27 B2 1089 968 13.73      
28 B2 583 518 2.86      
29 B2 509 452 0.17      
30 B2 247 220 0.14      

Unscaled Zero Point Vibrational Energy (zpe) 17677.9 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 15713.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.07362 0.05507 0.03150

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.751
C2 0.000 1.242 0.019
C3 0.000 -1.242 0.019
C4 0.000 1.231 -1.408
C5 0.000 -1.231 -1.408
C6 0.000 0.000 -2.107
F7 0.000 0.000 2.123
F8 0.000 2.426 0.711
F9 0.000 -2.426 0.711
H10 0.000 2.191 -1.945
H11 0.000 -2.191 -1.945
H12 0.000 0.000 -3.209

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 F7 F8 F9 H10 H11 H12
C11.44171.44172.48512.48512.85851.37182.42652.42653.47393.47393.9602
C21.44172.48411.42702.85502.46262.44311.37133.73282.18123.95503.4588
C31.44172.48412.85501.42702.46262.44313.73281.37133.95502.18123.4588
C42.48511.42702.85502.46151.41563.73912.43264.22631.10013.46342.1815
C52.48512.85501.42702.46151.41563.73914.22632.43263.46341.10012.1815
C62.85852.46262.46261.41561.41564.23033.71863.71862.19682.19681.1017
F71.37182.44312.44313.73913.73914.23032.80722.80724.62034.62035.3320
F82.42651.37133.73282.43264.22633.71862.80724.85232.66615.32624.6098
F92.42653.73281.37134.22632.43263.71862.80724.85235.32622.66614.6098
H103.47392.18123.95501.10013.46342.19684.62032.66615.32624.38162.5293
H113.47393.95502.18123.46341.10012.19684.62035.32622.66614.38162.5293
H123.96023.45883.45882.18152.18151.10175.33204.60984.60982.52932.5293

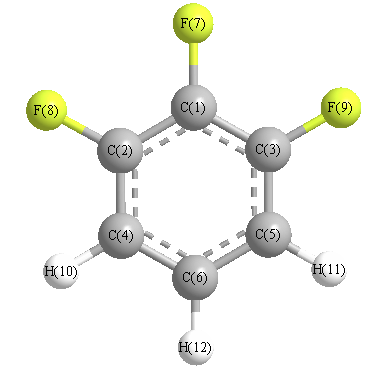
picture of Benzene trifluoride 123 state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 120.062 C1 C2 F8 119.197
C1 C3 C5 120.062 C1 C3 F9 119.197
C2 C1 C3 118.970 C2 C1 F7 120.515
C2 C4 C6 120.065 C2 C4 H10 118.771
C3 C1 F7 120.515 C3 C5 C6 120.065
C3 C5 H11 118.771 C4 C2 F8 120.741
C4 C6 C5 120.776 C4 C6 H12 119.612
C5 C3 F9 120.741 C5 C6 H12 119.612
C6 C4 H10 121.164 C6 C5 H11 121.164
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability