return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C5H8 (1,3-Pentadiene, (Z)-)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-191.898099
Energy at 298.15K-191.905247
HF Energy-191.596858
Nuclear repulsion energy153.044567
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3612 3210 0.29      
2 A' 3565 3169 1.76      
3 A' 3520 3129 1.82      
4 A' 3504 3115 2.11      
5 A' 3485 3098 6.51      
6 A' 3455 3071 10.77      
7 A' 3368 2994 2.49      
8 A' 1838 1633 0.66      
9 A' 1770 1573 1.39      
10 A' 1724 1532 2.55      
11 A' 1625 1444 0.55      
12 A' 1605 1426 4.81      
13 A' 1519 1350 9.86      
14 A' 1427 1269 4.44      
15 A' 1378 1225 0.03      
16 A' 1279 1137 1.29      
17 A' 1149 1021 0.61      
18 A' 1045 929 2.00      
19 A' 951 845 0.02      
20 A' 644 573 2.51      
21 A' 398 354 1.13      
22 A' 228 203 0.09      
23 A" 3527 3135 1.80      
24 A" 1722 1531 1.83      
25 A" 1183 1052 0.01      
26 A" 1137 1011 4.68      
27 A" 1082 962 0.35      
28 A" 981 872 11.92      
29 A" 843 749 3.15      
30 A" 672 598 8.83      
31 A" 372 331 0.05      
32 A" 120 107 0.00      
33 A" 94 83 0.13      

Unscaled Zero Point Vibrational Energy (zpe) 27410.3 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 24365.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.50678 0.08547 0.07417

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 2.429 0.127 0.000
C2 1.123 -0.235 0.000
C3 0.000 0.760 0.000
C4 -1.331 0.482 0.000
C5 -1.988 -0.907 0.000
H6 2.732 1.184 0.000
H7 3.234 -0.619 0.000
H8 0.862 -1.304 0.000
H9 0.310 1.817 0.000
H10 -2.034 1.332 0.000
H11 -1.242 -1.718 0.000
H12 -2.631 -1.028 0.891
H13 -2.631 -1.028 -0.891

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 H6 H7 H8 H9 H10 H11 H12 H13
C11.35432.50953.77594.53591.09901.09852.12182.71014.62214.10845.26585.2658
C21.35431.50032.55673.18322.14442.14561.10062.20753.52462.79213.93963.9396
C32.50951.50031.35932.59402.76463.51602.23611.10232.11292.77123.30353.3035
C43.77592.55671.35931.53674.12274.69602.82782.11541.10272.20152.18322.1832
C54.53593.18322.59401.53675.16205.23022.87733.56402.23891.10141.10551.1055
H61.09902.14442.76464.12275.16201.87153.11182.50354.76824.92055.86935.8693
H71.09852.14563.51604.69605.23021.87152.46923.80655.61794.60945.94685.9468
H82.12181.10062.23612.82782.87733.11182.46923.16963.91542.14453.61533.6153
H92.71012.20751.10232.11543.56402.50353.80653.16962.39383.86084.18844.1884
H104.62213.52462.11291.10272.23894.76825.61793.91542.39383.15022.59222.5922
H114.10842.79212.77122.20151.10144.92054.60942.14453.86083.15021.78811.7881
H125.26583.93963.30352.18321.10555.86935.94683.61534.18842.59221.78811.7816
H135.26583.93963.30352.18321.10555.86935.94683.61534.18842.59221.78811.7816

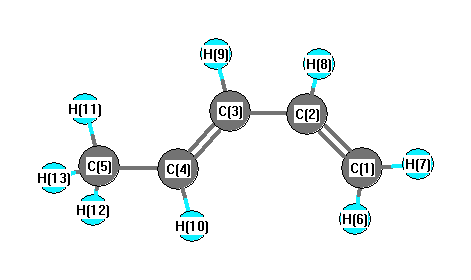
picture of 1,3-Pentadiene, (Z)- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 122.988 C1 C2 H8 119.254
C2 C1 H6 121.529 C2 C1 H7 121.684
C2 C3 C4 126.714 C2 C3 H9 115.176
C3 C2 H8 117.757 C3 C4 C5 127.097
C3 C4 H10 117.848 C4 C3 H9 118.110
C4 C5 H11 112.074 C4 C5 H12 110.377
C4 C5 H13 110.377 C5 C4 H10 115.055
H6 C1 H7 116.787 H11 C5 H12 108.239
H11 C5 H13 108.239 H12 C5 H13 107.382
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability