return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H6O (cis-2-butenal)

using model chemistry: MP2=FULL/STO-3G

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A
Energy calculated at MP2=FULL/STO-3G
 hartrees
Energy at 0K-227.181850
Energy at 298.15K-227.187310
Nuclear repulsion energy151.338851
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/STO-3G
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3553 3158 1.32      
2 A' 3510 3120 1.10      
3 A' 3487 3100 4.85      
4 A' 3364 2990 1.80      
5 A' 3299 2933 50.05      
6 A' 1827 1624 9.21      
7 A' 1726 1535 13.82      
8 A' 1719 1528 15.01      
9 A' 1610 1431 5.52      
10 A' 1502 1335 22.76      
11 A' 1417 1260 0.83      
12 A' 1410 1253 3.26      
13 A' 1230 1093 4.78      
14 A' 1083 962 7.72      
15 A' 957 850 12.83      
16 A' 749 666 14.98      
17 A' 398 354 0.78      
18 A' 180 160 1.44      
19 A" 3530 3138 1.22      
20 A" 1718 1527 1.87      
21 A" 1183 1052 0.11      
22 A" 1113 990 10.38      
23 A" 980 872 0.01      
24 A" 768 683 0.00      
25 A" 252 224 2.44      
26 A" 186 165 0.00      
27 A" 131 117 1.59      

Unscaled Zero Point Vibrational Energy (zpe) 21442.1 cm-1
Scaled (by 0.8889) Zero Point Vibrational Energy (zpe) 19059.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP2=FULL/STO-3G
ABC
0.60814 0.08266 0.07379

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP2=FULL/STO-3G

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -1.525 0.544 0.000
C2 0.000 0.704 0.000
C3 0.830 -0.367 0.000
C4 2.363 -0.289 0.000
O5 -2.120 -0.574 0.000
H6 -2.089 1.515 0.000
H7 0.396 1.732 0.000
H8 0.379 -1.375 0.000
H9 2.704 0.760 0.000
H10 2.778 -0.794 0.892
H11 2.778 -0.794 -0.892

Atom - Atom Distances (Å)
  C1 C2 C3 C4 O5 H6 H7 H8 H9 H10 H11
C11.53302.52413.97611.26651.12362.25802.70294.23454.59344.5934
C21.53301.35442.56362.47532.24131.10112.11322.70503.28003.2800
C32.52411.35441.53552.95663.47292.14261.10432.18702.18532.1853
C43.97612.56361.53554.49194.80442.82072.26141.10331.10541.1054
O51.26652.47532.95664.49192.08953.41232.62405.00514.98334.9833
H61.12362.24133.47294.80442.08952.49453.80074.85305.46095.4609
H72.25801.10112.14262.82073.41232.49453.10652.50503.58483.5848
H82.70292.11321.10432.26142.62403.80073.10653.15632.62462.6246
H94.23452.70502.18701.10335.00514.85302.50503.15631.79291.7929
H104.59343.28002.18531.10544.98335.46093.58482.62461.79291.7835
H114.59343.28002.18531.10544.98335.46093.58482.62461.79291.7835

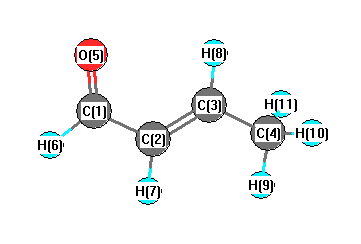
picture of cis-2-butenal state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 121.770 C1 C2 H7 117.063
C2 C1 O5 124.028 C2 C1 H6 114.176
C2 C3 C4 124.899 C2 C3 H8 118.157
C3 C2 H7 121.167 C3 C4 H9 110.889
C3 C4 H10 110.622 C3 C4 H11 110.622
C4 C3 H8 116.944 O5 C1 H6 121.796
H9 C4 H10 108.529 H9 C4 H11 108.529
H10 C4 H11 107.548
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at MP2=FULL/STO-3G Charges (e)
Number Element Mulliken CHELPG AIM ESP
1 C 0.119      
2 C -0.093      
3 C -0.026      
4 C -0.172      
5 O -0.206      
6 H 0.048      
7 H 0.057      
8 H 0.072      
9 H 0.065      
10 H 0.068      
11 H 0.068      


Electric dipole moments


Electric Quadrupole moment
Quadrupole components in D Å


Polarizabilities
Components of the polarizability tensor.
Units are Å3 (Angstrom cubed)
Change units.
  x y z
x 0.000 0.000 0.000
y 0.000 0.000 0.000
z 0.000 0.000 0.000


<r2> (average value of r2) Å2
<r2> 0.000
(<r2>)1/2 0.000